return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4Cl2 (1,4-dichlorobenzene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1AG
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-1149.874133
Energy at 298.15K-1149.878006
HF Energy-1148.587731
Nuclear repulsion energy449.027323
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3243 3072 0.00      
2 Ag 1638 1552 0.00      
3 Ag 1212 1148 0.00      
4 Ag 1138 1078 0.00      
5 Ag 768 727 0.00      
6 Ag 335 317 0.00      
7 Au 875 829 0.00      
8 Au 404 383 0.00      
9 B1g 797 755 0.00      
10 B1u 3229 3058 1.03      
11 B1u 1521 1441 111.45      
12 B1u 1136 1075 94.84      
13 B1u 1024 970 56.90      
14 B1u 569 539 22.67      
15 B2g 850 805 0.00      
16 B2g 409 387 0.00      
17 B2g 289 274 0.00      
18 B2u 3242 3071 1.92      
19 B2u 1470 1392 1.77      
20 B2u 1410 1335 5.68      
21 B2u 1135 1075 4.70      
22 B2u 222 210 0.45      
23 B3g 3230 3059 0.00      
24 B3g 1625 1539 0.00      
25 B3g 1322 1252 0.00      
26 B3g 637 603 0.00      
27 B3g 356 337 0.00      
28 B3u 768 727 69.88      
29 B3u 459 434 21.28      
30 B3u 100 95 0.29      

Unscaled Zero Point Vibrational Energy (zpe) 17705.0 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 16768.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.18890 0.02226 0.01991

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.388
C2 0.000 0.000 -1.388
C3 0.000 1.213 0.698
C4 0.000 -1.213 0.698
C5 0.000 -1.213 -0.698
C6 0.000 1.213 -0.698
Cl7 0.000 0.000 3.122
Cl8 0.000 0.000 -3.122
H9 0.000 2.149 1.246
H10 0.000 -2.149 1.246
H11 0.000 -2.149 -1.246
H12 0.000 2.149 -1.246

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C12.77571.39561.39562.41252.41251.73374.50942.15402.15403.39983.3998
C22.77572.41252.41251.39561.39564.50941.73373.39983.39982.15402.1540
C31.39562.41252.42592.79841.39512.71054.00711.08543.40673.88382.1577
C41.39562.41252.42591.39512.79842.71054.00713.40671.08542.15773.8838
C52.41251.39562.79841.39512.42594.00712.71053.88382.15771.08543.4067
C62.41251.39561.39512.79842.42594.00712.71052.15773.88383.40671.0854
Cl71.73374.50942.71052.71054.00714.00716.24312.85232.85234.86814.8681
Cl84.50941.73374.00714.00712.71052.71056.24314.86814.86812.85232.8523
H92.15403.39981.08543.40673.88382.15772.85234.86814.29864.96912.4928
H102.15403.39983.40671.08542.15773.88382.85234.86814.29862.49284.9691
H113.39982.15403.88382.15771.08543.40674.86812.85234.96912.49284.2986
H123.39982.15402.15773.88383.40671.08544.86812.85232.49284.96914.2986

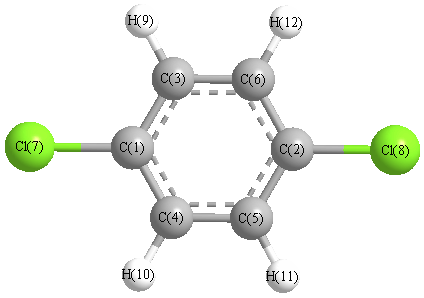
picture of 1,4-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C6 119.646 C1 C3 H9 119.978
C1 C4 C5 119.646 C1 C4 H10 119.978
C2 C5 C4 119.646 C2 C5 H11 119.978
C2 C6 C3 119.646 C2 C6 H12 119.978
C3 C1 C4 120.708 C3 C1 Cl7 119.646
C3 C6 H12 120.376 C4 C1 Cl7 119.646
C4 C5 H11 120.376 C5 C2 C6 120.708
C5 C2 Cl8 119.646 C5 C4 H10 120.376
C6 C2 Cl8 119.646 C6 C3 H9 120.376
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability