return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4O2 (parabenzoquinone)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1Ag
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-380.624115
Energy at 298.15K-380.628353
Nuclear repulsion energy321.858449
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3238 3066 0.00      
2 Ag 1712 1622 0.00      
3 Ag 1650 1563 0.00      
4 Ag 1174 1112 0.00      
5 Ag 785 744 0.00      
6 Ag 450 427 0.00      
7 Au 961 910 0.00      
8 Au 318 302 0.00      
9 B1g 747 707 0.00      
10 B1u 3219 3049 0.94      
11 B1u 1713 1622 160.22      
12 B1u 1389 1315 7.17      
13 B1u 950 900 11.85      
14 B1u 761 720 0.14      
15 B2g 957 907 0.00      
16 B2g 719 681 0.00      
17 B2g 171 162 0.00      
18 B2u 3236 3065 4.81      
19 B2u 1638 1551 21.95      
20 B2u 1344 1273 71.24      
21 B2u 1094 1036 51.16      
22 B2u 409 388 24.15      
23 B3g 3219 3049 0.00      
24 B3g 1422 1347 0.00      
25 B3g 1255 1189 0.00      
26 B3g 605 573 0.00      
27 B3g 445 422 0.00      
28 B3u 892 845 91.07      
29 B3u 500 474 6.78      
30 B3u 88 83 10.93      

Unscaled Zero Point Vibrational Energy (zpe) 18530.9 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 17550.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.17531 0.05490 0.04181

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.446
C2 0.000 0.000 -1.446
C3 0.000 1.266 0.674
C4 0.000 -1.266 0.674
C5 0.000 -1.266 -0.674
C6 0.000 1.266 -0.674
O7 0.000 0.000 2.673
O8 0.000 0.000 -2.673
H9 0.000 2.183 1.257
H10 0.000 -2.183 1.257
H11 0.000 -2.183 -1.257
H12 0.000 2.183 -1.257

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 O8 H9 H10 H11 H12
C12.89281.48291.48292.47002.47001.22704.11982.19162.19163.47503.4750
C22.89282.47002.47001.48291.48294.11981.22703.47503.47502.19162.1916
C31.48292.47002.53212.86901.34892.36623.57921.08673.49833.95342.1382
C41.48292.47002.53211.34892.86902.36623.57923.49831.08672.13823.9534
C52.47001.48292.86901.34892.53213.57922.36623.95342.13821.08673.4983
C62.47001.48291.34892.86902.53213.57922.36622.13823.95343.49831.0867
O71.22704.11982.36622.36623.57923.57925.34682.60252.60254.49624.4962
O84.11981.22703.57923.57922.36622.36625.34684.49624.49622.60252.6025
H92.19163.47501.08673.49833.95342.13822.60254.49624.36675.03872.5141
H102.19163.47503.49831.08672.13823.95342.60254.49624.36672.51415.0387
H113.47502.19163.95342.13821.08673.49834.49622.60255.03872.51414.3667
H123.47502.19162.13823.95343.49831.08674.49622.60252.51415.03874.3667

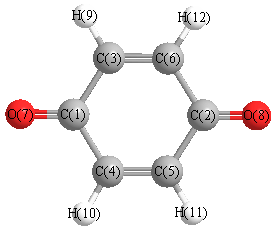
picture of parabenzoquinone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C6 121.372 C1 C3 H9 116.206
C1 C4 C5 121.372 C1 C4 H10 116.206
C2 C5 C4 121.372 C2 C5 H11 116.206
C2 C6 C3 121.372 C2 C6 H12 116.206
C3 C1 C4 117.256 C3 C1 O7 121.372
C3 C6 H12 122.422 C4 C1 O7 121.372
C4 C5 H11 122.422 C5 C2 C6 117.256
C5 C2 O8 121.372 C5 C4 H10 122.422
C6 C2 O8 121.372 C6 C3 H9 122.422
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability