Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -5224.174674 |
| Energy at 298.15K | -5224.184947 |
| HF Energy | -5222.821914 |
| Nuclear repulsion energy | 418.953039 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3149 |
2982 |
0.00 |
|
|
|
| 2 |
Ag |
1521 |
1441 |
0.00 |
|
|
|
| 3 |
Ag |
1350 |
1279 |
0.00 |
|
|
|
| 4 |
Ag |
1099 |
1041 |
0.00 |
|
|
|
| 5 |
Ag |
716 |
678 |
0.00 |
|
|
|
| 6 |
Ag |
199 |
188 |
0.00 |
|
|
|
| 7 |
Au |
3239 |
3067 |
0.81 |
|
|
|
| 8 |
Au |
1160 |
1099 |
6.39 |
|
|
|
| 9 |
Au |
779 |
738 |
5.64 |
|
|
|
| 10 |
Au |
115 |
109 |
4.33 |
|
|
|
| 11 |
Bg |
3218 |
3048 |
0.00 |
|
|
|
| 12 |
Bg |
1331 |
1260 |
0.00 |
|
|
|
| 13 |
Bg |
992 |
940 |
0.00 |
|
|
|
| 14 |
Bu |
3156 |
2989 |
7.83 |
|
|
|
| 15 |
Bu |
1519 |
1438 |
9.17 |
|
|
|
| 16 |
Bu |
1263 |
1197 |
54.89 |
|
|
|
| 17 |
Bu |
640 |
606 |
51.85 |
|
|
|
| 18 |
Bu |
189 |
179 |
6.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12817.2 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 12139.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.491 |
0.573 |
0.000 |
| C2 |
-0.491 |
-0.573 |
0.000 |
| Br3 |
-0.491 |
2.254 |
0.000 |
| Br4 |
0.491 |
-2.254 |
0.000 |
| H5 |
1.116 |
0.567 |
0.891 |
| H6 |
1.116 |
0.567 |
-0.891 |
| H7 |
-1.116 |
-0.567 |
0.891 |
| H8 |
-1.116 |
-0.567 |
-0.891 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5097 | 1.9473 | 2.8272 | 1.0881 | 1.0881 | 2.1629 | 2.1629 |
C2 | 1.5097 | | 2.8272 | 1.9473 | 2.1629 | 2.1629 | 1.0881 | 1.0881 | Br3 | 1.9473 | 2.8272 | | 4.6142 | 2.4950 | 2.4950 | 3.0238 | 3.0238 | Br4 | 2.8272 | 1.9473 | 4.6142 | | 3.0238 | 3.0238 | 2.4950 | 2.4950 | H5 | 1.0881 | 2.1629 | 2.4950 | 3.0238 | | 1.7813 | 2.5044 | 3.0733 | H6 | 1.0881 | 2.1629 | 2.4950 | 3.0238 | 1.7813 | | 3.0733 | 2.5044 | H7 | 2.1629 | 1.0881 | 3.0238 | 2.4950 | 2.5044 | 3.0733 | | 1.7813 | H8 | 2.1629 | 1.0881 | 3.0238 | 2.4950 | 3.0733 | 2.5044 | 1.7813 | |
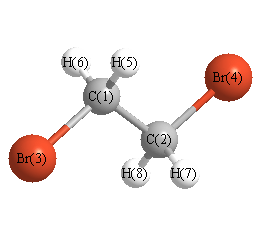 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
109.079 |
|
C1 |
C2 |
H7 |
111.700 |
| C1 |
C2 |
H8 |
111.700 |
|
C2 |
C1 |
Br3 |
109.079 |
| C2 |
C1 |
H5 |
111.700 |
|
C2 |
C1 |
H6 |
111.700 |
| Br3 |
C1 |
H5 |
107.139 |
|
Br3 |
C1 |
H6 |
107.139 |
| Br4 |
C2 |
H7 |
107.139 |
|
Br4 |
C2 |
H8 |
107.139 |
| H5 |
C1 |
H6 |
109.872 |
|
H7 |
C2 |
H8 |
109.872 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -5224.171180 |
| Energy at 298.15K | |
| HF Energy | -5222.817508 |
| Nuclear repulsion energy | 453.538529 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3200 |
3031 |
0.12 |
97.82 |
0.75 |
0.85 |
| 2 |
A |
3132 |
2966 |
16.17 |
196.96 |
0.02 |
0.04 |
| 3 |
A |
1506 |
1426 |
0.85 |
4.75 |
0.61 |
0.76 |
| 4 |
A |
1362 |
1290 |
23.02 |
3.48 |
0.70 |
0.82 |
| 5 |
A |
1238 |
1172 |
2.57 |
13.81 |
0.75 |
0.85 |
| 6 |
A |
1074 |
1017 |
0.08 |
2.73 |
0.72 |
0.84 |
| 7 |
A |
948 |
898 |
7.82 |
7.46 |
0.45 |
0.62 |
| 8 |
A |
590 |
559 |
6.45 |
15.67 |
0.08 |
0.15 |
| 9 |
A |
241 |
228 |
0.98 |
1.49 |
0.53 |
0.69 |
| 10 |
A |
87 |
82 |
0.20 |
1.31 |
0.73 |
0.84 |
| 11 |
B |
3211 |
3041 |
1.46 |
29.02 |
0.75 |
0.86 |
| 12 |
B |
3125 |
2960 |
2.80 |
45.72 |
0.75 |
0.86 |
| 13 |
B |
1501 |
1421 |
14.58 |
13.87 |
0.75 |
0.86 |
| 14 |
B |
1338 |
1267 |
55.71 |
1.17 |
0.75 |
0.86 |
| 15 |
B |
1167 |
1105 |
4.38 |
5.76 |
0.75 |
0.86 |
| 16 |
B |
881 |
834 |
19.52 |
1.58 |
0.75 |
0.86 |
| 17 |
B |
628 |
594 |
10.51 |
9.87 |
0.75 |
0.86 |
| 18 |
B |
371 |
352 |
5.16 |
2.36 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12798.1 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 12121.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.309 |
0.690 |
1.180 |
| C2 |
-0.309 |
-0.690 |
1.180 |
| Br3 |
-0.309 |
1.789 |
-0.294 |
| Br4 |
0.309 |
-1.789 |
-0.294 |
| H5 |
0.024 |
1.212 |
2.094 |
| H6 |
1.395 |
0.641 |
1.111 |
| H7 |
-0.024 |
-1.212 |
2.094 |
| H8 |
-1.395 |
-0.641 |
1.111 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5112 | 1.9394 | 2.8834 | 1.0913 | 1.0888 | 2.1363 | 2.1629 |
C2 | 1.5112 | | 2.8834 | 1.9394 | 2.1363 | 2.1629 | 1.0913 | 1.0888 | Br3 | 1.9394 | 2.8834 | | 3.6311 | 2.4790 | 2.4883 | 3.8459 | 3.0098 | Br4 | 2.8834 | 1.9394 | 3.6311 | | 3.8459 | 3.0098 | 2.4790 | 2.4883 | H5 | 1.0913 | 2.1363 | 2.4790 | 3.8459 | | 1.7809 | 2.4248 | 2.5326 | H6 | 1.0888 | 2.1629 | 2.4883 | 3.0098 | 1.7809 | | 2.5326 | 3.0700 | H7 | 2.1363 | 1.0913 | 3.8459 | 2.4790 | 2.4248 | 2.5326 | | 1.7809 | H8 | 2.1629 | 1.0888 | 3.0098 | 2.4883 | 2.5326 | 3.0700 | 1.7809 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
112.776 |
|
C1 |
C2 |
H7 |
109.282 |
| C1 |
C2 |
H8 |
111.549 |
|
C2 |
C1 |
Br3 |
112.776 |
| C2 |
C1 |
H5 |
109.282 |
|
C2 |
C1 |
H6 |
111.549 |
| Br3 |
C1 |
H5 |
106.374 |
|
Br3 |
C1 |
H6 |
107.144 |
| Br4 |
C2 |
H7 |
106.374 |
|
Br4 |
C2 |
H8 |
107.144 |
| H5 |
C1 |
H6 |
109.546 |
|
H7 |
C2 |
H8 |
109.546 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
