Jump to
S1C2
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -5226.153762 |
| Energy at 298.15K | |
| HF Energy | -5225.736679 |
| Nuclear repulsion energy | 415.649655 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3140 |
3140 |
0.00 |
148.07 |
0.07 |
0.14 |
| 2 |
Ag |
1517 |
1517 |
0.00 |
13.98 |
0.72 |
0.84 |
| 3 |
Ag |
1326 |
1326 |
0.00 |
34.08 |
0.51 |
0.68 |
| 4 |
Ag |
1080 |
1080 |
0.00 |
15.73 |
0.67 |
0.80 |
| 5 |
Ag |
675 |
675 |
0.00 |
95.19 |
0.28 |
0.44 |
| 6 |
Ag |
191 |
191 |
0.00 |
4.69 |
0.35 |
0.51 |
| 7 |
Au |
3224 |
3224 |
2.13 |
0.00 |
0.65 |
0.79 |
| 8 |
Au |
1138 |
1138 |
5.84 |
0.00 |
0.70 |
0.82 |
| 9 |
Au |
774 |
774 |
5.60 |
0.00 |
0.00 |
0.00 |
| 10 |
Au |
108 |
108 |
4.49 |
0.00 |
0.00 |
0.00 |
| 11 |
Bg |
3202 |
3202 |
0.00 |
93.30 |
0.75 |
0.86 |
| 12 |
Bg |
1319 |
1319 |
0.00 |
4.43 |
0.75 |
0.86 |
| 13 |
Bg |
973 |
973 |
0.00 |
6.75 |
0.75 |
0.86 |
| 14 |
Bu |
3148 |
3148 |
9.72 |
0.00 |
0.00 |
0.00 |
| 15 |
Bu |
1514 |
1514 |
8.25 |
0.00 |
0.00 |
0.00 |
| 16 |
Bu |
1247 |
1247 |
64.76 |
0.00 |
0.00 |
0.00 |
| 17 |
Bu |
599 |
599 |
71.46 |
0.00 |
0.00 |
0.00 |
| 18 |
Bu |
183 |
183 |
6.97 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 12679.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12679.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.494 |
0.570 |
0.000 |
| C2 |
-0.494 |
-0.570 |
0.000 |
| Br3 |
-0.494 |
2.275 |
0.000 |
| Br4 |
0.494 |
-2.275 |
0.000 |
| H5 |
1.116 |
0.574 |
0.890 |
| H6 |
1.116 |
0.574 |
-0.890 |
| H7 |
-1.116 |
-0.574 |
0.890 |
| H8 |
-1.116 |
-0.574 |
-0.890 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5085 | 1.9697 | 2.8448 | 1.0856 | 1.0856 | 2.1659 | 2.1659 |
C2 | 1.5085 | | 2.8448 | 1.9697 | 2.1659 | 2.1659 | 1.0856 | 1.0856 | Br3 | 1.9697 | 2.8448 | | 4.6551 | 2.5047 | 2.5047 | 3.0485 | 3.0485 | Br4 | 2.8448 | 1.9697 | 4.6551 | | 3.0485 | 3.0485 | 2.5047 | 2.5047 | H5 | 1.0856 | 2.1659 | 2.5047 | 3.0485 | | 1.7794 | 2.5095 | 3.0764 | H6 | 1.0856 | 2.1659 | 2.5047 | 3.0485 | 1.7794 | | 3.0764 | 2.5095 | H7 | 2.1659 | 1.0856 | 3.0485 | 2.5047 | 2.5095 | 3.0764 | | 1.7794 | H8 | 2.1659 | 1.0856 | 3.0485 | 2.5047 | 3.0764 | 2.5095 | 1.7794 | |
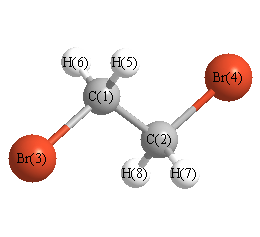 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
109.033 |
|
C1 |
C2 |
H7 |
112.191 |
| C1 |
C2 |
H8 |
112.191 |
|
C2 |
C1 |
Br3 |
109.033 |
| C2 |
C1 |
H5 |
112.191 |
|
C2 |
C1 |
H6 |
112.191 |
| Br3 |
C1 |
H5 |
106.512 |
|
Br3 |
C1 |
H6 |
106.512 |
| Br4 |
C2 |
H7 |
106.512 |
|
Br4 |
C2 |
H8 |
106.512 |
| H5 |
C1 |
H6 |
110.076 |
|
H7 |
C2 |
H8 |
110.076 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -5226.149813 |
| Energy at 298.15K | |
| HF Energy | -5225.732488 |
| Nuclear repulsion energy | 448.954449 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3181 |
3181 |
0.15 |
102.22 |
0.75 |
0.86 |
| 2 |
A |
3120 |
3120 |
19.96 |
216.91 |
0.02 |
0.04 |
| 3 |
A |
1500 |
1500 |
0.76 |
4.68 |
0.63 |
0.77 |
| 4 |
A |
1345 |
1345 |
26.26 |
4.04 |
0.63 |
0.77 |
| 5 |
A |
1223 |
1223 |
3.21 |
14.56 |
0.75 |
0.86 |
| 6 |
A |
1053 |
1053 |
0.32 |
2.64 |
0.68 |
0.81 |
| 7 |
A |
928 |
928 |
9.01 |
8.63 |
0.44 |
0.61 |
| 8 |
A |
561 |
561 |
8.75 |
17.65 |
0.08 |
0.15 |
| 9 |
A |
233 |
233 |
1.11 |
1.71 |
0.48 |
0.64 |
| 10 |
A |
81 |
81 |
0.23 |
1.47 |
0.73 |
0.84 |
| 11 |
B |
3194 |
3194 |
3.16 |
28.13 |
0.75 |
0.86 |
| 12 |
B |
3111 |
3111 |
2.92 |
53.40 |
0.75 |
0.86 |
| 13 |
B |
1494 |
1494 |
12.92 |
14.47 |
0.75 |
0.86 |
| 14 |
B |
1317 |
1317 |
71.92 |
1.91 |
0.75 |
0.86 |
| 15 |
B |
1151 |
1151 |
4.13 |
5.82 |
0.75 |
0.86 |
| 16 |
B |
867 |
867 |
21.44 |
1.55 |
0.75 |
0.86 |
| 17 |
B |
597 |
597 |
16.16 |
11.46 |
0.75 |
0.86 |
| 18 |
B |
361 |
361 |
6.46 |
2.82 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12656.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12656.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.311 |
0.688 |
1.181 |
| C2 |
-0.311 |
-0.688 |
1.181 |
| Br3 |
-0.311 |
1.816 |
-0.294 |
| Br4 |
0.311 |
-1.816 |
-0.294 |
| H5 |
0.030 |
1.212 |
2.094 |
| H6 |
1.394 |
0.645 |
1.106 |
| H7 |
-0.030 |
-1.212 |
2.094 |
| H8 |
-1.394 |
-0.645 |
1.106 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5106 | 1.9582 | 2.9067 | 1.0893 | 1.0864 | 2.1356 | 2.1659 |
C2 | 1.5106 | | 2.9067 | 1.9582 | 2.1356 | 2.1659 | 1.0893 | 1.0864 | Br3 | 1.9582 | 2.9067 | | 3.6858 | 2.4865 | 2.4973 | 3.8668 | 3.0319 | Br4 | 2.9067 | 1.9582 | 3.6858 | | 3.8668 | 3.0319 | 2.4865 | 2.4973 | H5 | 1.0893 | 2.1356 | 2.4865 | 3.8668 | | 1.7771 | 2.4249 | 2.5406 | H6 | 1.0864 | 2.1659 | 2.4973 | 3.0319 | 1.7771 | | 2.5406 | 3.0722 | H7 | 2.1356 | 1.0893 | 3.8668 | 2.4865 | 2.4249 | 2.5406 | | 1.7771 | H8 | 2.1659 | 1.0864 | 3.0319 | 2.4973 | 2.5406 | 3.0722 | 1.7771 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
113.221 |
|
C1 |
C2 |
H7 |
109.388 |
| C1 |
C2 |
H8 |
111.993 |
|
C2 |
C1 |
Br3 |
113.221 |
| C2 |
C1 |
H5 |
109.388 |
|
C2 |
C1 |
H6 |
111.993 |
| Br3 |
C1 |
H5 |
105.795 |
|
Br3 |
C1 |
H6 |
106.681 |
| Br4 |
C2 |
H7 |
105.795 |
|
Br4 |
C2 |
H8 |
106.681 |
| H5 |
C1 |
H6 |
109.525 |
|
H7 |
C2 |
H8 |
109.525 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
