return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CH2Br (n-propyl bromide)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-2691.068823
Energy at 298.15K-2691.079335
HF Energy-2690.075154
Nuclear repulsion energy236.729107
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3173 3006 23.40      
2 A' 3129 2964 19.68      
3 A' 3102 2938 8.48      
4 A' 3080 2917 21.72      
5 A' 1547 1465 10.18      
6 A' 1531 1450 1.58      
7 A' 1519 1439 2.52      
8 A' 1450 1373 5.52      
9 A' 1399 1325 11.15      
10 A' 1298 1229 32.08      
11 A' 1145 1084 2.40      
12 A' 1078 1021 1.57      
13 A' 930 881 9.59      
14 A' 694 657 17.65      
15 A' 321 304 1.10      
16 A' 221 209 1.29      
17 A" 3202 3032 14.19      
18 A" 3171 3003 29.03      
19 A" 3149 2982 0.02      
20 A" 1541 1459 11.59      
21 A" 1356 1284 0.00      
22 A" 1277 1210 0.68      
23 A" 1100 1042 3.36      
24 A" 880 833 0.00      
25 A" 758 718 4.90      
26 A" 244 231 0.05      
27 A" 120 114 0.76      

Unscaled Zero Point Vibrational Energy (zpe) 21207.9 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 20086.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.84890 0.05479 0.05300

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.831 0.000
C2 1.506 0.655 0.000
C3 2.212 2.010 0.000
Br4 -0.926 -0.883 0.000
H5 -0.342 1.362 0.888
H6 -0.342 1.362 -0.888
H7 1.801 0.075 0.879
H8 1.801 0.075 -0.879
H9 3.297 1.882 0.000
H10 1.946 2.595 -0.884
H11 1.946 2.595 0.884

Atom - Atom Distances (Å)
  C1 C2 C3 Br4 H5 H6 H7 H8 H9 H10 H11
C11.51662.50681.94841.09021.09022.14182.14183.46012.77162.7716
C21.51661.52782.87822.16962.16961.09341.09342.17032.17692.1769
C32.50681.52784.26862.78132.78132.16482.16481.09211.09301.0930
Br41.94842.87824.26862.48422.48423.02123.02125.04764.59704.5970
H51.09022.16962.78132.48421.77682.50033.06173.78203.14652.5998
H61.09022.16962.78132.48421.77683.06172.50033.78202.59983.1465
H72.14181.09342.16483.02122.50033.06171.75722.50483.07902.5245
H82.14181.09342.16483.02123.06172.50031.75722.50482.52453.0790
H93.46012.17031.09215.04763.78203.78202.50482.50481.76471.7647
H102.77162.17691.09304.59703.14652.59983.07902.52451.76471.7681
H112.77162.17691.09304.59702.59983.14652.52453.07901.76471.7681

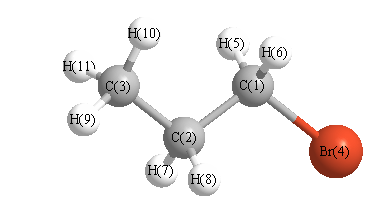
picture of n-propyl bromide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 110.857 C1 C2 H7 109.223
C1 C2 H8 109.223 C2 C1 Br4 111.723
C2 C1 H5 111.625 C2 C1 H6 111.625
C2 C3 H9 110.769 C2 C3 H10 111.245
C2 C3 H11 111.245 C3 C2 H7 110.251
C3 C2 H8 110.251 Br4 C1 H5 106.214
Br4 C1 H6 106.214 H5 C1 H6 109.151
H7 C2 H8 106.946 H9 C3 H10 107.730
H9 C3 H11 107.730 H10 C3 H11 107.966
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability