Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -157.960702 |
| Energy at 298.15K | -157.971403 |
| HF Energy | -157.325399 |
| Nuclear repulsion energy | 130.743910 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
120 |
114 |
0.00 |
|
|
|
|
Au |
233 |
221 |
0.01 |
|
|
|
|
Bu |
265 |
251 |
0.02 |
|
|
|
|
Bg |
266 |
252 |
0.00 |
|
|
|
|
Ag |
435 |
412 |
0.00 |
|
|
|
|
Au |
741 |
701 |
4.01 |
|
|
|
|
Bg |
827 |
783 |
0.00 |
|
|
|
|
Ag |
868 |
822 |
0.00 |
|
|
|
|
Au |
982 |
930 |
1.10 |
|
|
|
|
Bu |
996 |
943 |
8.27 |
|
|
|
|
Bu |
1057 |
1001 |
0.16 |
|
|
|
|
Ag |
1109 |
1050 |
0.00 |
|
|
|
|
Ag |
1196 |
1133 |
0.00 |
|
|
|
|
Bg |
1240 |
1174 |
0.00 |
|
|
|
|
Au |
1321 |
1251 |
0.16 |
|
|
|
|
Bu |
1341 |
1271 |
0.39 |
|
|
|
|
Bg |
1364 |
1291 |
0.00 |
|
|
|
|
Ag |
1425 |
1350 |
0.00 |
|
|
|
|
Ag |
1447 |
1371 |
0.00 |
|
|
|
|
Bu |
1450 |
1373 |
11.99 |
|
|
|
|
Ag |
1527 |
1446 |
0.00 |
|
|
|
|
Bu |
1530 |
1449 |
2.83 |
|
|
|
|
Bg |
1541 |
1459 |
0.00 |
|
|
|
|
Au |
1542 |
1461 |
20.19 |
|
|
|
|
Ag |
1544 |
1462 |
0.00 |
|
|
|
|
Bu |
1550 |
1468 |
13.65 |
|
|
|
|
Ag |
3067 |
2905 |
0.00 |
|
|
|
|
Bu |
3073 |
2911 |
22.58 |
|
|
|
|
Ag |
3076 |
2913 |
0.00 |
|
|
|
|
Bu |
3076 |
2914 |
93.62 |
|
|
|
|
Bg |
3107 |
2942 |
0.00 |
|
|
|
|
Au |
3128 |
2962 |
21.14 |
|
|
|
|
Bg |
3162 |
2995 |
0.00 |
|
|
|
|
Ag |
3165 |
2997 |
0.00 |
|
|
|
|
Au |
3165 |
2997 |
98.58 |
|
|
|
|
Bu |
3165 |
2998 |
66.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29549.0 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 27985.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.422 |
0.636 |
0.000 |
| C2 |
0.422 |
-0.636 |
0.000 |
| C3 |
0.422 |
1.908 |
0.000 |
| C4 |
-0.422 |
-1.908 |
0.000 |
| H5 |
-1.079 |
0.630 |
0.878 |
| H6 |
-1.079 |
0.630 |
-0.878 |
| H7 |
1.079 |
-0.630 |
0.878 |
| H8 |
1.079 |
-0.630 |
-0.878 |
| H9 |
-0.202 |
2.805 |
0.000 |
| H10 |
1.065 |
1.949 |
0.883 |
| H11 |
1.065 |
1.949 |
-0.883 |
| H12 |
0.202 |
-2.805 |
0.000 |
| H13 |
-1.065 |
-1.949 |
0.883 |
| H14 |
-1.065 |
-1.949 |
-0.883 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5264 | 1.5261 | 2.5437 | 1.0963 | 1.0963 | 2.1504 | 2.1504 | 2.1797 | 2.1713 | 2.1713 | 3.4969 | 2.8061 | 2.8061 |
C2 | 1.5264 | | 2.5437 | 1.5261 | 2.1504 | 2.1504 | 1.0963 | 1.0963 | 3.4969 | 2.8061 | 2.8061 | 2.1797 | 2.1713 | 2.1713 | C3 | 1.5261 | 2.5437 | | 3.9076 | 2.1577 | 2.1577 | 2.7641 | 2.7641 | 1.0928 | 1.0934 | 1.0934 | 4.7175 | 4.2265 | 4.2265 | C4 | 2.5437 | 1.5261 | 3.9076 | | 2.7641 | 2.7641 | 2.1577 | 2.1577 | 4.7175 | 4.2265 | 4.2265 | 1.0928 | 1.0934 | 1.0934 | H5 | 1.0963 | 2.1504 | 2.1577 | 2.7641 | | 1.7552 | 2.4983 | 3.0533 | 2.5038 | 2.5175 | 3.0720 | 3.7692 | 2.5784 | 3.1221 | H6 | 1.0963 | 2.1504 | 2.1577 | 2.7641 | 1.7552 | | 3.0533 | 2.4983 | 2.5038 | 3.0720 | 2.5175 | 3.7692 | 3.1221 | 2.5784 | H7 | 2.1504 | 1.0963 | 2.7641 | 2.1577 | 2.4983 | 3.0533 | | 1.7552 | 3.7692 | 2.5784 | 3.1221 | 2.5038 | 2.5175 | 3.0720 | H8 | 2.1504 | 1.0963 | 2.7641 | 2.1577 | 3.0533 | 2.4983 | 1.7552 | | 3.7692 | 3.1221 | 2.5784 | 2.5038 | 3.0720 | 2.5175 | H9 | 2.1797 | 3.4969 | 1.0928 | 4.7175 | 2.5038 | 2.5038 | 3.7692 | 3.7692 | | 1.7663 | 1.7663 | 5.6239 | 4.9111 | 4.9111 | H10 | 2.1713 | 2.8061 | 1.0934 | 4.2265 | 2.5175 | 3.0720 | 2.5784 | 3.1221 | 1.7663 | | 1.7659 | 4.9111 | 4.4418 | 4.7800 | H11 | 2.1713 | 2.8061 | 1.0934 | 4.2265 | 3.0720 | 2.5175 | 3.1221 | 2.5784 | 1.7663 | 1.7659 | | 4.9111 | 4.7800 | 4.4418 | H12 | 3.4969 | 2.1797 | 4.7175 | 1.0928 | 3.7692 | 3.7692 | 2.5038 | 2.5038 | 5.6239 | 4.9111 | 4.9111 | | 1.7663 | 1.7663 | H13 | 2.8061 | 2.1713 | 4.2265 | 1.0934 | 2.5784 | 3.1221 | 2.5175 | 3.0720 | 4.9111 | 4.4418 | 4.7800 | 1.7663 | | 1.7659 | H14 | 2.8061 | 2.1713 | 4.2265 | 1.0934 | 3.1221 | 2.5784 | 3.0720 | 2.5175 | 4.9111 | 4.7800 | 4.4418 | 1.7663 | 1.7659 | |
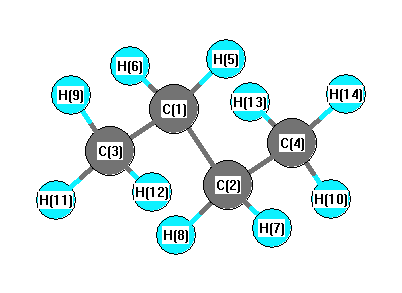 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
112.885 |
|
C1 |
C2 |
H7 |
109.052 |
| C1 |
C2 |
H8 |
109.052 |
|
C1 |
C3 |
H9 |
111.599 |
| C1 |
C3 |
H11 |
110.892 |
|
C1 |
C3 |
H12 |
30.895 |
| C2 |
C1 |
C3 |
112.885 |
|
C2 |
C1 |
H5 |
109.052 |
| C2 |
C1 |
H6 |
109.052 |
|
C2 |
C4 |
H10 |
17.280 |
| C2 |
C4 |
H13 |
110.892 |
|
C2 |
C4 |
H14 |
110.892 |
| C3 |
C1 |
H5 |
109.641 |
|
C3 |
C1 |
H6 |
109.641 |
| C4 |
C2 |
H7 |
109.641 |
|
C4 |
C2 |
H8 |
109.641 |
| H5 |
C1 |
H6 |
106.356 |
|
H7 |
C2 |
H8 |
106.356 |
| H9 |
C3 |
H11 |
107.791 |
|
H9 |
C3 |
H12 |
142.494 |
| H10 |
C4 |
H13 |
94.163 |
|
H10 |
C4 |
H14 |
114.205 |
| H11 |
C3 |
H12 |
93.714 |
|
H13 |
C4 |
H14 |
107.712 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -157.959826 |
| Energy at 298.15K | |
| Nuclear repulsion energy | 132.774322 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3177 |
3009 |
29.69 |
|
|
|
| 2 |
A |
3161 |
2994 |
0.11 |
|
|
|
| 3 |
A |
3116 |
2951 |
21.88 |
|
|
|
| 4 |
A |
3079 |
2916 |
21.36 |
|
|
|
| 5 |
A |
3072 |
2909 |
27.16 |
|
|
|
| 6 |
A |
1557 |
1474 |
2.17 |
|
|
|
| 7 |
A |
1546 |
1464 |
6.41 |
|
|
|
| 8 |
A |
1528 |
1447 |
0.11 |
|
|
|
| 9 |
A |
1451 |
1374 |
8.04 |
|
|
|
| 10 |
A |
1408 |
1334 |
0.10 |
|
|
|
| 11 |
A |
1341 |
1270 |
0.62 |
|
|
|
| 12 |
A |
1226 |
1161 |
0.05 |
|
|
|
| 13 |
A |
1128 |
1068 |
0.14 |
|
|
|
| 14 |
A |
1016 |
962 |
0.16 |
|
|
|
| 15 |
A |
858 |
813 |
0.02 |
|
|
|
| 16 |
A |
817 |
774 |
2.05 |
|
|
|
| 17 |
A |
323 |
306 |
0.01 |
|
|
|
| 18 |
A |
290 |
275 |
0.03 |
|
|
|
| 19 |
A |
121 |
114 |
0.01 |
|
|
|
| 20 |
B |
3169 |
3002 |
48.83 |
|
|
|
| 21 |
B |
3162 |
2994 |
73.57 |
|
|
|
| 22 |
B |
3123 |
2958 |
26.77 |
|
|
|
| 23 |
B |
3076 |
2914 |
22.36 |
|
|
|
| 24 |
B |
3070 |
2907 |
28.18 |
|
|
|
| 25 |
B |
1548 |
1466 |
20.12 |
|
|
|
| 26 |
B |
1541 |
1459 |
10.01 |
|
|
|
| 27 |
B |
1526 |
1446 |
0.40 |
|
|
|
| 28 |
B |
1451 |
1374 |
8.44 |
|
|
|
| 29 |
B |
1398 |
1324 |
0.30 |
|
|
|
| 30 |
B |
1318 |
1249 |
0.21 |
|
|
|
| 31 |
B |
1179 |
1117 |
2.62 |
|
|
|
| 32 |
B |
1002 |
949 |
1.02 |
|
|
|
| 33 |
B |
985 |
933 |
3.92 |
|
|
|
| 34 |
B |
758 |
718 |
4.72 |
|
|
|
| 35 |
B |
449 |
426 |
0.41 |
|
|
|
| 36 |
B |
217 |
206 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29593.3 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 28027.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
