Jump to
S1C2
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -158.292709 |
| Energy at 298.15K | -158.303371 |
| HF Energy | -158.085904 |
| Nuclear repulsion energy | 130.554621 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
123 |
123 |
0.01 |
|
|
|
|
Au |
225 |
225 |
0.00 |
|
|
|
|
Bg |
261 |
261 |
0.00 |
|
|
|
|
Bu |
262 |
262 |
0.01 |
|
|
|
|
Ag |
430 |
430 |
0.00 |
|
|
|
|
Au |
742 |
742 |
3.40 |
|
|
|
|
Bg |
825 |
825 |
0.00 |
|
|
|
|
Ag |
855 |
855 |
0.00 |
|
|
|
|
Au |
980 |
980 |
0.95 |
|
|
|
|
Bu |
993 |
993 |
6.60 |
|
|
|
|
Bu |
1036 |
1036 |
0.18 |
|
|
|
|
Ag |
1084 |
1084 |
0.00 |
|
|
|
|
Ag |
1187 |
1187 |
0.00 |
|
|
|
|
Bg |
1231 |
1231 |
0.00 |
|
|
|
|
Au |
1316 |
1316 |
0.19 |
|
|
|
|
Bu |
1343 |
1343 |
1.45 |
|
|
|
|
Bg |
1360 |
1360 |
0.00 |
|
|
|
|
Ag |
1420 |
1420 |
0.00 |
|
|
|
|
Ag |
1441 |
1441 |
0.00 |
|
|
|
|
Bu |
1444 |
1444 |
8.10 |
|
|
|
|
Ag |
1519 |
1519 |
0.00 |
|
|
|
|
Bu |
1525 |
1525 |
2.27 |
|
|
|
|
Bg |
1534 |
1534 |
0.00 |
|
|
|
|
Au |
1536 |
1536 |
17.72 |
|
|
|
|
Ag |
1539 |
1539 |
0.00 |
|
|
|
|
Bu |
1545 |
1545 |
11.12 |
|
|
|
|
Ag |
3038 |
3038 |
0.00 |
|
|
|
|
Bu |
3046 |
3046 |
45.35 |
|
|
|
|
Bu |
3052 |
3052 |
93.08 |
|
|
|
|
Ag |
3053 |
3053 |
0.00 |
|
|
|
|
Bg |
3062 |
3062 |
0.00 |
|
|
|
|
Au |
3084 |
3084 |
21.97 |
|
|
|
|
Bg |
3119 |
3119 |
0.00 |
|
|
|
|
Au |
3123 |
3123 |
137.39 |
|
|
|
|
Ag |
3123 |
3123 |
0.00 |
|
|
|
|
Bu |
3124 |
3124 |
93.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29289.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 29289.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.421 |
0.638 |
0.000 |
| C2 |
0.421 |
-0.638 |
0.000 |
| C3 |
0.421 |
1.913 |
0.000 |
| C4 |
-0.421 |
-1.913 |
0.000 |
| H5 |
-1.080 |
0.634 |
0.876 |
| H6 |
-1.080 |
0.634 |
-0.876 |
| H7 |
1.080 |
-0.634 |
0.876 |
| H8 |
1.080 |
-0.634 |
-0.876 |
| H9 |
-0.205 |
2.808 |
0.000 |
| H10 |
1.066 |
1.960 |
0.882 |
| H11 |
1.066 |
1.960 |
-0.882 |
| H12 |
0.205 |
-2.808 |
0.000 |
| H13 |
-1.066 |
-1.960 |
0.882 |
| H14 |
-1.066 |
-1.960 |
-0.882 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5295 | 1.5283 | 2.5515 | 1.0957 | 1.0957 | 2.1540 | 2.1540 | 2.1807 | 2.1759 | 2.1759 | 3.5028 | 2.8181 | 2.8181 |
C2 | 1.5295 | | 2.5515 | 1.5283 | 2.1540 | 2.1540 | 1.0957 | 1.0957 | 3.5028 | 2.8181 | 2.8181 | 2.1807 | 2.1759 | 2.1759 | C3 | 1.5283 | 2.5515 | | 3.9182 | 2.1580 | 2.1580 | 2.7730 | 2.7730 | 1.0922 | 1.0930 | 1.0930 | 4.7263 | 4.2413 | 4.2413 | C4 | 2.5515 | 1.5283 | 3.9182 | | 2.7730 | 2.7730 | 2.1580 | 2.1580 | 4.7263 | 4.2413 | 4.2413 | 1.0922 | 1.0930 | 1.0930 | H5 | 1.0957 | 2.1540 | 2.1580 | 2.7730 | | 1.7510 | 2.5050 | 3.0563 | 2.5018 | 2.5222 | 3.0739 | 3.7771 | 2.5939 | 3.1330 | H6 | 1.0957 | 2.1540 | 2.1580 | 2.7730 | 1.7510 | | 3.0563 | 2.5050 | 2.5018 | 3.0739 | 2.5222 | 3.7771 | 3.1330 | 2.5939 | H7 | 2.1540 | 1.0957 | 2.7730 | 2.1580 | 2.5050 | 3.0563 | | 1.7510 | 3.7771 | 2.5939 | 3.1330 | 2.5018 | 2.5222 | 3.0739 | H8 | 2.1540 | 1.0957 | 2.7730 | 2.1580 | 3.0563 | 2.5050 | 1.7510 | | 3.7771 | 3.1330 | 2.5939 | 2.5018 | 3.0739 | 2.5222 | H9 | 2.1807 | 3.5028 | 1.0922 | 4.7263 | 2.5018 | 2.5018 | 3.7771 | 3.7771 | | 1.7638 | 1.7638 | 5.6312 | 4.9245 | 4.9245 | H10 | 2.1759 | 2.8181 | 1.0930 | 4.2413 | 2.5222 | 3.0739 | 2.5939 | 3.1330 | 1.7638 | | 1.7633 | 4.9245 | 4.4614 | 4.7973 | H11 | 2.1759 | 2.8181 | 1.0930 | 4.2413 | 3.0739 | 2.5222 | 3.1330 | 2.5939 | 1.7638 | 1.7633 | | 4.9245 | 4.7973 | 4.4614 | H12 | 3.5028 | 2.1807 | 4.7263 | 1.0922 | 3.7771 | 3.7771 | 2.5018 | 2.5018 | 5.6312 | 4.9245 | 4.9245 | | 1.7638 | 1.7638 | H13 | 2.8181 | 2.1759 | 4.2413 | 1.0930 | 2.5939 | 3.1330 | 2.5222 | 3.0739 | 4.9245 | 4.4614 | 4.7973 | 1.7638 | | 1.7633 | H14 | 2.8181 | 2.1759 | 4.2413 | 1.0930 | 3.1330 | 2.5939 | 3.0739 | 2.5222 | 4.9245 | 4.7973 | 4.4614 | 1.7638 | 1.7633 | |
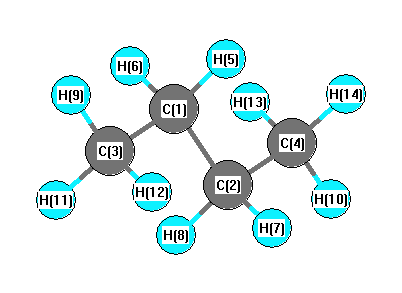 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
113.112 |
|
C1 |
C2 |
H7 |
109.153 |
| C1 |
C2 |
H8 |
109.153 |
|
C1 |
C3 |
H9 |
111.562 |
| C1 |
C3 |
H11 |
111.131 |
|
C1 |
C3 |
H12 |
30.834 |
| C2 |
C1 |
C3 |
113.112 |
|
C2 |
C1 |
H5 |
109.153 |
| C2 |
C1 |
H6 |
109.153 |
|
C2 |
C4 |
H10 |
17.229 |
| C2 |
C4 |
H13 |
111.131 |
|
C2 |
C4 |
H14 |
111.131 |
| C3 |
C1 |
H5 |
109.550 |
|
C3 |
C1 |
H6 |
109.550 |
| C4 |
C2 |
H7 |
109.550 |
|
C4 |
C2 |
H8 |
109.550 |
| H5 |
C1 |
H6 |
106.079 |
|
H7 |
C2 |
H8 |
106.079 |
| H9 |
C3 |
H11 |
107.644 |
|
H9 |
C3 |
H12 |
142.396 |
| H10 |
C4 |
H13 |
94.464 |
|
H10 |
C4 |
H14 |
114.405 |
| H11 |
C3 |
H12 |
93.985 |
|
H13 |
C4 |
H14 |
107.539 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -158.291523 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3136 |
3136 |
46.20 |
|
|
|
| 2 |
A |
3120 |
3120 |
0.63 |
|
|
|
| 3 |
A |
3076 |
3076 |
31.07 |
|
|
|
| 4 |
A |
3057 |
3057 |
27.26 |
|
|
|
| 5 |
A |
3042 |
3042 |
23.92 |
|
|
|
| 6 |
A |
1549 |
1549 |
1.91 |
|
|
|
| 7 |
A |
1540 |
1540 |
5.92 |
|
|
|
| 8 |
A |
1521 |
1521 |
0.00 |
|
|
|
| 9 |
A |
1445 |
1445 |
5.29 |
|
|
|
| 10 |
A |
1402 |
1402 |
0.40 |
|
|
|
| 11 |
A |
1337 |
1337 |
0.39 |
|
|
|
| 12 |
A |
1217 |
1217 |
0.07 |
|
|
|
| 13 |
A |
1105 |
1105 |
0.12 |
|
|
|
| 14 |
A |
1010 |
1010 |
0.24 |
|
|
|
| 15 |
A |
846 |
846 |
0.03 |
|
|
|
| 16 |
A |
806 |
806 |
1.70 |
|
|
|
| 17 |
A |
323 |
323 |
0.01 |
|
|
|
| 18 |
A |
275 |
275 |
0.02 |
|
|
|
| 19 |
A |
113 |
113 |
0.01 |
|
|
|
| 20 |
B |
3128 |
3128 |
64.66 |
|
|
|
| 21 |
B |
3122 |
3122 |
102.73 |
|
|
|
| 22 |
B |
3081 |
3081 |
33.88 |
|
|
|
| 23 |
B |
3054 |
3054 |
26.64 |
|
|
|
| 24 |
B |
3041 |
3041 |
31.62 |
|
|
|
| 25 |
B |
1541 |
1541 |
14.81 |
|
|
|
| 26 |
B |
1534 |
1534 |
10.29 |
|
|
|
| 27 |
B |
1518 |
1518 |
0.40 |
|
|
|
| 28 |
B |
1444 |
1444 |
7.37 |
|
|
|
| 29 |
B |
1396 |
1396 |
1.36 |
|
|
|
| 30 |
B |
1313 |
1313 |
0.33 |
|
|
|
| 31 |
B |
1172 |
1172 |
2.33 |
|
|
|
| 32 |
B |
986 |
986 |
0.35 |
|
|
|
| 33 |
B |
980 |
980 |
3.88 |
|
|
|
| 34 |
B |
758 |
758 |
3.97 |
|
|
|
| 35 |
B |
441 |
441 |
0.31 |
|
|
|
| 36 |
B |
216 |
216 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29321.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 29321.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
