return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHCH2CH3 (1-Butene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-156.753556
Energy at 298.15K-156.761544
HF Energy-156.133832
Nuclear repulsion energy117.211327
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3277 3104 16.34      
2 A 3182 3014 4.37      
3 A 3175 3007 40.24      
4 A 3170 3003 32.51      
5 A 3166 2999 13.68      
6 A 3127 2961 7.89      
7 A 3079 2916 28.06      
8 A 3075 2912 25.43      
9 A 1710 1619 6.29      
10 A 1545 1463 8.79      
11 A 1540 1458 9.22      
12 A 1525 1445 3.83      
13 A 1477 1399 3.01      
14 A 1444 1368 5.52      
15 A 1368 1296 0.60      
16 A 1333 1263 0.76      
17 A 1318 1248 0.03      
18 A 1224 1159 0.15      
19 A 1116 1057 4.28      
20 A 1062 1005 1.50      
21 A 1021 967 20.78      
22 A 1004 951 3.58      
23 A 905 858 45.59      
24 A 882 836 1.62      
25 A 807 764 3.20      
26 A 655 621 11.84      
27 A 437 414 0.58      
28 A 325 308 0.71      
29 A 239 227 0.04      
30 A 107 102 0.07      

Unscaled Zero Point Vibrational Energy (zpe) 24147.8 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 22870.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.73618 0.13972 0.13692

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.705 -0.262 -0.300
C2 0.544 0.533 0.304
C3 -0.715 -0.279 0.369
C4 -1.843 0.009 -0.293
H5 -0.682 -1.175 0.991
H6 1.468 -0.568 -1.321
H7 2.621 0.333 -0.324
H8 1.906 -1.164 0.283
H9 0.818 0.861 1.314
H10 0.363 1.438 -0.285
H11 -1.918 0.890 -0.923
H12 -2.721 -0.624 -0.222

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.53102.51083.55752.86221.09221.09281.09312.15692.16573.85214.4406
C21.53101.49962.51542.21112.17002.17952.17651.09651.09482.77403.5033
C32.51081.49961.33871.09072.77643.46222.76832.13102.13002.11742.1190
C43.55752.51541.33872.09653.51434.47563.96993.22272.62791.08621.0845
H52.86222.21111.09072.09653.21453.86122.68272.54863.08913.07482.4354
H61.09222.17002.77643.51433.21451.77081.76663.06772.51413.70814.3307
H71.09282.17953.46224.47563.86121.77081.76672.49312.51474.61235.4277
H81.09312.17652.76833.96992.68271.76661.76672.51983.07834.50554.6855
H92.15691.09652.13103.22272.54863.06772.49312.51981.75983.53424.1333
H102.16571.09482.13002.62793.08912.51412.51473.07831.75982.43083.7097
H113.85212.77402.11741.08623.07483.70814.61234.50553.53422.43081.8515
H124.44063.50332.11901.08452.43544.33075.42774.68554.13333.70971.8515

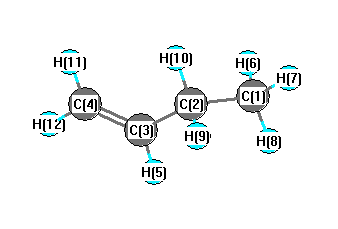
picture of 1-Butene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.876 C1 C2 H9 109.232
C1 C2 H10 110.016 C2 C1 H6 110.515
C2 C1 H7 111.229 C2 C1 H8 110.976
C2 C3 C4 124.707 C2 C3 H5 116.324
C3 C2 H9 109.357 C3 C2 H10 109.377
C3 C4 H11 121.315 C3 C4 H12 121.617
C4 C3 H5 118.956 H6 C1 H7 108.273
H6 C1 H8 107.877 H7 C1 H8 107.843
H9 C2 H10 106.850 H11 C4 H12 117.068
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability