Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -155.552074 |
| Energy at 298.15K | -155.557387 |
| HF Energy | -154.947629 |
| Nuclear repulsion energy | 103.507150 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3283 |
3109 |
0.00 |
|
|
|
| 2 |
Ag |
3188 |
3020 |
0.00 |
|
|
|
| 3 |
Ag |
3177 |
3009 |
0.00 |
|
|
|
| 4 |
Ag |
1718 |
1627 |
0.00 |
|
|
|
| 5 |
Ag |
1495 |
1416 |
0.00 |
|
|
|
| 6 |
Ag |
1325 |
1255 |
0.00 |
|
|
|
| 7 |
Ag |
1248 |
1182 |
0.00 |
|
|
|
| 8 |
Ag |
914 |
865 |
0.00 |
|
|
|
| 9 |
Ag |
521 |
493 |
0.00 |
|
|
|
| 10 |
Au |
1038 |
984 |
50.45 |
|
|
|
| 11 |
Au |
891 |
844 |
89.75 |
|
|
|
| 12 |
Au |
525 |
498 |
14.87 |
|
|
|
| 13 |
Au |
157 |
148 |
0.43 |
|
|
|
| 14 |
Bg |
980 |
928 |
0.00 |
|
|
|
| 15 |
Bg |
891 |
844 |
0.00 |
|
|
|
| 16 |
Bg |
764 |
724 |
0.00 |
|
|
|
| 17 |
Bu |
3283 |
3110 |
23.79 |
|
|
|
| 18 |
Bu |
3191 |
3022 |
11.82 |
|
|
|
| 19 |
Bu |
3181 |
3013 |
16.24 |
|
|
|
| 20 |
Bu |
1651 |
1564 |
11.26 |
|
|
|
| 21 |
Bu |
1429 |
1354 |
6.13 |
|
|
|
| 22 |
Bu |
1335 |
1265 |
2.11 |
|
|
|
| 23 |
Bu |
1016 |
962 |
3.10 |
|
|
|
| 24 |
Bu |
299 |
283 |
1.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18750.0 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 17758.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.729 |
0.000 |
| C2 |
0.000 |
-0.729 |
0.000 |
| C3 |
1.118 |
1.476 |
0.000 |
| C4 |
-1.118 |
-1.476 |
0.000 |
| H5 |
-0.972 |
1.221 |
0.000 |
| H6 |
0.972 |
-1.221 |
0.000 |
| H7 |
1.076 |
2.559 |
0.000 |
| H8 |
2.102 |
1.018 |
0.000 |
| H9 |
-1.076 |
-2.559 |
0.000 |
| H10 |
-2.102 |
-1.018 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4584 | 1.3443 | 2.4723 | 1.0896 | 2.1789 | 2.1230 | 2.1222 | 3.4600 | 2.7336 |
C2 | 1.4584 | | 2.4723 | 1.3443 | 2.1789 | 1.0896 | 3.4600 | 2.7336 | 2.1230 | 2.1222 | C3 | 1.3443 | 2.4723 | | 3.7029 | 2.1058 | 2.7005 | 1.0840 | 1.0860 | 4.5930 | 4.0730 | C4 | 2.4723 | 1.3443 | 3.7029 | | 2.7005 | 2.1058 | 4.5930 | 4.0730 | 1.0840 | 1.0860 | H5 | 1.0896 | 2.1789 | 2.1058 | 2.7005 | | 3.1214 | 2.4472 | 3.0816 | 3.7812 | 2.5076 | H6 | 2.1789 | 1.0896 | 2.7005 | 2.1058 | 3.1214 | | 3.7812 | 2.5076 | 2.4472 | 3.0816 | H7 | 2.1230 | 3.4600 | 1.0840 | 4.5930 | 2.4472 | 3.7812 | | 1.8517 | 5.5525 | 4.7853 | H8 | 2.1222 | 2.7336 | 1.0860 | 4.0730 | 3.0816 | 2.5076 | 1.8517 | | 4.7853 | 4.6718 | H9 | 3.4600 | 2.1230 | 4.5930 | 1.0840 | 3.7812 | 2.4472 | 5.5525 | 4.7853 | | 1.8517 | H10 | 2.7336 | 2.1222 | 4.0730 | 1.0860 | 2.5076 | 3.0816 | 4.7853 | 4.6718 | 1.8517 | |
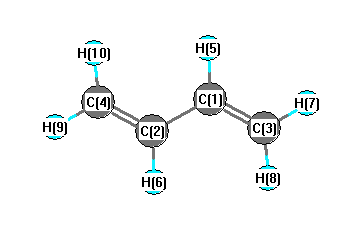 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
123.744 |
|
C1 |
C2 |
H6 |
116.811 |
| C1 |
C3 |
H7 |
121.545 |
|
C1 |
C3 |
H8 |
121.306 |
| C2 |
C1 |
C3 |
123.744 |
|
C2 |
C1 |
H5 |
116.811 |
| C2 |
C4 |
H9 |
121.545 |
|
C2 |
C4 |
H10 |
121.306 |
| C3 |
C1 |
H5 |
119.445 |
|
C4 |
C2 |
H6 |
119.445 |
| H7 |
C3 |
H8 |
117.148 |
|
H9 |
C4 |
H10 |
117.148 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -155.547990 |
| Energy at 298.15K | -155.553353 |
| HF Energy | -154.942478 |
| Nuclear repulsion energy | 104.761437 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3286 |
3112 |
4.47 |
|
|
|
| 2 |
A |
3197 |
3028 |
6.56 |
|
|
|
| 3 |
A |
3183 |
3015 |
14.61 |
|
|
|
| 4 |
A |
1687 |
1597 |
2.32 |
|
|
|
| 5 |
A |
1487 |
1408 |
10.93 |
|
|
|
| 6 |
A |
1350 |
1278 |
0.00 |
|
|
|
| 7 |
A |
1086 |
1028 |
0.04 |
|
|
|
| 8 |
A |
1001 |
948 |
5.33 |
|
|
|
| 9 |
A |
914 |
866 |
6.69 |
|
|
|
| 10 |
A |
905 |
857 |
4.20 |
|
|
|
| 11 |
A |
754 |
714 |
5.70 |
|
|
|
| 12 |
A |
277 |
262 |
0.01 |
|
|
|
| 13 |
A |
190 |
180 |
0.08 |
|
|
|
| 14 |
B |
3285 |
3111 |
18.96 |
|
|
|
| 15 |
B |
3188 |
3020 |
11.08 |
|
|
|
| 16 |
B |
3179 |
3011 |
4.59 |
|
|
|
| 17 |
B |
1688 |
1598 |
2.72 |
|
|
|
| 18 |
B |
1453 |
1376 |
2.25 |
|
|
|
| 19 |
B |
1317 |
1247 |
0.26 |
|
|
|
| 20 |
B |
1112 |
1053 |
4.60 |
|
|
|
| 21 |
B |
1024 |
970 |
42.78 |
|
|
|
| 22 |
B |
910 |
862 |
78.26 |
|
|
|
| 23 |
B |
632 |
599 |
9.14 |
|
|
|
| 24 |
B |
462 |
438 |
13.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18783.0 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 17789.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.125 |
0.724 |
0.555 |
| C2 |
-0.125 |
-0.724 |
0.555 |
| C3 |
-0.125 |
1.529 |
-0.491 |
| C4 |
0.125 |
-1.529 |
-0.491 |
| H5 |
0.518 |
1.159 |
1.473 |
| H6 |
-0.518 |
-1.159 |
1.473 |
| H7 |
0.096 |
2.590 |
-0.452 |
| H8 |
-0.571 |
1.145 |
-1.403 |
| H9 |
-0.096 |
-2.590 |
-0.452 |
| H10 |
0.571 |
-1.145 |
-1.403 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4703 | 1.3429 | 2.4845 | 1.0891 | 2.1915 | 2.1200 | 2.1195 | 3.4710 | 2.7432 |
C2 | 1.4703 | | 2.4845 | 1.3429 | 2.1915 | 1.0891 | 3.4710 | 2.7432 | 2.1200 | 2.1195 | C3 | 1.3429 | 2.4845 | | 3.0688 | 2.0992 | 3.3522 | 1.0843 | 1.0854 | 4.1196 | 2.9097 | C4 | 2.4845 | 1.3429 | 3.0688 | | 3.3522 | 2.0992 | 4.1196 | 2.9097 | 1.0843 | 1.0854 | H5 | 1.0891 | 2.1915 | 2.0992 | 3.3522 | | 2.5384 | 2.4352 | 3.0748 | 4.2586 | 3.6851 | H6 | 2.1915 | 1.0891 | 3.3522 | 2.0992 | 2.5384 | | 4.2586 | 3.6851 | 2.4352 | 3.0748 | H7 | 2.1200 | 3.4710 | 1.0843 | 4.1196 | 2.4352 | 4.2586 | | 1.8542 | 5.1837 | 3.8832 | H8 | 2.1195 | 2.7432 | 1.0854 | 2.9097 | 3.0748 | 3.6851 | 1.8542 | | 3.8832 | 2.5586 | H9 | 3.4710 | 2.1200 | 4.1196 | 1.0843 | 4.2586 | 2.4352 | 5.1837 | 3.8832 | | 1.8542 | H10 | 2.7432 | 2.1195 | 2.9097 | 1.0854 | 3.6851 | 3.0748 | 3.8832 | 2.5586 | 1.8542 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
123.990 |
|
C1 |
C2 |
H6 |
117.021 |
| C1 |
C3 |
H7 |
121.351 |
|
C1 |
C3 |
H8 |
121.214 |
| C2 |
C1 |
C3 |
123.990 |
|
C2 |
C1 |
H5 |
117.021 |
| C2 |
C4 |
H9 |
121.351 |
|
C2 |
C4 |
H10 |
121.214 |
| C3 |
C1 |
H5 |
118.977 |
|
C4 |
C2 |
H6 |
118.977 |
| H7 |
C3 |
H8 |
117.429 |
|
H9 |
C4 |
H10 |
117.429 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
