Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -191.463691 |
| Energy at 298.15K | |
| HF Energy | -190.802460 |
| Nuclear repulsion energy | 102.797699 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3286 |
3112 |
5.71 |
58.78 |
0.58 |
0.73 |
| 2 |
A' |
3227 |
3056 |
1.35 |
95.32 |
0.31 |
0.47 |
| 3 |
A' |
3183 |
3015 |
3.68 |
70.02 |
0.14 |
0.24 |
| 4 |
A' |
2950 |
2794 |
101.86 |
106.72 |
0.27 |
0.43 |
| 5 |
A' |
1767 |
1674 |
130.71 |
50.47 |
0.42 |
0.59 |
| 6 |
A' |
1679 |
1590 |
5.37 |
5.59 |
0.15 |
0.26 |
| 7 |
A' |
1478 |
1399 |
11.42 |
4.16 |
0.64 |
0.78 |
| 8 |
A' |
1410 |
1335 |
4.06 |
16.29 |
0.42 |
0.60 |
| 9 |
A' |
1311 |
1241 |
4.12 |
6.43 |
0.24 |
0.39 |
| 10 |
A' |
1190 |
1127 |
29.22 |
10.43 |
0.55 |
0.71 |
| 11 |
A' |
938 |
889 |
21.79 |
0.91 |
0.06 |
0.11 |
| 12 |
A' |
576 |
545 |
4.97 |
4.95 |
0.34 |
0.51 |
| 13 |
A' |
322 |
305 |
9.29 |
0.38 |
0.33 |
0.50 |
| 14 |
A" |
1035 |
980 |
9.14 |
2.35 |
0.75 |
0.86 |
| 15 |
A" |
1015 |
961 |
24.25 |
2.23 |
0.75 |
0.86 |
| 16 |
A" |
959 |
908 |
38.86 |
0.64 |
0.75 |
0.86 |
| 17 |
A" |
604 |
572 |
12.41 |
2.72 |
0.75 |
0.86 |
| 18 |
A" |
165 |
156 |
3.72 |
1.00 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13546.7 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 12830.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.150 |
-0.747 |
0.000 |
| C2 |
0.000 |
0.721 |
0.000 |
| C3 |
1.218 |
1.285 |
0.000 |
| O4 |
-1.221 |
-1.323 |
0.000 |
| H5 |
0.805 |
-1.315 |
0.000 |
| H6 |
-0.914 |
1.309 |
0.000 |
| H7 |
1.355 |
2.360 |
0.000 |
| H8 |
2.116 |
0.673 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4759 | 2.4491 | 1.2162 | 1.1111 | 2.1935 | 3.4521 | 2.6737 |
C2 | 1.4759 | | 1.3417 | 2.3810 | 2.1899 | 1.0866 | 2.1262 | 2.1163 | C3 | 2.4491 | 1.3417 | | 3.5702 | 2.6327 | 2.1317 | 1.0839 | 1.0867 | O4 | 1.2162 | 2.3810 | 3.5702 | | 2.0258 | 2.6498 | 4.4942 | 3.8880 | H5 | 1.1111 | 2.1899 | 2.6327 | 2.0258 | | 3.1372 | 3.7163 | 2.3815 | H6 | 2.1935 | 1.0866 | 2.1317 | 2.6498 | 3.1372 | | 2.5003 | 3.0957 | H7 | 3.4521 | 2.1262 | 1.0839 | 4.4942 | 3.7163 | 2.5003 | | 1.8507 | H8 | 2.6737 | 2.1163 | 1.0867 | 3.8880 | 2.3815 | 3.0957 | 1.8507 | |
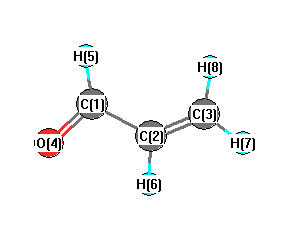 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.657 |
|
C1 |
C2 |
H6 |
116.926 |
| C2 |
C1 |
O4 |
124.087 |
|
C2 |
C1 |
H5 |
114.946 |
| C2 |
C3 |
H7 |
122.100 |
|
C2 |
C3 |
H8 |
120.901 |
| C3 |
C2 |
H6 |
122.418 |
|
O4 |
C1 |
H5 |
120.967 |
| H7 |
C3 |
H8 |
116.998 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -191.461336 |
| Energy at 298.15K | |
| HF Energy | -190.799891 |
| Nuclear repulsion energy | 104.414188 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3299 |
3124 |
2.48 |
50.29 |
0.65 |
0.79 |
| 2 |
A' |
3219 |
3049 |
10.06 |
115.54 |
0.26 |
0.41 |
| 3 |
A' |
3192 |
3023 |
3.09 |
52.51 |
0.17 |
0.29 |
| 4 |
A' |
2983 |
2826 |
142.98 |
168.03 |
0.30 |
0.47 |
| 5 |
A' |
1766 |
1673 |
64.12 |
10.77 |
0.50 |
0.67 |
| 6 |
A' |
1682 |
1593 |
41.70 |
29.90 |
0.22 |
0.36 |
| 7 |
A' |
1457 |
1380 |
39.25 |
6.53 |
0.57 |
0.73 |
| 8 |
A' |
1447 |
1370 |
1.39 |
10.09 |
0.35 |
0.52 |
| 9 |
A' |
1325 |
1255 |
1.66 |
11.99 |
0.37 |
0.54 |
| 10 |
A' |
1084 |
1026 |
4.54 |
3.71 |
0.73 |
0.84 |
| 11 |
A' |
950 |
900 |
55.94 |
2.79 |
0.18 |
0.30 |
| 12 |
A' |
690 |
653 |
9.65 |
1.24 |
0.56 |
0.72 |
| 13 |
A' |
295 |
280 |
6.84 |
2.70 |
0.48 |
0.65 |
| 14 |
A" |
1028 |
973 |
2.33 |
5.08 |
0.75 |
0.86 |
| 15 |
A" |
1023 |
969 |
38.57 |
1.88 |
0.75 |
0.86 |
| 16 |
A" |
964 |
913 |
22.68 |
0.98 |
0.75 |
0.86 |
| 17 |
A" |
563 |
533 |
12.78 |
4.58 |
0.75 |
0.86 |
| 18 |
A" |
168 |
159 |
6.88 |
0.49 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13567.4 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 12849.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.893 |
-0.286 |
0.000 |
| C2 |
0.000 |
0.903 |
0.000 |
| C3 |
1.334 |
0.765 |
0.000 |
| O4 |
-0.497 |
-1.437 |
0.000 |
| H5 |
-1.979 |
-0.071 |
0.000 |
| H6 |
-0.468 |
1.884 |
0.000 |
| H7 |
1.998 |
1.621 |
0.000 |
| H8 |
1.775 |
-0.227 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4868 | 2.4622 | 1.2169 | 1.1078 | 2.2110 | 3.4635 | 2.6683 |
C2 | 1.4868 | | 1.3411 | 2.3918 | 2.2059 | 1.0871 | 2.1236 | 2.1038 | C3 | 2.4622 | 1.3411 | | 2.8634 | 3.4171 | 2.1214 | 1.0838 | 1.0851 | O4 | 1.2169 | 2.3918 | 2.8634 | | 2.0159 | 3.3207 | 3.9468 | 2.5739 | H5 | 1.1078 | 2.2059 | 3.4171 | 2.0159 | | 2.4706 | 4.3227 | 3.7575 | H6 | 2.2110 | 1.0871 | 2.1214 | 3.3207 | 2.4706 | | 2.4807 | 3.0799 | H7 | 3.4635 | 2.1236 | 1.0838 | 3.9468 | 4.3227 | 2.4807 | | 1.8612 | H8 | 2.6683 | 2.1038 | 1.0851 | 2.5739 | 3.7575 | 3.0799 | 1.8612 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.998 |
|
C1 |
C2 |
H6 |
117.577 |
| C2 |
C1 |
O4 |
124.112 |
|
C2 |
C1 |
H5 |
115.699 |
| C2 |
C3 |
H7 |
121.907 |
|
C2 |
C3 |
H8 |
119.880 |
| C3 |
C2 |
H6 |
121.425 |
|
O4 |
C1 |
H5 |
120.189 |
| H7 |
C3 |
H8 |
118.213 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
