Jump to
S1C2
Energy calculated at PBE1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -191.728588 |
| Energy at 298.15K | |
| HF Energy | -191.728588 |
| Nuclear repulsion energy | 103.320172 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3251 |
3116 |
7.94 |
64.05 |
0.54 |
0.71 |
| 2 |
A' |
3201 |
3068 |
2.65 |
103.34 |
0.34 |
0.51 |
| 3 |
A' |
3155 |
3024 |
7.88 |
70.73 |
0.14 |
0.24 |
| 4 |
A' |
2905 |
2784 |
111.34 |
105.70 |
0.29 |
0.45 |
| 5 |
A' |
1829 |
1753 |
245.89 |
61.37 |
0.43 |
0.60 |
| 6 |
A' |
1706 |
1635 |
2.45 |
32.07 |
0.19 |
0.32 |
| 7 |
A' |
1464 |
1403 |
13.17 |
5.90 |
0.60 |
0.75 |
| 8 |
A' |
1399 |
1341 |
8.27 |
19.49 |
0.44 |
0.61 |
| 9 |
A' |
1303 |
1249 |
4.11 |
7.16 |
0.25 |
0.40 |
| 10 |
A' |
1181 |
1132 |
35.27 |
11.30 |
0.53 |
0.69 |
| 11 |
A' |
929 |
891 |
22.21 |
1.17 |
0.06 |
0.11 |
| 12 |
A' |
577 |
553 |
6.39 |
5.46 |
0.34 |
0.50 |
| 13 |
A' |
319 |
306 |
9.63 |
1.01 |
0.37 |
0.54 |
| 14 |
A" |
1042 |
999 |
10.27 |
1.67 |
0.75 |
0.86 |
| 15 |
A" |
1027 |
984 |
10.11 |
3.70 |
0.75 |
0.86 |
| 16 |
A" |
993 |
952 |
49.32 |
1.10 |
0.75 |
0.86 |
| 17 |
A" |
611 |
585 |
11.84 |
2.41 |
0.75 |
0.86 |
| 18 |
A" |
172 |
165 |
3.78 |
1.38 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13531.4 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 12969.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.152 |
-0.744 |
0.000 |
| C2 |
0.000 |
0.718 |
0.000 |
| C3 |
1.207 |
1.282 |
0.000 |
| O4 |
-1.210 |
-1.320 |
0.000 |
| H5 |
0.807 |
-1.310 |
0.000 |
| H6 |
-0.916 |
1.303 |
0.000 |
| H7 |
1.345 |
2.358 |
0.000 |
| H8 |
2.109 |
0.673 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4701 | 2.4392 | 1.2046 | 1.1135 | 2.1850 | 3.4441 | 2.6680 |
C2 | 1.4701 | | 1.3322 | 2.3698 | 2.1833 | 1.0868 | 2.1208 | 2.1094 | C3 | 2.4392 | 1.3322 | | 3.5507 | 2.6227 | 2.1234 | 1.0849 | 1.0879 | O4 | 1.2046 | 2.3698 | 3.5507 | | 2.0169 | 2.6387 | 4.4778 | 3.8709 | H5 | 1.1135 | 2.1833 | 2.6227 | 2.0169 | | 3.1303 | 3.7074 | 2.3725 | H6 | 2.1850 | 1.0868 | 2.1234 | 2.6387 | 3.1303 | | 2.4955 | 3.0899 | H7 | 3.4441 | 2.1208 | 1.0849 | 4.4778 | 3.7074 | 2.4955 | | 1.8496 | H8 | 2.6680 | 2.1094 | 1.0879 | 3.8709 | 2.3725 | 3.0899 | 1.8496 | |
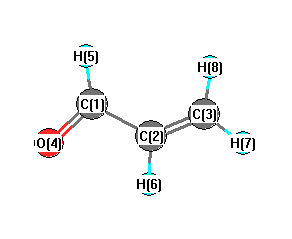 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.938 |
|
C1 |
C2 |
H6 |
116.625 |
| C2 |
C1 |
O4 |
124.456 |
|
C2 |
C1 |
H5 |
114.654 |
| C2 |
C3 |
H7 |
122.340 |
|
C2 |
C3 |
H8 |
120.966 |
| C3 |
C2 |
H6 |
122.437 |
|
O4 |
C1 |
H5 |
120.890 |
| H7 |
C3 |
H8 |
116.694 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.106 |
|
|
|
| 2 |
C |
-0.259 |
|
|
|
| 3 |
C |
-0.419 |
|
|
|
| 4 |
O |
-0.298 |
|
|
|
| 5 |
H |
0.162 |
|
|
|
| 6 |
H |
0.229 |
|
|
|
| 7 |
H |
0.244 |
|
|
|
| 8 |
H |
0.235 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.416 |
2.055 |
0.000 |
3.172 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.141 |
-2.572 |
0.000 |
| y |
-2.572 |
-23.728 |
0.000 |
| z |
0.000 |
0.000 |
-24.946 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.195 |
-2.572 |
0.000 |
| y |
-2.572 |
0.816 |
0.000 |
| z |
0.000 |
0.000 |
-1.012 |
|
| Polar |
|---|
| 3z2-r2 | -2.023 |
|---|
| x2-y2 | -0.414 |
|---|
| xy | -2.572 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.926 |
1.685 |
0.000 |
| y |
1.685 |
6.287 |
0.000 |
| z |
0.000 |
0.000 |
2.544 |
<r2> (average value of r
2) Å
2
| <r2> |
83.126 |
| (<r2>)1/2 |
9.117 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -191.725990 |
| Energy at 298.15K | |
| HF Energy | -191.725990 |
| Nuclear repulsion energy | 104.920052 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3264 |
3129 |
3.66 |
54.03 |
0.65 |
0.79 |
| 2 |
A' |
3190 |
3058 |
12.59 |
124.94 |
0.26 |
0.41 |
| 3 |
A' |
3166 |
3034 |
7.72 |
51.60 |
0.17 |
0.29 |
| 4 |
A' |
2933 |
2811 |
159.81 |
170.52 |
0.32 |
0.48 |
| 5 |
A' |
1830 |
1754 |
123.71 |
14.70 |
0.60 |
0.75 |
| 6 |
A' |
1697 |
1627 |
68.53 |
47.06 |
0.20 |
0.33 |
| 7 |
A' |
1444 |
1384 |
34.86 |
8.38 |
0.73 |
0.84 |
| 8 |
A' |
1434 |
1374 |
14.11 |
7.64 |
0.25 |
0.40 |
| 9 |
A' |
1321 |
1266 |
2.66 |
15.89 |
0.37 |
0.54 |
| 10 |
A' |
1076 |
1031 |
5.73 |
4.32 |
0.68 |
0.81 |
| 11 |
A' |
946 |
907 |
61.96 |
4.05 |
0.19 |
0.32 |
| 12 |
A' |
690 |
661 |
11.14 |
0.81 |
0.71 |
0.83 |
| 13 |
A' |
288 |
276 |
7.07 |
3.48 |
0.46 |
0.63 |
| 14 |
A" |
1042 |
999 |
5.49 |
8.43 |
0.75 |
0.86 |
| 15 |
A" |
1031 |
988 |
33.24 |
0.19 |
0.75 |
0.86 |
| 16 |
A" |
1007 |
965 |
21.33 |
0.80 |
0.75 |
0.86 |
| 17 |
A" |
562 |
538 |
11.88 |
4.08 |
0.75 |
0.86 |
| 18 |
A" |
182 |
174 |
7.61 |
0.46 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13550.3 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 12988.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.884 |
-0.291 |
0.000 |
| C2 |
0.000 |
0.896 |
0.000 |
| C3 |
1.325 |
0.769 |
0.000 |
| O4 |
-0.495 |
-1.432 |
0.000 |
| H5 |
-1.973 |
-0.072 |
0.000 |
| H6 |
-0.477 |
1.873 |
0.000 |
| H7 |
1.989 |
1.627 |
0.000 |
| H8 |
1.774 |
-0.220 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4806 | 2.4508 | 1.2053 | 1.1108 | 2.2024 | 3.4546 | 2.6595 |
C2 | 1.4806 | | 1.3315 | 2.3801 | 2.1984 | 1.0875 | 2.1188 | 2.0964 | C3 | 2.4508 | 1.3315 | | 2.8556 | 3.4044 | 2.1145 | 1.0848 | 1.0861 | O4 | 1.2053 | 2.3801 | 2.8556 | | 2.0085 | 3.3051 | 3.9400 | 2.5720 | H5 | 1.1108 | 2.1984 | 3.4044 | 2.0085 | | 2.4544 | 4.3114 | 3.7506 | H6 | 2.2024 | 1.0875 | 2.1145 | 3.3051 | 2.4544 | | 2.4786 | 3.0748 | H7 | 3.4546 | 2.1188 | 1.0848 | 3.9400 | 4.3114 | 2.4786 | | 1.8598 | H8 | 2.6595 | 2.0964 | 1.0861 | 2.5720 | 3.7506 | 3.0748 | 1.8598 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.174 |
|
C1 |
C2 |
H6 |
117.275 |
| C2 |
C1 |
O4 |
124.460 |
|
C2 |
C1 |
H5 |
115.327 |
| C2 |
C3 |
H7 |
122.196 |
|
C2 |
C3 |
H8 |
119.910 |
| C3 |
C2 |
H6 |
121.551 |
|
O4 |
C1 |
H5 |
120.213 |
| H7 |
C3 |
H8 |
117.894 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.101 |
|
|
|
| 2 |
C |
-0.306 |
|
|
|
| 3 |
C |
-0.370 |
|
|
|
| 4 |
O |
-0.294 |
|
|
|
| 5 |
H |
0.170 |
|
|
|
| 6 |
H |
0.215 |
|
|
|
| 7 |
H |
0.236 |
|
|
|
| 8 |
H |
0.249 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.249 |
2.646 |
0.000 |
2.658 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.010 |
-0.618 |
0.000 |
| y |
-0.618 |
-25.856 |
0.000 |
| z |
0.000 |
0.000 |
-24.919 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.377 |
-0.618 |
0.000 |
| y |
-0.618 |
-3.391 |
0.000 |
| z |
0.000 |
0.000 |
-1.986 |
|
| Polar |
|---|
| 3z2-r2 | -3.972 |
|---|
| x2-y2 | 5.846 |
|---|
| xy | -0.618 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.112 |
0.455 |
0.000 |
| y |
0.455 |
5.638 |
0.000 |
| z |
0.000 |
0.000 |
2.532 |
<r2> (average value of r
2) Å
2
| <r2> |
74.637 |
| (<r2>)1/2 |
8.639 |
