Jump to
S1C2
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -191.785680 |
| Energy at 298.15K | |
| HF Energy | -191.567865 |
| Nuclear repulsion energy | 102.999942 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3257 |
3257 |
9.05 |
65.63 |
0.56 |
0.72 |
| 2 |
A' |
3204 |
3204 |
2.74 |
101.61 |
0.34 |
0.50 |
| 3 |
A' |
3162 |
3162 |
7.54 |
73.09 |
0.14 |
0.24 |
| 4 |
A' |
2915 |
2915 |
113.03 |
109.20 |
0.29 |
0.45 |
| 5 |
A' |
1772 |
1772 |
200.82 |
60.38 |
0.42 |
0.59 |
| 6 |
A' |
1684 |
1684 |
3.13 |
17.44 |
0.15 |
0.27 |
| 7 |
A' |
1475 |
1475 |
10.72 |
6.27 |
0.57 |
0.73 |
| 8 |
A' |
1407 |
1407 |
6.16 |
19.63 |
0.42 |
0.59 |
| 9 |
A' |
1311 |
1311 |
3.30 |
8.43 |
0.24 |
0.39 |
| 10 |
A' |
1183 |
1183 |
34.91 |
12.01 |
0.53 |
0.70 |
| 11 |
A' |
932 |
932 |
23.69 |
1.01 |
0.06 |
0.11 |
| 12 |
A' |
574 |
574 |
5.46 |
5.66 |
0.34 |
0.50 |
| 13 |
A' |
322 |
322 |
9.75 |
0.80 |
0.35 |
0.52 |
| 14 |
A" |
1039 |
1039 |
11.43 |
1.65 |
0.75 |
0.86 |
| 15 |
A" |
1022 |
1022 |
15.00 |
3.52 |
0.75 |
0.86 |
| 16 |
A" |
983 |
983 |
41.33 |
1.07 |
0.75 |
0.86 |
| 17 |
A" |
610 |
610 |
11.52 |
2.65 |
0.75 |
0.86 |
| 18 |
A" |
170 |
170 |
3.87 |
1.29 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13511.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13511.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.148 |
-0.748 |
0.000 |
| C2 |
0.000 |
0.719 |
0.000 |
| C3 |
1.209 |
1.290 |
0.000 |
| O4 |
-1.213 |
-1.326 |
0.000 |
| H5 |
0.808 |
-1.313 |
0.000 |
| H6 |
-0.917 |
1.300 |
0.000 |
| H7 |
1.340 |
2.365 |
0.000 |
| H8 |
2.113 |
0.688 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4745 | 2.4484 | 1.2117 | 1.1108 | 2.1872 | 3.4502 | 2.6782 |
C2 | 1.4745 | | 1.3369 | 2.3779 | 2.1867 | 1.0851 | 2.1223 | 2.1127 | C3 | 2.4484 | 1.3369 | | 3.5650 | 2.6332 | 2.1256 | 1.0828 | 1.0857 | O4 | 1.2117 | 2.3779 | 3.5650 | | 2.0214 | 2.6426 | 4.4879 | 3.8880 | H5 | 1.1108 | 2.1867 | 2.6332 | 2.0214 | | 3.1305 | 3.7157 | 2.3883 | H6 | 2.1872 | 1.0851 | 2.1256 | 2.6426 | 3.1305 | | 2.4954 | 3.0903 | H7 | 3.4502 | 2.1223 | 1.0828 | 4.4879 | 3.7157 | 2.4954 | | 1.8460 | H8 | 2.6782 | 2.1127 | 1.0857 | 3.8880 | 2.3883 | 3.0903 | 1.8460 | |
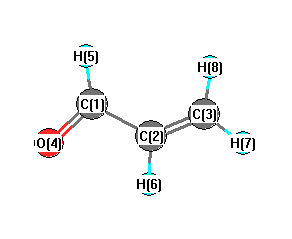 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.041 |
|
C1 |
C2 |
H6 |
116.586 |
| C2 |
C1 |
O4 |
124.266 |
|
C2 |
C1 |
H5 |
114.792 |
| C2 |
C3 |
H7 |
122.233 |
|
C2 |
C3 |
H8 |
121.060 |
| C3 |
C2 |
H6 |
122.373 |
|
O4 |
C1 |
H5 |
120.942 |
| H7 |
C3 |
H8 |
116.707 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -191.783260 |
| Energy at 298.15K | |
| HF Energy | -191.565280 |
| Nuclear repulsion energy | 104.572142 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3270 |
3270 |
4.39 |
54.71 |
0.65 |
0.79 |
| 2 |
A' |
3193 |
3193 |
13.45 |
130.22 |
0.25 |
0.40 |
| 3 |
A' |
3172 |
3172 |
7.30 |
49.21 |
0.19 |
0.32 |
| 4 |
A' |
2945 |
2945 |
160.85 |
176.18 |
0.32 |
0.48 |
| 5 |
A' |
1774 |
1774 |
88.19 |
10.70 |
0.64 |
0.78 |
| 6 |
A' |
1673 |
1673 |
68.50 |
42.75 |
0.21 |
0.35 |
| 7 |
A' |
1454 |
1454 |
44.74 |
8.08 |
0.57 |
0.72 |
| 8 |
A' |
1443 |
1443 |
1.01 |
9.70 |
0.33 |
0.50 |
| 9 |
A' |
1328 |
1328 |
2.75 |
15.83 |
0.36 |
0.53 |
| 10 |
A' |
1084 |
1084 |
5.22 |
4.47 |
0.68 |
0.81 |
| 11 |
A' |
939 |
939 |
61.04 |
3.76 |
0.18 |
0.31 |
| 12 |
A' |
690 |
690 |
10.36 |
0.95 |
0.68 |
0.81 |
| 13 |
A' |
292 |
292 |
6.99 |
3.34 |
0.46 |
0.63 |
| 14 |
A" |
1034 |
1034 |
6.55 |
7.94 |
0.75 |
0.86 |
| 15 |
A" |
1029 |
1029 |
33.72 |
0.13 |
0.75 |
0.86 |
| 16 |
A" |
994 |
994 |
18.22 |
1.13 |
0.75 |
0.86 |
| 17 |
A" |
564 |
564 |
11.73 |
4.45 |
0.75 |
0.86 |
| 18 |
A" |
183 |
183 |
7.35 |
0.47 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13530.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13530.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.888 |
-0.291 |
0.000 |
| C2 |
0.000 |
0.899 |
0.000 |
| C3 |
1.331 |
0.772 |
0.000 |
| O4 |
-0.497 |
-1.438 |
0.000 |
| H5 |
-1.975 |
-0.073 |
0.000 |
| H6 |
-0.476 |
1.875 |
0.000 |
| H7 |
1.990 |
1.631 |
0.000 |
| H8 |
1.781 |
-0.213 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4850 | 2.4605 | 1.2129 | 1.1078 | 2.2045 | 3.4610 | 2.6706 |
C2 | 1.4850 | | 1.3366 | 2.3898 | 2.2009 | 1.0858 | 2.1202 | 2.1000 | C3 | 2.4605 | 1.3366 | | 2.8681 | 3.4115 | 2.1168 | 1.0828 | 1.0839 | O4 | 1.2129 | 2.3898 | 2.8681 | | 2.0124 | 3.3135 | 3.9504 | 2.5860 | H5 | 1.1078 | 2.2009 | 3.4115 | 2.0124 | | 2.4573 | 4.3151 | 3.7582 | H6 | 2.2045 | 1.0858 | 2.1168 | 3.3135 | 2.4573 | | 2.4782 | 3.0753 | H7 | 3.4610 | 2.1202 | 1.0828 | 3.9504 | 4.3151 | 2.4782 | | 1.8567 | H8 | 2.6706 | 2.1000 | 1.0839 | 2.5860 | 3.7582 | 3.0753 | 1.8567 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.301 |
|
C1 |
C2 |
H6 |
117.228 |
| C2 |
C1 |
O4 |
124.399 |
|
C2 |
C1 |
H5 |
115.403 |
| C2 |
C3 |
H7 |
122.055 |
|
C2 |
C3 |
H8 |
120.002 |
| C3 |
C2 |
H6 |
121.471 |
|
O4 |
C1 |
H5 |
120.198 |
| H7 |
C3 |
H8 |
117.943 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
