Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -3110.979175 |
| Energy at 298.15K | |
| HF Energy | -3109.958538 |
| Nuclear repulsion energy | 289.746372 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3154 |
2987 |
11.29 |
7.66 |
0.08 |
0.16 |
| 2 |
A' |
3146 |
2979 |
1.21 |
143.67 |
0.09 |
0.16 |
| 3 |
A' |
1522 |
1441 |
0.53 |
15.34 |
0.71 |
0.83 |
| 4 |
A' |
1521 |
1441 |
7.74 |
0.78 |
0.71 |
0.83 |
| 5 |
A' |
1385 |
1312 |
2.81 |
10.83 |
0.66 |
0.79 |
| 6 |
A' |
1286 |
1218 |
43.46 |
2.24 |
0.56 |
0.71 |
| 7 |
A' |
1103 |
1044 |
1.09 |
8.64 |
0.74 |
0.85 |
| 8 |
A' |
796 |
754 |
24.51 |
41.16 |
0.29 |
0.45 |
| 9 |
A' |
686 |
650 |
42.72 |
12.14 |
0.25 |
0.40 |
| 10 |
A' |
259 |
245 |
0.72 |
3.93 |
0.38 |
0.55 |
| 11 |
A' |
203 |
192 |
6.96 |
0.13 |
0.30 |
0.46 |
| 12 |
A" |
3235 |
3064 |
2.11 |
1.92 |
0.75 |
0.86 |
| 13 |
A" |
3214 |
3044 |
0.34 |
93.67 |
0.75 |
0.86 |
| 14 |
A" |
1339 |
1268 |
0.22 |
6.34 |
0.75 |
0.86 |
| 15 |
A" |
1188 |
1125 |
5.15 |
0.05 |
0.75 |
0.86 |
| 16 |
A" |
1023 |
969 |
0.23 |
6.27 |
0.75 |
0.86 |
| 17 |
A" |
789 |
748 |
4.84 |
0.03 |
0.75 |
0.86 |
| 18 |
A" |
125 |
118 |
5.49 |
0.02 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12986.4 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 12299.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.639 |
0.000 |
| C2 |
1.233 |
-0.234 |
0.000 |
| Br3 |
-1.585 |
-0.486 |
0.000 |
| Cl4 |
2.684 |
0.809 |
0.000 |
| H5 |
-0.046 |
1.264 |
0.890 |
| H6 |
-0.046 |
1.264 |
-0.890 |
| H7 |
1.274 |
-0.860 |
0.890 |
| H8 |
1.274 |
-0.860 |
-0.890 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5110 | 1.9439 | 2.6894 | 1.0883 | 1.0883 | 2.1588 | 2.1588 |
C2 | 1.5110 | | 2.8296 | 1.7872 | 2.1611 | 2.1611 | 1.0885 | 1.0885 | Br3 | 1.9439 | 2.8296 | | 4.4614 | 2.4945 | 2.4945 | 3.0173 | 3.0173 | Cl4 | 2.6894 | 1.7872 | 4.4614 | | 2.9069 | 2.9069 | 2.3594 | 2.3594 | H5 | 1.0883 | 2.1611 | 2.4945 | 2.9069 | | 1.7806 | 2.4995 | 3.0686 | H6 | 1.0883 | 2.1611 | 2.4945 | 2.9069 | 1.7806 | | 3.0686 | 2.4995 | H7 | 2.1588 | 1.0885 | 3.0173 | 2.3594 | 2.4995 | 3.0686 | | 1.7797 | H8 | 2.1588 | 1.0885 | 3.0173 | 2.3594 | 3.0686 | 2.4995 | 1.7797 | |
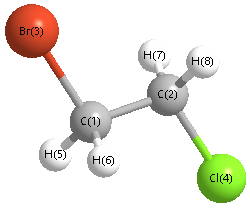 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
108.973 |
|
C1 |
C2 |
H7 |
111.253 |
| C1 |
C2 |
H8 |
111.253 |
|
C2 |
C1 |
Br3 |
109.330 |
| C2 |
C1 |
Cl4 |
38.933 |
|
C2 |
C1 |
H6 |
111.454 |
| Br3 |
C1 |
H5 |
107.320 |
|
Br3 |
C1 |
H6 |
107.320 |
| Cl4 |
C2 |
H7 |
107.781 |
|
Cl4 |
C2 |
H8 |
107.781 |
| H5 |
C1 |
H6 |
109.787 |
|
H7 |
C2 |
H8 |
109.672 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -3110.976292 |
| Energy at 298.15K | |
| HF Energy | -3109.954909 |
| Nuclear repulsion energy | 306.108985 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3211 |
3041 |
2.28 |
35.15 |
0.75 |
0.86 |
| 2 |
A |
3196 |
3027 |
1.38 |
91.13 |
0.75 |
0.86 |
| 3 |
A |
3134 |
2968 |
14.55 |
152.65 |
0.04 |
0.08 |
| 4 |
A |
3123 |
2958 |
9.46 |
82.65 |
0.25 |
0.40 |
| 5 |
A |
1507 |
1428 |
2.02 |
6.04 |
0.68 |
0.81 |
| 6 |
A |
1501 |
1422 |
12.63 |
12.64 |
0.73 |
0.85 |
| 7 |
A |
1394 |
1320 |
24.64 |
2.96 |
0.72 |
0.84 |
| 8 |
A |
1353 |
1281 |
39.35 |
1.46 |
0.64 |
0.78 |
| 9 |
A |
1260 |
1193 |
4.15 |
13.40 |
0.74 |
0.85 |
| 10 |
A |
1193 |
1130 |
3.50 |
5.58 |
0.74 |
0.85 |
| 11 |
A |
1084 |
1027 |
0.97 |
2.75 |
0.75 |
0.85 |
| 12 |
A |
975 |
923 |
9.47 |
6.39 |
0.43 |
0.60 |
| 13 |
A |
908 |
860 |
22.45 |
2.15 |
0.74 |
0.85 |
| 14 |
A |
720 |
682 |
15.38 |
9.21 |
0.42 |
0.59 |
| 15 |
A |
609 |
577 |
8.89 |
14.00 |
0.20 |
0.34 |
| 16 |
A |
401 |
380 |
5.83 |
2.28 |
0.71 |
0.83 |
| 17 |
A |
255 |
242 |
1.05 |
1.32 |
0.57 |
0.72 |
| 18 |
A |
109 |
103 |
0.47 |
1.74 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 12967.3 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 12281.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.008 |
1.099 |
-0.384 |
| C2 |
1.281 |
0.896 |
0.404 |
| Br3 |
-1.347 |
-0.224 |
0.036 |
| Cl4 |
2.153 |
-0.576 |
-0.087 |
| H5 |
-0.421 |
2.071 |
-0.138 |
| H6 |
0.192 |
1.039 |
-1.455 |
| H7 |
1.948 |
1.741 |
0.219 |
| H8 |
1.077 |
0.822 |
1.471 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5107 | 1.9390 | 2.7375 | 1.0908 | 1.0889 | 2.1304 | 2.1583 |
C2 | 1.5107 | | 2.8799 | 1.7796 | 2.1383 | 2.1591 | 1.0917 | 1.0891 | Br3 | 1.9390 | 2.8799 | | 3.5191 | 2.4809 | 2.4874 | 3.8399 | 3.0042 | Cl4 | 2.7375 | 1.7796 | 3.5191 | | 3.6926 | 2.8857 | 2.3461 | 2.3533 | H5 | 1.0908 | 2.1383 | 2.4809 | 3.6926 | | 1.7816 | 2.4181 | 2.5286 | H6 | 1.0889 | 2.1591 | 2.4874 | 2.8857 | 1.7816 | | 2.5255 | 3.0645 | H7 | 2.1304 | 1.0917 | 3.8399 | 2.3461 | 2.4181 | 2.5255 | | 1.7800 | H8 | 2.1583 | 1.0891 | 3.0042 | 2.3533 | 2.5286 | 3.0645 | 1.7800 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.352 |
|
C1 |
C2 |
H7 |
108.830 |
| C1 |
C2 |
H8 |
111.200 |
|
C2 |
C1 |
Br3 |
112.609 |
| C2 |
C1 |
Cl4 |
36.959 |
|
C2 |
C1 |
H6 |
111.277 |
| Br3 |
C1 |
H5 |
106.553 |
|
Br3 |
C1 |
H6 |
107.099 |
| Cl4 |
C2 |
H7 |
107.143 |
|
Cl4 |
C2 |
H8 |
107.795 |
| H5 |
C1 |
H6 |
109.647 |
|
H7 |
C2 |
H8 |
109.410 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
