return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H5Cl (1-Propene, 3-chloro-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-576.664444
Energy at 298.15K-576.669493
HF Energy-576.018176
Nuclear repulsion energy143.638254
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3287 3113 9.71      
2 A 3213 3043 6.69      
3 A 3196 3026 3.80      
4 A 3185 3017 3.57      
5 A 3128 2962 17.37      
6 A 1709 1619 1.13      
7 A 1517 1437 6.11      
8 A 1475 1397 11.15      
9 A 1360 1288 26.37      
10 A 1329 1258 7.54      
11 A 1267 1200 1.45      
12 A 1157 1096 2.16      
13 A 1020 966 23.55      
14 A 976 925 6.70      
15 A 931 882 3.38      
16 A 927 878 50.12      
17 A 792 750 49.18      
18 A 618 585 5.61      
19 A 412 391 0.54      
20 A 293 277 4.50      
21 A 105 100 0.43      

Unscaled Zero Point Vibrational Energy (zpe) 15947.4 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15103.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.71233 0.09387 0.09104

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.213 -0.201 -0.309
C2 1.120 -0.118 0.459
C3 -0.047 0.746 0.124
Cl4 -1.527 -0.244 -0.111
H5 2.297 0.353 -1.238
H6 3.052 -0.826 -0.028
H7 1.048 -0.690 1.380
H8 -0.274 1.443 0.931
H9 0.123 1.302 -0.798

Atom - Atom Distances (Å)
  C1 C2 C3 Cl4 H5 H6 H7 H8 H9
C11.33772.48783.74491.08571.08392.10913.22852.6196
C21.33771.48992.71032.11812.11491.08762.14542.1417
C32.48781.48991.79632.73893.47852.20051.09041.0897
Cl43.74492.71031.79634.03084.61663.00882.34582.3634
H51.08572.11812.73894.03081.85153.08323.53592.4116
H61.08392.11493.47854.61661.85152.45314.13943.7013
H72.10911.08762.20053.00883.08322.45312.55013.0935
H83.22852.14541.09042.34583.53594.13942.55011.7801
H92.61962.14171.08972.36342.41163.70133.09351.7801

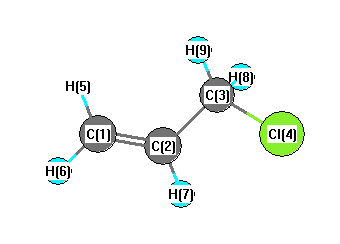
picture of 1-Propene, 3-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 123.150 C1 C2 H7 120.476
C2 C1 H5 121.505 C2 C1 H6 121.336
C2 C3 Cl4 110.779 C2 C3 H8 111.557
C2 C3 H9 111.306 C3 C2 H7 116.373
Cl4 C3 H8 106.106 Cl4 C3 H9 107.406
H5 C1 H6 117.158 H8 C3 H9 109.472
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability