return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH2CH2CH2CH3 (1-Propanamine)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-173.984251
Energy at 298.15K-173.994747
HF Energy-173.316837
Nuclear repulsion energy131.242791
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3541 3354 1.18      
2 A' 3169 3001 29.98      
3 A' 3082 2919 66.88      
4 A' 3077 2914 13.41      
5 A' 3066 2904 17.07      
6 A' 1743 1651 34.50      
7 A' 1548 1466 9.14      
8 A' 1533 1452 1.86      
9 A' 1522 1442 0.23      
10 A' 1449 1372 6.68      
11 A' 1426 1351 6.22      
12 A' 1348 1277 4.44      
13 A' 1174 1112 6.30      
14 A' 1130 1071 12.86      
15 A' 1072 1015 1.63      
16 A' 958 907 174.50      
17 A' 882 836 59.83      
18 A' 456 432 3.25      
19 A' 270 256 4.44      
20 A" 3638 3445 0.00      
21 A" 3164 2996 54.54      
22 A" 3139 2973 34.88      
23 A" 3109 2945 2.88      
24 A" 1543 1461 10.38      
25 A" 1428 1352 0.55      
26 A" 1357 1285 0.69      
27 A" 1274 1206 0.00      
28 A" 1061 1005 0.01      
29 A" 878 832 1.86      
30 A" 750 711 2.32      
31 A" 300 284 50.50      
32 A" 234 222 4.78      
33 A" 137 129 0.67      

Unscaled Zero Point Vibrational Energy (zpe) 27227.6 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 25787.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.83023 0.12322 0.11618

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.429 1.287 0.000
C2 0.000 0.750 0.000
C3 -0.055 -0.778 0.000
N4 -1.389 -1.375 0.000
H5 1.449 2.379 0.000
H6 1.975 0.943 0.883
H7 1.975 0.943 -0.883
H8 -0.540 1.122 0.879
H9 -0.540 1.122 -0.879
H10 0.481 -1.159 -0.877
H11 0.481 -1.159 0.877
H12 -1.902 -1.057 -0.814
H13 -1.902 -1.057 0.814

Atom - Atom Distances (Å)
  C1 C2 C3 N4 H5 H6 H7 H8 H9 H10 H11 H12 H13
C11.52602.54213.87581.09231.09341.09342.16232.16232.76562.76564.15304.1530
C21.52601.52922.53882.18012.17152.17151.09671.09672.15512.15512.74702.7470
C32.54211.52921.46163.49662.80332.80332.14932.14931.09541.09542.03782.0378
N43.87582.53881.46164.70594.17914.17912.78022.78022.07572.07571.01331.0133
H51.09232.18013.49664.70591.76531.76532.51182.51183.77133.77134.86814.8681
H61.09342.17152.80334.17911.76531.76542.52133.07573.12192.57904.68044.3626
H71.09342.17152.80334.17911.76531.76543.07572.52132.57903.12194.36264.6804
H82.16231.09672.14932.78022.51182.52133.07571.75773.05422.49943.07732.5708
H92.16231.09672.14932.78022.51183.07572.52131.75772.49943.05422.57083.0773
H102.76562.15511.09542.07573.77133.12192.57903.05422.49941.75312.38522.9228
H112.76562.15511.09542.07573.77132.57903.12192.49943.05421.75312.92282.3852
H124.15302.74702.03781.01334.86814.68044.36263.07732.57082.38522.92281.6278
H134.15302.74702.03781.01334.86814.36264.68042.57083.07732.92282.38521.6278

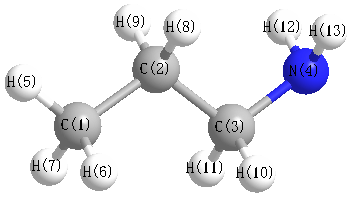
picture of 1-Propanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.625 C1 C2 H8 109.990
C1 C2 H9 109.990 C2 C1 H5 111.670
C2 C1 H6 110.922 C2 C1 H7 110.922
C2 C3 N4 116.158 C2 C3 H10 109.285
C2 C3 H11 109.285 C3 C2 H8 108.758
C3 C2 H9 108.758 C3 N4 H12 109.514
C3 N4 H13 109.514 N4 C3 H10 107.679
N4 C3 H11 107.679 H5 C1 H6 107.741
H5 C1 H7 107.741 H6 C1 H7 107.672
H8 C2 H9 106.522 H10 C3 H11 106.305
H12 N4 H13 106.875
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability