Jump to
S1C2
Vibrational Frequencies calculated at B3PW91/6-311G*
Geometric Data calculated at B3PW91/6-311G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -207.920347 |
| Energy at 298.15K | -207.923738 |
| HF Energy | -207.920347 |
| Nuclear repulsion energy | 102.387254 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3810 |
3668 |
26.83 |
|
|
|
| 2 |
A |
3120 |
3003 |
5.36 |
|
|
|
| 3 |
A |
3024 |
2911 |
29.69 |
|
|
|
| 4 |
A |
2364 |
2276 |
0.37 |
|
|
|
| 5 |
A |
1493 |
1437 |
5.14 |
|
|
|
| 6 |
A |
1428 |
1375 |
47.41 |
|
|
|
| 7 |
A |
1385 |
1333 |
1.37 |
|
|
|
| 8 |
A |
1237 |
1191 |
23.88 |
|
|
|
| 9 |
A |
1098 |
1057 |
103.52 |
|
|
|
| 10 |
A |
992 |
955 |
23.73 |
|
|
|
| 11 |
A |
906 |
872 |
16.25 |
|
|
|
| 12 |
A |
590 |
568 |
2.34 |
|
|
|
| 13 |
A |
403 |
388 |
77.28 |
|
|
|
| 14 |
A |
316 |
304 |
98.55 |
|
|
|
| 15 |
A |
217 |
209 |
9.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11190.8 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 10773.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.568 |
0.586 |
0.038 |
| C2 |
0.824 |
0.116 |
-0.007 |
| O3 |
-1.505 |
-0.450 |
-0.111 |
| H4 |
-0.707 |
1.151 |
0.970 |
| H5 |
-0.732 |
1.277 |
-0.791 |
| H6 |
-1.408 |
-1.068 |
0.621 |
| N7 |
1.907 |
-0.281 |
-0.014 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4704 | 1.4050 | 1.0987 | 1.0919 | 1.9446 | 2.6230 |
C2 | 1.4704 | | 2.3993 | 2.0908 | 2.0946 | 2.6034 | 1.1530 | O3 | 1.4050 | 2.3993 | | 2.0905 | 2.0110 | 0.9626 | 3.4175 | H4 | 1.0987 | 2.0908 | 2.0905 | | 1.7656 | 2.3538 | 3.1390 | H5 | 1.0919 | 2.0946 | 2.0110 | 1.7656 | | 2.8198 | 3.1617 | H6 | 1.9446 | 2.6034 | 0.9626 | 2.3538 | 2.8198 | | 3.4657 | N7 | 2.6230 | 1.1530 | 3.4175 | 3.1390 | 3.1617 | 3.4657 | |
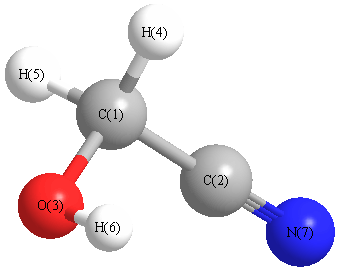 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
178.013 |
|
C1 |
O3 |
H6 |
109.002 |
| C2 |
C1 |
O3 |
113.096 |
|
C2 |
C1 |
H4 |
108.074 |
| C2 |
C1 |
H5 |
108.768 |
|
O3 |
C1 |
H4 |
112.656 |
| O3 |
C1 |
H5 |
106.626 |
|
H4 |
C1 |
H5 |
107.410 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.230 |
|
|
|
| 2 |
C |
0.110 |
|
|
|
| 3 |
O |
-0.532 |
|
|
|
| 4 |
H |
0.248 |
|
|
|
| 5 |
H |
0.269 |
|
|
|
| 6 |
H |
0.403 |
|
|
|
| 7 |
N |
-0.269 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.378 |
1.108 |
1.442 |
2.994 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.843 |
1.279 |
-2.119 |
| y |
1.279 |
-20.444 |
-1.860 |
| z |
-2.119 |
-1.860 |
-21.661 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.790 |
1.279 |
-2.119 |
| y |
1.279 |
6.308 |
-1.860 |
| z |
-2.119 |
-1.860 |
4.482 |
|
| Polar |
|---|
| 3z2-r2 | 8.964 |
|---|
| x2-y2 | -11.398 |
|---|
| xy | 1.279 |
|---|
| xz | -2.119 |
|---|
| yz | -1.860 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.459 |
-0.457 |
-0.037 |
| y |
-0.457 |
3.665 |
-0.127 |
| z |
-0.037 |
-0.127 |
3.119 |
<r2> (average value of r
2) Å
2
| <r2> |
78.476 |
| (<r2>)1/2 |
8.859 |
