Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Geometric Data calculated at MP2=FULL/6-311G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -207.510919 |
| Energy at 298.15K | -207.514311 |
| HF Energy | -206.809563 |
| Nuclear repulsion energy | 101.829118 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3847 |
3644 |
35.34 |
55.66 |
0.30 |
0.46 |
| 2 |
A |
3187 |
3019 |
4.05 |
72.00 |
0.45 |
0.62 |
| 3 |
A |
3090 |
2926 |
24.67 |
105.48 |
0.14 |
0.24 |
| 4 |
A |
2200 |
2084 |
8.79 |
31.74 |
0.23 |
0.38 |
| 5 |
A |
1533 |
1452 |
3.94 |
10.95 |
0.69 |
0.82 |
| 6 |
A |
1450 |
1373 |
45.56 |
6.56 |
0.73 |
0.84 |
| 7 |
A |
1407 |
1333 |
1.65 |
5.66 |
0.75 |
0.86 |
| 8 |
A |
1257 |
1190 |
24.72 |
3.09 |
0.66 |
0.80 |
| 9 |
A |
1114 |
1055 |
96.28 |
5.55 |
0.31 |
0.47 |
| 10 |
A |
1009 |
956 |
24.33 |
0.43 |
0.48 |
0.65 |
| 11 |
A |
913 |
864 |
19.19 |
2.60 |
0.13 |
0.23 |
| 12 |
A |
585 |
554 |
3.98 |
1.74 |
0.26 |
0.42 |
| 13 |
A |
403 |
382 |
111.29 |
0.94 |
0.73 |
0.85 |
| 14 |
A |
314 |
297 |
68.62 |
4.37 |
0.75 |
0.86 |
| 15 |
A |
214 |
203 |
8.71 |
3.90 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11261.3 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 10665.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.578 |
0.591 |
0.038 |
| C2 |
0.821 |
0.121 |
-0.006 |
| O3 |
-1.506 |
-0.458 |
-0.111 |
| H4 |
-0.722 |
1.149 |
0.970 |
| H5 |
-0.747 |
1.274 |
-0.794 |
| H6 |
-1.399 |
-1.066 |
0.625 |
| N7 |
1.923 |
-0.281 |
-0.014 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4760 | 1.4088 | 1.0959 | 1.0896 | 1.9406 | 2.6487 |
C2 | 1.4760 | | 2.4005 | 2.0947 | 2.0995 | 2.5956 | 1.1731 | O3 | 1.4088 | 2.4005 | | 2.0896 | 2.0107 | 0.9610 | 3.4350 | H4 | 1.0959 | 2.0947 | 2.0896 | | 1.7688 | 2.3418 | 3.1633 | H5 | 1.0896 | 2.0995 | 2.0107 | 1.7688 | | 2.8136 | 3.1866 | H6 | 1.9406 | 2.5956 | 0.9610 | 2.3418 | 2.8136 | | 3.4729 | N7 | 2.6487 | 1.1731 | 3.4350 | 3.1633 | 3.1866 | 3.4729 | |
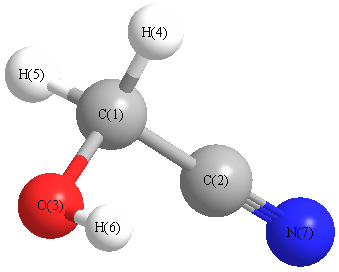 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
178.012 |
|
C1 |
O3 |
H6 |
108.465 |
| C2 |
C1 |
O3 |
112.611 |
|
C2 |
C1 |
H4 |
108.160 |
| C2 |
C1 |
H5 |
108.903 |
|
O3 |
C1 |
H4 |
112.480 |
| O3 |
C1 |
H5 |
106.476 |
|
H4 |
C1 |
H5 |
108.064 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
