Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
Geometric Data calculated at B2PLYP=FULL/6-311G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -207.834413 |
| Energy at 298.15K | -207.837801 |
| HF Energy | -207.604449 |
| Nuclear repulsion energy | 102.076293 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3817 |
3817 |
27.33 |
62.74 |
0.30 |
0.46 |
| 2 |
A |
3153 |
3153 |
5.81 |
79.61 |
0.40 |
0.57 |
| 3 |
A |
3058 |
3058 |
29.13 |
110.35 |
0.16 |
0.27 |
| 4 |
A |
2290 |
2290 |
1.40 |
47.83 |
0.23 |
0.38 |
| 5 |
A |
1521 |
1521 |
3.72 |
11.69 |
0.68 |
0.81 |
| 6 |
A |
1444 |
1444 |
47.12 |
6.95 |
0.72 |
0.84 |
| 7 |
A |
1400 |
1400 |
1.91 |
5.94 |
0.75 |
0.86 |
| 8 |
A |
1249 |
1249 |
25.84 |
3.19 |
0.66 |
0.79 |
| 9 |
A |
1091 |
1091 |
96.54 |
6.42 |
0.32 |
0.48 |
| 10 |
A |
1005 |
1005 |
26.25 |
0.48 |
0.55 |
0.71 |
| 11 |
A |
903 |
903 |
18.83 |
2.23 |
0.13 |
0.23 |
| 12 |
A |
584 |
584 |
2.77 |
2.00 |
0.26 |
0.42 |
| 13 |
A |
401 |
401 |
88.86 |
1.04 |
0.73 |
0.84 |
| 14 |
A |
316 |
316 |
86.84 |
4.79 |
0.75 |
0.86 |
| 15 |
A |
216 |
216 |
9.02 |
4.26 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11223.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 11223.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.572 |
0.590 |
0.038 |
| C2 |
0.825 |
0.118 |
-0.006 |
| O3 |
-1.507 |
-0.455 |
-0.111 |
| H4 |
-0.714 |
1.150 |
0.968 |
| H5 |
-0.739 |
1.275 |
-0.792 |
| H6 |
-1.412 |
-1.065 |
0.626 |
| N7 |
1.914 |
-0.282 |
-0.014 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4754 | 1.4103 | 1.0953 | 1.0890 | 1.9470 | 2.6351 |
C2 | 1.4754 | | 2.4036 | 2.0943 | 2.0985 | 2.6080 | 1.1600 | O3 | 1.4103 | 2.4036 | | 2.0905 | 2.0114 | 0.9619 | 3.4269 | H4 | 1.0953 | 2.0943 | 2.0905 | | 1.7652 | 2.3473 | 3.1503 | H5 | 1.0890 | 2.0985 | 2.0114 | 1.7652 | | 2.8177 | 3.1732 | H6 | 1.9470 | 2.6080 | 0.9619 | 2.3473 | 2.8177 | | 3.4762 | N7 | 2.6351 | 1.1600 | 3.4269 | 3.1503 | 3.1732 | 3.4762 | |
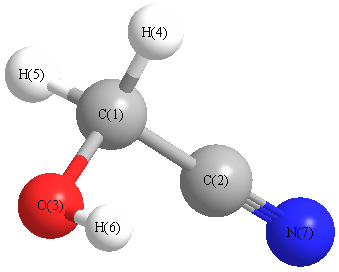 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
178.026 |
|
C1 |
O3 |
H6 |
108.855 |
| C2 |
C1 |
O3 |
112.784 |
|
C2 |
C1 |
H4 |
108.198 |
| C2 |
C1 |
H5 |
108.905 |
|
O3 |
C1 |
H4 |
112.489 |
| O3 |
C1 |
H5 |
106.472 |
|
H4 |
C1 |
H5 |
107.827 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
