return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6O (2-Propen-1-ol)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-192.628690
Energy at 298.15K-192.634957
HF Energy-191.956894
Nuclear repulsion energy116.669212
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3854 3650 16.19      
2 A 3282 3109 14.07      
3 A 3207 3037 6.48      
4 A 3183 3014 8.68      
5 A 3082 2919 49.46      
6 A 3033 2872 56.58      
7 A 1718 1627 0.72      
8 A 1557 1475 1.79      
9 A 1506 1427 13.17      
10 A 1452 1375 3.47      
11 A 1325 1255 2.66      
12 A 1308 1239 26.01      
13 A 1268 1201 67.48      
14 A 1188 1125 15.43      
15 A 1101 1042 75.92      
16 A 1024 970 26.28      
17 A 984 932 5.76      
18 A 940 891 3.20      
19 A 920 872 44.82      
20 A 657 623 5.13      
21 A 453 429 4.15      
22 A 341 323 14.06      
23 A 270 256 145.86      
24 A 115 109 1.17      

Unscaled Zero Point Vibrational Energy (zpe) 18884.1 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 17885.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.92136 0.14433 0.13872

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.588 0.456 0.286
C2 -0.662 -0.362 0.260
C3 -1.831 0.077 -0.218
O4 1.616 -0.301 -0.342
H5 0.419 1.411 -0.228
H6 0.855 0.676 1.331
H7 -0.574 -1.366 0.670
H8 -2.720 -0.543 -0.198
H9 -1.932 1.070 -0.645
H10 2.454 0.133 -0.163

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8 H9 H10
C11.49392.50031.42311.09831.09972.19443.48982.75621.9468
C21.49391.33712.35722.13332.12671.08802.11652.11713.1835
C32.50031.33713.47062.61653.15772.10931.08411.08594.2864
O41.42312.35723.47062.09252.08082.63724.34603.81650.9601
H51.09832.13332.61652.09251.77803.08303.69852.41242.4045
H61.09972.12673.15772.08081.77802.57764.07473.43882.2550
H72.19441.08802.10932.63723.08302.57762.45683.08323.4798
H83.48982.11651.08414.34603.69854.07472.45681.85045.2188
H92.75622.11711.08593.81652.41243.43883.08321.85044.5115
H101.94683.18354.28640.96012.40452.25503.47985.21884.5115

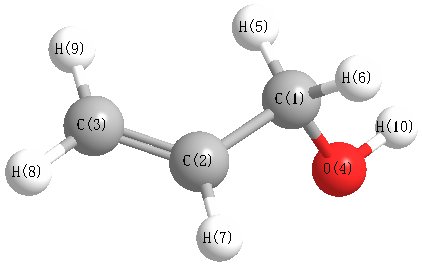
picture of 2-Propen-1-ol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 123.965 C1 C2 H7 115.508
C1 O4 H10 107.972 C2 C1 O4 107.799
C2 C1 H5 109.822 C2 C1 H6 109.221
C2 C3 H8 121.536 C2 C3 H9 121.450
C3 C2 H7 120.525 O4 C1 H5 111.524
O4 C1 H6 110.483 H5 C1 H6 107.974
H8 C3 H9 117.014
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability