Jump to
S1C2
Vibrational Frequencies calculated at B3PW91/6-311G*
Geometric Data calculated at B3PW91/6-311G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -271.740109 |
| Energy at 298.15K | -271.750648 |
| Nuclear repulsion energy | 236.146782 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3157 |
3039 |
14.22 |
|
|
|
| 2 |
A |
3113 |
2996 |
61.52 |
|
|
|
| 3 |
A |
3109 |
2993 |
51.88 |
|
|
|
| 4 |
A |
3107 |
2991 |
9.30 |
|
|
|
| 5 |
A |
3101 |
2985 |
5.25 |
|
|
|
| 6 |
A |
3067 |
2952 |
6.86 |
|
|
|
| 7 |
A |
3043 |
2930 |
4.29 |
|
|
|
| 8 |
A |
3037 |
2923 |
52.03 |
|
|
|
| 9 |
A |
3035 |
2922 |
12.99 |
|
|
|
| 10 |
A |
3020 |
2907 |
6.56 |
|
|
|
| 11 |
A |
1813 |
1745 |
193.38 |
|
|
|
| 12 |
A |
1519 |
1462 |
8.18 |
|
|
|
| 13 |
A |
1512 |
1456 |
10.34 |
|
|
|
| 14 |
A |
1504 |
1448 |
1.93 |
|
|
|
| 15 |
A |
1491 |
1436 |
16.61 |
|
|
|
| 16 |
A |
1478 |
1423 |
15.80 |
|
|
|
| 17 |
A |
1473 |
1418 |
3.01 |
|
|
|
| 18 |
A |
1420 |
1367 |
5.70 |
|
|
|
| 19 |
A |
1393 |
1341 |
54.37 |
|
|
|
| 20 |
A |
1385 |
1333 |
4.83 |
|
|
|
| 21 |
A |
1339 |
1289 |
3.72 |
|
|
|
| 22 |
A |
1302 |
1254 |
15.07 |
|
|
|
| 23 |
A |
1276 |
1229 |
23.16 |
|
|
|
| 24 |
A |
1229 |
1183 |
42.04 |
|
|
|
| 25 |
A |
1136 |
1094 |
3.28 |
|
|
|
| 26 |
A |
1123 |
1081 |
8.15 |
|
|
|
| 27 |
A |
1062 |
1022 |
0.60 |
|
|
|
| 28 |
A |
1004 |
967 |
3.27 |
|
|
|
| 29 |
A |
970 |
933 |
6.12 |
|
|
|
| 30 |
A |
923 |
888 |
1.39 |
|
|
|
| 31 |
A |
858 |
826 |
2.56 |
|
|
|
| 32 |
A |
788 |
758 |
3.09 |
|
|
|
| 33 |
A |
746 |
718 |
2.87 |
|
|
|
| 34 |
A |
574 |
553 |
4.71 |
|
|
|
| 35 |
A |
526 |
507 |
12.73 |
|
|
|
| 36 |
A |
411 |
396 |
1.62 |
|
|
|
| 37 |
A |
320 |
308 |
0.34 |
|
|
|
| 38 |
A |
252 |
243 |
0.00 |
|
|
|
| 39 |
A |
182 |
175 |
1.05 |
|
|
|
| 40 |
A |
132 |
127 |
0.22 |
|
|
|
| 41 |
A |
85 |
82 |
0.77 |
|
|
|
| 42 |
A |
52 |
50 |
2.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31031.3 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 29873.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.748 |
-0.290 |
-0.006 |
| C2 |
-1.404 |
0.426 |
0.053 |
| C3 |
-0.225 |
-0.531 |
-0.044 |
| C4 |
1.134 |
0.147 |
-0.017 |
| C5 |
2.338 |
-0.770 |
0.027 |
| O6 |
1.252 |
1.350 |
-0.024 |
| H7 |
-3.578 |
0.418 |
0.067 |
| H8 |
-2.870 |
-0.842 |
-0.944 |
| H9 |
-2.859 |
-1.009 |
0.813 |
| H10 |
-1.332 |
1.163 |
-0.752 |
| H11 |
-1.323 |
1.001 |
0.981 |
| H12 |
-0.255 |
-1.279 |
0.761 |
| H13 |
-0.280 |
-1.121 |
-0.970 |
| H14 |
3.250 |
-0.195 |
-0.133 |
| H15 |
2.396 |
-1.258 |
1.006 |
| H16 |
2.265 |
-1.566 |
-0.720 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
| C1 | | 1.5242 | 2.5353 | 3.9067 | 5.1090 | 4.3234 | 1.0936 | 1.0953 | 1.0952 | 2.1627 | 2.1613 | 2.7895 | 2.7771 | 6.0002 | 5.3315 | 5.2224 |
C2 | 1.5242 | | 1.5218 | 2.5544 | 3.9288 | 2.8132 | 2.1744 | 2.1796 | 2.1805 | 1.0943 | 1.0951 | 2.1744 | 2.1689 | 4.6988 | 4.2645 | 4.2462 | C3 | 2.5353 | 1.5218 | | 1.5183 | 2.5749 | 2.3909 | 3.4872 | 2.8117 | 2.8115 | 2.1440 | 2.1459 | 1.0992 | 1.1001 | 3.4918 | 2.9155 | 2.7800 | C4 | 3.9067 | 2.5544 | 1.5183 | | 1.5139 | 1.2087 | 4.7210 | 4.2273 | 4.2392 | 2.7665 | 2.7868 | 2.1368 | 2.1252 | 2.1463 | 2.1477 | 2.1697 | C5 | 5.1090 | 3.9288 | 2.5749 | 1.5139 | | 2.3821 | 6.0348 | 5.2985 | 5.2619 | 4.2203 | 4.1778 | 2.7425 | 2.8238 | 1.0898 | 1.0958 | 1.0942 | O6 | 4.3234 | 2.8132 | 2.3909 | 1.2087 | 2.3821 | | 4.9204 | 4.7584 | 4.8131 | 2.6909 | 2.7867 | 3.1296 | 3.0576 | 2.5273 | 3.0279 | 3.1644 | H7 | 1.0936 | 2.1744 | 3.4872 | 4.7210 | 6.0348 | 4.9204 | | 1.7637 | 1.7638 | 2.5050 | 2.5022 | 3.7956 | 3.7846 | 6.8586 | 6.2759 | 6.2213 | H8 | 1.0953 | 2.1796 | 2.8117 | 4.2273 | 5.2985 | 4.7584 | 1.7637 | | 1.7650 | 2.5348 | 3.0815 | 3.1523 | 2.6051 | 6.2074 | 5.6312 | 5.1910 | H9 | 1.0952 | 2.1805 | 2.8115 | 4.2392 | 5.2619 | 4.8131 | 1.7638 | 1.7650 | | 3.0828 | 2.5351 | 2.6187 | 3.1377 | 6.2353 | 5.2648 | 5.3777 | H10 | 2.1627 | 1.0943 | 2.1440 | 2.7665 | 4.2203 | 2.6909 | 2.5050 | 2.5348 | 3.0828 | | 1.7415 | 3.0680 | 2.5241 | 4.8184 | 4.7803 | 4.5154 | H11 | 2.1613 | 1.0951 | 2.1459 | 2.7868 | 4.1778 | 2.7867 | 2.5022 | 3.0815 | 2.5351 | 1.7415 | | 2.5273 | 3.0662 | 4.8566 | 4.3518 | 4.7290 | H12 | 2.7895 | 2.1744 | 1.0992 | 2.1368 | 2.7425 | 3.1296 | 3.7956 | 3.1523 | 2.6187 | 3.0680 | 2.5273 | | 1.7389 | 3.7759 | 2.6625 | 2.9371 | H13 | 2.7771 | 2.1689 | 1.1001 | 2.1252 | 2.8238 | 3.0576 | 3.7846 | 2.6051 | 3.1377 | 2.5241 | 3.0662 | 1.7389 | | 3.7446 | 3.3302 | 2.5962 | H14 | 6.0002 | 4.6988 | 3.4918 | 2.1463 | 1.0898 | 2.5273 | 6.8586 | 6.2074 | 6.2353 | 4.8184 | 4.8566 | 3.7759 | 3.7446 | | 1.7768 | 1.7877 | H15 | 5.3315 | 4.2645 | 2.9155 | 2.1477 | 1.0958 | 3.0279 | 6.2759 | 5.6312 | 5.2648 | 4.7803 | 4.3518 | 2.6625 | 3.3302 | 1.7768 | | 1.7584 | H16 | 5.2224 | 4.2462 | 2.7800 | 2.1697 | 1.0942 | 3.1644 | 6.2213 | 5.1910 | 5.3777 | 4.5154 | 4.7290 | 2.9371 | 2.5962 | 1.7877 | 1.7584 | |
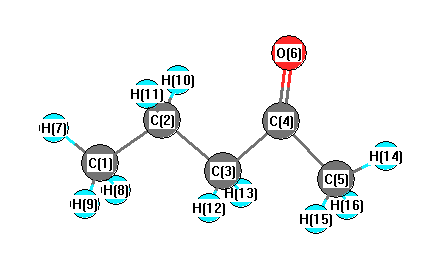 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.683 |
|
C1 |
C2 |
H10 |
110.290 |
| C1 |
C2 |
H11 |
110.133 |
|
C2 |
C1 |
H7 |
111.262 |
| C2 |
C1 |
H8 |
111.585 |
|
C2 |
C1 |
H9 |
111.657 |
| C2 |
C3 |
C4 |
114.326 |
|
C2 |
C3 |
H12 |
111.089 |
| C2 |
C3 |
H13 |
110.599 |
|
C3 |
C2 |
H10 |
108.985 |
| C3 |
C2 |
H11 |
109.085 |
|
C3 |
C4 |
C5 |
116.241 |
| C3 |
C4 |
O6 |
122.092 |
|
C4 |
C3 |
H12 |
108.376 |
| C4 |
C3 |
H13 |
107.431 |
|
C4 |
C5 |
H14 |
109.970 |
| C4 |
C5 |
H15 |
109.733 |
|
C4 |
C5 |
H16 |
111.581 |
| C5 |
C4 |
O6 |
121.666 |
|
H7 |
C1 |
H8 |
107.365 |
| H7 |
C1 |
H9 |
107.374 |
|
H8 |
C1 |
H9 |
107.362 |
| H10 |
C2 |
H11 |
105.396 |
|
H12 |
C3 |
H13 |
104.490 |
| H14 |
C5 |
H15 |
108.767 |
|
H14 |
C5 |
H16 |
109.877 |
| H15 |
C5 |
H16 |
106.826 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.648 |
|
|
|
| 2 |
H |
0.219 |
|
|
|
| 3 |
H |
0.226 |
|
|
|
| 4 |
H |
0.226 |
|
|
|
| 5 |
C |
-0.456 |
|
|
|
| 6 |
H |
0.224 |
|
|
|
| 7 |
H |
0.229 |
|
|
|
| 8 |
C |
-0.528 |
|
|
|
| 9 |
H |
0.244 |
|
|
|
| 10 |
C |
-0.747 |
|
|
|
| 11 |
H |
0.253 |
|
|
|
| 12 |
H |
0.235 |
|
|
|
| 13 |
C |
0.344 |
|
|
|
| 14 |
O |
-0.302 |
|
|
|
| 15 |
H |
0.230 |
|
|
|
| 16 |
H |
0.249 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.938 |
2.163 |
0.617 |
2.969 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.303 |
-4.622 |
-1.359 |
| y |
-4.622 |
-39.959 |
-0.616 |
| z |
-1.359 |
-0.616 |
-37.413 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.617 |
-4.622 |
-1.359 |
| y |
-4.622 |
-0.101 |
-0.616 |
| z |
-1.359 |
-0.616 |
3.718 |
|
| Polar |
|---|
| 3z2-r2 | 7.436 |
|---|
| x2-y2 | -2.344 |
|---|
| xy | -4.622 |
|---|
| xz | -1.359 |
|---|
| yz | -0.616 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
