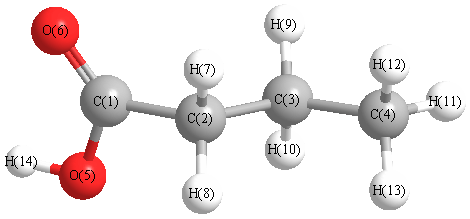Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3755 |
3615 |
36.75 |
|
|
|
| 2 |
A |
3130 |
3013 |
35.31 |
|
|
|
| 3 |
A |
3112 |
2996 |
36.44 |
|
|
|
| 4 |
A |
3108 |
2992 |
35.28 |
|
|
|
| 5 |
A |
3082 |
2967 |
0.10 |
|
|
|
| 6 |
A |
3061 |
2947 |
25.07 |
|
|
|
| 7 |
A |
3044 |
2930 |
12.15 |
|
|
|
| 8 |
A |
3036 |
2923 |
28.18 |
|
|
|
| 9 |
A |
1850 |
1781 |
336.93 |
|
|
|
| 10 |
A |
1520 |
1463 |
10.48 |
|
|
|
| 11 |
A |
1513 |
1456 |
10.98 |
|
|
|
| 12 |
A |
1506 |
1450 |
1.98 |
|
|
|
| 13 |
A |
1484 |
1429 |
7.96 |
|
|
|
| 14 |
A |
1424 |
1371 |
5.03 |
|
|
|
| 15 |
A |
1395 |
1343 |
39.91 |
|
|
|
| 16 |
A |
1383 |
1331 |
19.84 |
|
|
|
| 17 |
A |
1338 |
1288 |
1.81 |
|
|
|
| 18 |
A |
1299 |
1250 |
0.19 |
|
|
|
| 19 |
A |
1273 |
1226 |
41.73 |
|
|
|
| 20 |
A |
1219 |
1173 |
148.34 |
|
|
|
| 21 |
A |
1122 |
1080 |
8.15 |
|
|
|
| 22 |
A |
1089 |
1048 |
85.66 |
|
|
|
| 23 |
A |
1061 |
1022 |
5.42 |
|
|
|
| 24 |
A |
929 |
895 |
1.49 |
|
|
|
| 25 |
A |
900 |
866 |
2.37 |
|
|
|
| 26 |
A |
880 |
848 |
9.54 |
|
|
|
| 27 |
A |
763 |
734 |
34.28 |
|
|
|
| 28 |
A |
743 |
716 |
31.81 |
|
|
|
| 29 |
A |
642 |
618 |
80.44 |
|
|
|
| 30 |
A |
588 |
566 |
48.21 |
|
|
|
| 31 |
A |
431 |
415 |
3.17 |
|
|
|
| 32 |
A |
332 |
320 |
1.05 |
|
|
|
| 33 |
A |
246 |
237 |
0.02 |
|
|
|
| 34 |
A |
187 |
180 |
0.02 |
|
|
|
| 35 |
A |
98 |
94 |
0.22 |
|
|
|
| 36 |
A |
34 |
33 |
0.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26286.3 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 25305.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.426 |
|
|
|
| 2 |
C |
-0.503 |
|
|
|
| 3 |
C |
-0.451 |
|
|
|
| 4 |
C |
-0.653 |
|
|
|
| 5 |
O |
-0.511 |
|
|
|
| 6 |
O |
-0.342 |
|
|
|
| 7 |
H |
0.249 |
|
|
|
| 8 |
H |
0.243 |
|
|
|
| 9 |
H |
0.230 |
|
|
|
| 10 |
H |
0.238 |
|
|
|
| 11 |
H |
0.228 |
|
|
|
| 12 |
H |
0.223 |
|
|
|
| 13 |
H |
0.221 |
|
|
|
| 14 |
H |
0.402 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.048 |
1.314 |
0.231 |
1.697 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.645 |
5.365 |
0.116 |
| y |
5.365 |
-39.275 |
-1.513 |
| z |
0.116 |
-1.513 |
-36.137 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.061 |
5.365 |
0.116 |
| y |
5.365 |
-3.384 |
-1.513 |
| z |
0.116 |
-1.513 |
1.323 |
|
| Polar |
|---|
| 3z2-r2 | 2.647 |
|---|
| x2-y2 | 3.630 |
|---|
| xy | 5.365 |
|---|
| xz | 0.116 |
|---|
| yz | -1.513 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.911 |
-0.063 |
-0.331 |
| y |
-0.063 |
7.285 |
0.042 |
| z |
-0.331 |
0.042 |
6.188 |
<r2> (average value of r
2) Å
2
| <r2> |
208.063 |
| (<r2>)1/2 |
14.424 |