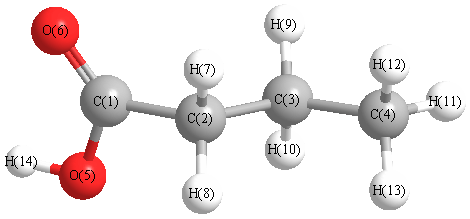Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3788 |
3631 |
41.22 |
|
|
|
| 2 |
A |
3146 |
3016 |
33.52 |
|
|
|
| 3 |
A |
3130 |
3000 |
34.72 |
|
|
|
| 4 |
A |
3125 |
2995 |
32.90 |
|
|
|
| 5 |
A |
3099 |
2971 |
0.06 |
|
|
|
| 6 |
A |
3074 |
2946 |
25.62 |
|
|
|
| 7 |
A |
3059 |
2932 |
10.22 |
|
|
|
| 8 |
A |
3050 |
2924 |
26.93 |
|
|
|
| 9 |
A |
1871 |
1793 |
345.51 |
|
|
|
| 10 |
A |
1522 |
1458 |
10.91 |
|
|
|
| 11 |
A |
1515 |
1452 |
11.53 |
|
|
|
| 12 |
A |
1508 |
1445 |
2.27 |
|
|
|
| 13 |
A |
1485 |
1423 |
9.58 |
|
|
|
| 14 |
A |
1427 |
1368 |
7.57 |
|
|
|
| 15 |
A |
1407 |
1348 |
49.64 |
|
|
|
| 16 |
A |
1390 |
1332 |
13.51 |
|
|
|
| 17 |
A |
1341 |
1286 |
1.49 |
|
|
|
| 18 |
A |
1300 |
1246 |
0.26 |
|
|
|
| 19 |
A |
1278 |
1225 |
54.63 |
|
|
|
| 20 |
A |
1229 |
1178 |
141.05 |
|
|
|
| 21 |
A |
1129 |
1082 |
8.12 |
|
|
|
| 22 |
A |
1098 |
1052 |
68.59 |
|
|
|
| 23 |
A |
1074 |
1029 |
7.73 |
|
|
|
| 24 |
A |
939 |
900 |
1.83 |
|
|
|
| 25 |
A |
904 |
866 |
2.28 |
|
|
|
| 26 |
A |
886 |
849 |
8.16 |
|
|
|
| 27 |
A |
766 |
734 |
39.91 |
|
|
|
| 28 |
A |
743 |
712 |
28.99 |
|
|
|
| 29 |
A |
645 |
618 |
80.78 |
|
|
|
| 30 |
A |
590 |
566 |
48.15 |
|
|
|
| 31 |
A |
434 |
416 |
3.20 |
|
|
|
| 32 |
A |
335 |
321 |
1.09 |
|
|
|
| 33 |
A |
251 |
241 |
0.03 |
|
|
|
| 34 |
A |
189 |
181 |
0.02 |
|
|
|
| 35 |
A |
103 |
98 |
0.21 |
|
|
|
| 36 |
A |
40 |
38 |
0.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26434.1 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 25337.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.431 |
|
|
|
| 2 |
C |
-0.517 |
|
|
|
| 3 |
C |
-0.460 |
|
|
|
| 4 |
C |
-0.664 |
|
|
|
| 5 |
O |
-0.513 |
|
|
|
| 6 |
O |
-0.342 |
|
|
|
| 7 |
H |
0.253 |
|
|
|
| 8 |
H |
0.248 |
|
|
|
| 9 |
H |
0.233 |
|
|
|
| 10 |
H |
0.242 |
|
|
|
| 11 |
H |
0.232 |
|
|
|
| 12 |
H |
0.227 |
|
|
|
| 13 |
H |
0.225 |
|
|
|
| 14 |
H |
0.405 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.059 |
1.313 |
0.257 |
1.706 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.986 |
5.488 |
0.269 |
| y |
5.488 |
-39.097 |
-1.477 |
| z |
0.269 |
-1.477 |
-36.245 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.684 |
5.488 |
0.269 |
| y |
5.488 |
-2.981 |
-1.477 |
| z |
0.269 |
-1.477 |
1.297 |
|
| Polar |
|---|
| 3z2-r2 | 2.594 |
|---|
| x2-y2 | 3.111 |
|---|
| xy | 5.488 |
|---|
| xz | 0.269 |
|---|
| yz | -1.477 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.849 |
-0.070 |
-0.325 |
| y |
-0.070 |
7.266 |
0.044 |
| z |
-0.325 |
0.044 |
6.155 |
<r2> (average value of r
2) Å
2
| <r2> |
207.201 |
| (<r2>)1/2 |
14.394 |