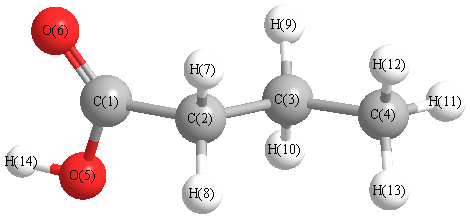return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8O2 (Butanoic acid)

using model chemistry: B2PLYP=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP=FULL/6-311G*
 hartrees
Energy at 0K-307.505839
Energy at 298.15K-307.515281
HF Energy-307.163356
Nuclear repulsion energy238.947308
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3756 3756 40.76      
2 A 3149 3149 37.41      
3 A 3131 3131 36.59      
4 A 3128 3128 34.22      
5 A 3104 3104 0.12      
6 A 3083 3083 23.72      
7 A 3064 3064 12.92      
8 A 3056 3056 27.09      
9 A 1829 1829 301.53      
10 A 1542 1542 9.57      
11 A 1535 1535 9.98      
12 A 1528 1528 1.58      
13 A 1509 1509 6.67      
14 A 1446 1446 4.11      
15 A 1410 1410 19.79      
16 A 1393 1393 29.53      
17 A 1351 1351 2.81      
18 A 1315 1315 0.14      
19 A 1288 1288 27.30      
20 A 1229 1229 148.42      
21 A 1133 1133 7.11      
22 A 1100 1100 101.84      
23 A 1064 1064 5.54      
24 A 932 932 1.24      
25 A 908 908 2.41      
26 A 884 884 11.36      
27 A 767 767 19.41      
28 A 745 745 43.55      
29 A 640 640 80.96      
30 A 588 588 48.48      
31 A 434 434 2.84      
32 A 335 335 1.18      
33 A 248 248 0.03      
34 A 189 189 0.03      
35 A 97 97 0.22      
36 A 35 35 0.27      

Unscaled Zero Point Vibrational Energy (zpe) 26472.2 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 26472.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULL/6-311G*
ABC
0.27434 0.06204 0.05539

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.181 -0.146 0.087
C2 -0.236 -0.341 0.558
C3 -1.258 0.199 -0.455
C4 -2.696 -0.030 0.007
O5 1.530 1.166 0.092
O6 1.937 -1.012 -0.281
H7 -0.381 -1.409 0.719
H8 -0.363 0.174 1.514
H9 -1.097 -0.292 -1.420
H10 -1.078 1.265 -0.610
H11 -3.410 0.356 -0.724
H12 -2.904 -1.094 0.144
H13 -2.890 0.472 0.958
H14 2.435 1.214 -0.245

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 O6 H7 H8 H9 H10 H11 H12 H13 H14
C11.50602.52253.87961.35711.20672.10602.12702.73522.75314.68884.19384.20901.8792
C21.50601.53672.54002.36712.42381.09021.09332.15742.15633.49292.80282.80453.1930
C32.52251.53671.52703.00083.42122.17532.16291.09421.09222.17392.17632.17553.8359
C43.87962.54001.52704.39204.74432.78712.78492.15822.16171.09161.09261.09285.2855
O51.35712.36713.00084.39202.24653.26692.56693.36262.70205.07114.97604.55690.9674
O61.20672.42383.42124.74432.24652.55523.14933.31983.79235.53654.85985.19962.2811
H72.10601.09022.17532.78713.26692.55521.77172.51663.06603.79062.60663.14503.9673
H82.12701.09332.16292.78492.56693.14931.77173.05972.49233.78453.15282.60473.4651
H92.73522.15741.09422.15823.36263.31982.51663.05971.75482.50032.52003.07394.0152
H102.75312.15631.09222.16172.70203.79233.06602.49231.75482.50523.07622.52373.5324
H114.68883.49292.17391.09165.07115.53653.79063.78452.50032.50521.76341.76365.9268
H124.19382.80282.17631.09264.97604.85982.60663.15282.52003.07621.76341.76465.8291
H134.20902.80452.17551.09284.55695.19963.14502.60473.07392.52371.76361.76465.5096
H141.87923.19303.83595.28550.96742.28113.96733.46514.01523.53245.92685.82915.5096

picture of Butanoic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.992 C1 C2 H7 107.343
C1 C2 H8 108.796 C1 O5 H14 106.681
C2 C1 O5 111.430 C2 C1 O6 126.281
C2 C3 C4 112.003 C2 C3 H9 109.013
C2 C3 H10 109.045 C3 C2 H7 110.651
C3 C2 H8 109.496 C3 C4 H11 111.142
C3 C4 H12 111.275 C3 C4 H13 111.201
C4 C3 H9 109.741 C4 C3 H10 110.136
O5 C1 O6 122.276 H7 C2 H8 108.468
H9 C3 H10 106.754 H11 C4 H12 107.670
H11 C4 H13 107.677 H12 C4 H13 107.698
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability