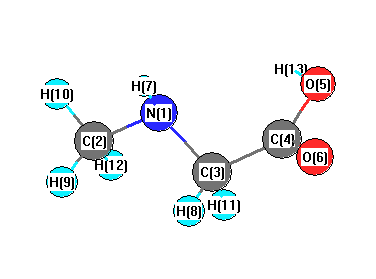
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4056 |
3668 |
166.29 |
|
|
|
| 2 |
A |
3768 |
3408 |
1.85 |
|
|
|
| 3 |
A |
3270 |
2957 |
46.16 |
|
|
|
| 4 |
A |
3242 |
2932 |
27.91 |
|
|
|
| 5 |
A |
3233 |
2924 |
42.94 |
|
|
|
| 6 |
A |
3202 |
2896 |
31.09 |
|
|
|
| 7 |
A |
3151 |
2850 |
79.38 |
|
|
|
| 8 |
A |
2048 |
1852 |
458.24 |
|
|
|
| 9 |
A |
1668 |
1508 |
30.50 |
|
|
|
| 10 |
A |
1643 |
1486 |
4.26 |
|
|
|
| 11 |
A |
1634 |
1478 |
17.27 |
|
|
|
| 12 |
A |
1621 |
1466 |
8.15 |
|
|
|
| 13 |
A |
1597 |
1444 |
1.29 |
|
|
|
| 14 |
A |
1547 |
1399 |
481.49 |
|
|
|
| 15 |
A |
1493 |
1350 |
74.42 |
|
|
|
| 16 |
A |
1414 |
1279 |
9.74 |
|
|
|
| 17 |
A |
1337 |
1209 |
1.24 |
|
|
|
| 18 |
A |
1297 |
1173 |
14.14 |
|
|
|
| 19 |
A |
1263 |
1142 |
28.05 |
|
|
|
| 20 |
A |
1228 |
1110 |
33.93 |
|
|
|
| 21 |
A |
1100 |
995 |
26.81 |
|
|
|
| 22 |
A |
1063 |
962 |
11.68 |
|
|
|
| 23 |
A |
962 |
870 |
31.88 |
|
|
|
| 24 |
A |
928 |
839 |
71.22 |
|
|
|
| 25 |
A |
759 |
686 |
124.43 |
|
|
|
| 26 |
A |
715 |
647 |
4.77 |
|
|
|
| 27 |
A |
642 |
581 |
21.75 |
|
|
|
| 28 |
A |
511 |
462 |
14.30 |
|
|
|
| 29 |
A |
396 |
358 |
2.76 |
|
|
|
| 30 |
A |
288 |
260 |
2.48 |
|
|
|
| 31 |
A |
219 |
198 |
2.86 |
|
|
|
| 32 |
A |
131 |
118 |
3.10 |
|
|
|
| 33 |
A |
90 |
81 |
8.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25756.1 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 23293.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.677 |
|
|
|
| 2 |
C |
-0.451 |
|
|
|
| 3 |
C |
-0.365 |
|
|
|
| 4 |
C |
0.610 |
|
|
|
| 5 |
O |
-0.611 |
|
|
|
| 6 |
O |
-0.437 |
|
|
|
| 7 |
H |
0.339 |
|
|
|
| 8 |
H |
0.240 |
|
|
|
| 9 |
H |
0.199 |
|
|
|
| 10 |
H |
0.220 |
|
|
|
| 11 |
H |
0.249 |
|
|
|
| 12 |
H |
0.222 |
|
|
|
| 13 |
H |
0.462 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
5.570 |
0.050 |
-0.372 |
5.582 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.798 |
-1.189 |
-0.884 |
| y |
-1.189 |
-37.487 |
-0.616 |
| z |
-0.884 |
-0.616 |
-33.241 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.434 |
-1.189 |
-0.884 |
| y |
-1.189 |
1.533 |
-0.616 |
| z |
-0.884 |
-0.616 |
7.901 |
|
| Polar |
|---|
| 3z2-r2 | 15.803 |
|---|
| x2-y2 | -7.311 |
|---|
| xy | -1.189 |
|---|
| xz | -0.884 |
|---|
| yz | -0.616 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.419 |
0.529 |
0.087 |
| y |
0.529 |
6.138 |
-0.003 |
| z |
0.087 |
-0.003 |
5.006 |
<r2> (average value of r
2) Å
2
| <r2> |
197.887 |
| (<r2>)1/2 |
14.067 |