return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5Br (bromobenzene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-2803.954466
Energy at 298.15K-2803.962077
Nuclear repulsion energy430.708244
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3240 3068 5.40      
2 A1 3230 3059 9.42      
3 A1 3209 3040 0.12      
4 A1 1634 1548 22.21      
5 A1 1513 1433 40.91      
6 A1 1214 1149 0.04      
7 A1 1105 1046 17.26      
8 A1 1045 989 15.46      
9 A1 1007 954 13.65      
10 A1 684 648 15.46      
11 A1 324 307 0.98      
12 A2 867 821 0.00      
13 A2 806 763 0.00      
14 A2 393 372 0.00      
15 B1 866 820 0.05      
16 B1 838 793 0.19      
17 B1 714 676 101.70      
18 B1 437 414 7.60      
19 B1 372 353 0.04      
20 B1 165 156 0.61      
21 B2 3237 3065 7.36      
22 B2 3218 3048 7.21      
23 B2 1637 1550 2.38      
24 B2 1480 1402 9.73      
25 B2 1452 1375 0.33      
26 B2 1337 1267 0.67      
27 B2 1197 1133 0.11      
28 B2 1105 1046 4.39      
29 B2 620 587 0.27      
30 B2 253 239 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 19598.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 18561.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.18878 0.03304 0.02812

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Br1 0.000 0.000 1.802
C2 0.000 0.000 -0.093
C3 0.000 1.217 -0.777
C4 0.000 -1.217 -0.777
C5 0.000 1.208 -2.175
C6 0.000 -1.208 -2.175
C7 0.000 0.000 -2.876
H8 0.000 2.153 -0.227
H9 0.000 -2.153 -0.227
H10 0.000 2.152 -2.713
H11 0.000 -2.152 -2.713
H12 0.000 0.000 -3.962

Atom - Atom Distances (Å)
  Br1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12
Br11.89572.85252.85254.15664.15664.67862.95922.95925.00195.00195.7647
C21.89571.39641.39642.40672.40672.78302.15742.15743.39033.39033.8690
C32.85251.39642.43481.39752.79962.42651.08533.41522.14953.88613.4098
C42.85251.39642.43482.79961.39752.42653.41521.08533.88612.14953.4098
C54.15662.40671.39752.79962.41681.39732.16433.88491.08653.40352.1577
C64.15662.40672.79961.39752.41681.39733.88492.16433.40351.08652.1577
C74.67862.78302.42652.42651.39731.39733.41363.41362.15852.15851.0861
H82.95922.15741.08533.41522.16433.88493.41364.30642.48534.97144.3111
H92.95922.15743.41521.08533.88492.16433.41364.30644.97142.48534.3111
H105.00193.39032.14953.88611.08653.40352.15852.48534.97144.30462.4887
H115.00193.39033.88612.14953.40351.08652.15854.97142.48534.30462.4887
H125.76473.86903.40983.40982.15772.15771.08614.31114.31112.48872.4887

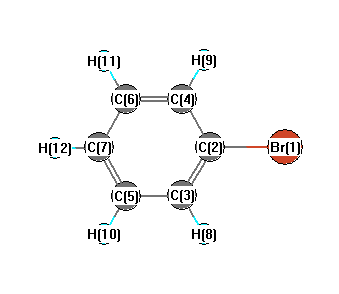
picture of bromobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Br1 C2 C3 119.327 Br1 C2 C4 119.327
C2 C3 C5 118.957 C2 C3 H8 120.241
C2 C4 C6 118.957 C2 C4 H9 120.241
C3 C2 C4 121.347 C3 C5 C7 120.508
C3 C5 H10 119.315 C4 C6 C7 120.508
C4 C6 H11 119.315 C5 C3 H8 120.802
C5 C7 C6 119.723 C5 C7 H12 120.138
C6 C4 H9 120.802 C6 C7 H12 120.138
C7 C5 H10 120.177 C7 C6 H11 120.177
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability