return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5Cl (chlorobenzene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-690.760965
Energy at 298.15K-690.765933
HF Energy-689.665564
Nuclear repulsion energy315.290685
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3240 3069 4.57      
2 A1 3231 3060 9.91      
3 A1 3211 3041 0.16      
4 A1 1642 1555 24.09      
5 A1 1520 1439 47.85      
6 A1 1211 1147 0.01      
7 A1 1134 1074 37.78      
8 A1 1050 994 13.11      
9 A1 1011 957 7.36      
10 A1 721 683 19.55      
11 A1 424 402 1.98      
12 A2 875 829 0.00      
13 A2 810 767 0.00      
14 A2 398 377 0.00      
15 B1 869 823 0.03      
16 B1 841 797 0.50      
17 B1 718 680 102.63      
18 B1 449 425 9.54      
19 B1 414 392 0.02      
20 B1 185 175 0.35      
21 B2 3237 3066 8.43      
22 B2 3219 3049 6.31      
23 B2 1640 1553 2.57      
24 B2 1485 1406 7.79      
25 B2 1453 1377 0.53      
26 B2 1338 1267 0.16      
27 B2 1196 1133 0.11      
28 B2 1106 1047 4.36      
29 B2 622 589 0.28      
30 B2 298 282 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 19772.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 18726.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.18917 0.05238 0.04102

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Cl1 0.000 0.000 2.250
C2 0.000 0.000 0.513
C3 0.000 1.215 -0.173
C4 0.000 -1.215 -0.173
C5 0.000 1.208 -1.569
C6 0.000 -1.208 -1.569
C7 0.000 0.000 -2.271
H8 0.000 2.148 0.382
H9 0.000 -2.148 0.382
H10 0.000 2.152 -2.106
H11 0.000 -2.152 -2.106
H12 0.000 0.000 -3.357

Atom - Atom Distances (Å)
  Cl1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12
Cl11.73712.71062.71064.00614.00614.52172.84712.84714.85954.85955.6077
C21.73711.39551.39552.40762.40762.78462.15232.15233.39063.39063.8706
C32.71061.39552.43061.39672.79702.42521.08543.40902.14903.88343.4087
C42.71061.39552.43062.79701.39672.42523.40901.08543.88342.14903.4087
C54.00612.40761.39672.79702.41621.39732.16593.88241.08643.40322.1580
C64.00612.40762.79701.39672.41621.39733.88242.16593.40321.08642.1580
C74.52172.78462.42522.42521.39731.39733.41403.41402.15872.15871.0860
H82.84712.15231.08543.40902.16593.88243.41404.29672.48854.96874.3125
H92.84712.15233.40901.08543.88242.16593.41404.29674.96872.48854.3125
H104.85953.39062.14903.88341.08643.40322.15872.48854.96874.30482.4894
H114.85953.39063.88342.14903.40321.08642.15874.96872.48854.30482.4894
H125.60773.87063.40873.40872.15802.15801.08604.31254.31252.48942.4894

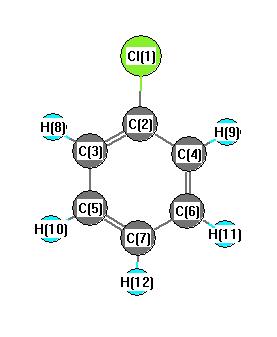
picture of chlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Cl1 C2 C3 119.440 Cl1 C2 C4 119.440
C2 C3 C5 119.145 C2 C3 H8 119.834
C2 C4 C6 119.145 C2 C4 H9 119.834
C3 C2 C4 121.121 C3 C5 C7 120.458
C3 C5 H10 119.343 C4 C6 C7 120.458
C4 C6 H11 119.343 C5 C3 H8 121.021
C5 C7 C6 119.673 C5 C7 H12 120.164
C6 C4 H9 121.021 C6 C7 H12 120.164
C7 C5 H10 120.199 C7 C6 H11 120.199
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability