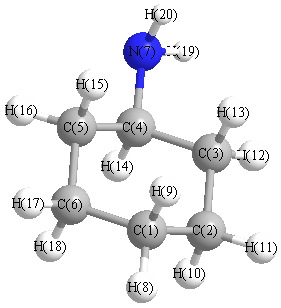
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3800 |
3437 |
0.41 |
|
|
|
| 2 |
A |
3728 |
3372 |
1.47 |
|
|
|
| 3 |
A |
3228 |
2920 |
77.61 |
|
|
|
| 4 |
A |
3211 |
2904 |
158.41 |
|
|
|
| 5 |
A |
3205 |
2898 |
96.02 |
|
|
|
| 6 |
A |
3202 |
2896 |
81.38 |
|
|
|
| 7 |
A |
3191 |
2886 |
94.06 |
|
|
|
| 8 |
A |
3167 |
2864 |
72.94 |
|
|
|
| 9 |
A |
3160 |
2858 |
23.16 |
|
|
|
| 10 |
A |
3157 |
2856 |
12.28 |
|
|
|
| 11 |
A |
3157 |
2855 |
8.97 |
|
|
|
| 12 |
A |
3147 |
2846 |
34.47 |
|
|
|
| 13 |
A |
3141 |
2841 |
13.31 |
|
|
|
| 14 |
A |
1851 |
1674 |
40.20 |
|
|
|
| 15 |
A |
1645 |
1488 |
1.63 |
|
|
|
| 16 |
A |
1628 |
1473 |
9.25 |
|
|
|
| 17 |
A |
1626 |
1471 |
4.02 |
|
|
|
| 18 |
A |
1622 |
1467 |
0.98 |
|
|
|
| 19 |
A |
1615 |
1461 |
0.10 |
|
|
|
| 20 |
A |
1567 |
1417 |
6.28 |
|
|
|
| 21 |
A |
1522 |
1377 |
4.97 |
|
|
|
| 22 |
A |
1519 |
1374 |
0.94 |
|
|
|
| 23 |
A |
1514 |
1369 |
1.78 |
|
|
|
| 24 |
A |
1500 |
1356 |
0.72 |
|
|
|
| 25 |
A |
1491 |
1349 |
2.68 |
|
|
|
| 26 |
A |
1457 |
1318 |
1.52 |
|
|
|
| 27 |
A |
1416 |
1281 |
1.12 |
|
|
|
| 28 |
A |
1398 |
1264 |
1.63 |
|
|
|
| 29 |
A |
1397 |
1263 |
2.55 |
|
|
|
| 30 |
A |
1334 |
1206 |
5.74 |
|
|
|
| 31 |
A |
1306 |
1181 |
2.31 |
|
|
|
| 32 |
A |
1224 |
1107 |
18.51 |
|
|
|
| 33 |
A |
1212 |
1096 |
6.99 |
|
|
|
| 34 |
A |
1166 |
1054 |
0.02 |
|
|
|
| 35 |
A |
1158 |
1048 |
2.36 |
|
|
|
| 36 |
A |
1128 |
1020 |
12.04 |
|
|
|
| 37 |
A |
1106 |
1001 |
1.85 |
|
|
|
| 38 |
A |
1061 |
960 |
2.04 |
|
|
|
| 39 |
A |
1011 |
914 |
62.61 |
|
|
|
| 40 |
A |
992 |
897 |
83.08 |
|
|
|
| 41 |
A |
953 |
861 |
2.54 |
|
|
|
| 42 |
A |
941 |
851 |
47.24 |
|
|
|
| 43 |
A |
903 |
817 |
4.46 |
|
|
|
| 44 |
A |
856 |
774 |
0.41 |
|
|
|
| 45 |
A |
836 |
756 |
2.24 |
|
|
|
| 46 |
A |
591 |
535 |
1.10 |
|
|
|
| 47 |
A |
491 |
444 |
1.19 |
|
|
|
| 48 |
A |
481 |
435 |
1.82 |
|
|
|
| 49 |
A |
434 |
393 |
0.19 |
|
|
|
| 50 |
A |
367 |
332 |
13.88 |
|
|
|
| 51 |
A |
354 |
320 |
0.95 |
|
|
|
| 52 |
A |
277 |
250 |
40.91 |
|
|
|
| 53 |
A |
240 |
217 |
5.94 |
|
|
|
| 54 |
A |
169 |
153 |
0.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 43925.7 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 39726.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.404 |
|
|
|
| 2 |
C |
-0.415 |
|
|
|
| 3 |
C |
-0.397 |
|
|
|
| 4 |
C |
-0.021 |
|
|
|
| 5 |
C |
-0.401 |
|
|
|
| 6 |
C |
-0.413 |
|
|
|
| 7 |
N |
-0.829 |
|
|
|
| 8 |
H |
0.211 |
|
|
|
| 9 |
H |
0.199 |
|
|
|
| 10 |
H |
0.201 |
|
|
|
| 11 |
H |
0.212 |
|
|
|
| 12 |
H |
0.228 |
|
|
|
| 13 |
H |
0.187 |
|
|
|
| 14 |
H |
0.198 |
|
|
|
| 15 |
H |
0.186 |
|
|
|
| 16 |
H |
0.199 |
|
|
|
| 17 |
H |
0.211 |
|
|
|
| 18 |
H |
0.201 |
|
|
|
| 19 |
H |
0.327 |
|
|
|
| 20 |
H |
0.322 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.531 |
1.170 |
0.462 |
1.366 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.547 |
-4.173 |
-1.560 |
| y |
-4.173 |
-47.023 |
-0.689 |
| z |
-1.560 |
-0.689 |
-44.718 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.676 |
-4.173 |
-1.560 |
| y |
-4.173 |
-0.391 |
-0.689 |
| z |
-1.560 |
-0.689 |
3.067 |
|
| Polar |
|---|
| 3z2-r2 | 6.134 |
|---|
| x2-y2 | -1.524 |
|---|
| xy | -4.173 |
|---|
| xz | -1.560 |
|---|
| yz | -0.689 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.903 |
-0.163 |
-0.097 |
| y |
-0.163 |
10.108 |
-0.101 |
| z |
-0.097 |
-0.101 |
9.265 |
<r2> (average value of r
2) Å
2
| <r2> |
232.424 |
| (<r2>)1/2 |
15.245 |