return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H13N (cyclohexanamine)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-290.420269
Energy at 298.15K-290.436780
HF Energy-289.282795
Nuclear repulsion energy330.840977
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3616 3425 0.12      
2 A 3519 3333 1.65      
3 A 3135 2969 47.82      
4 A 3128 2962 50.65      
5 A 3121 2956 29.53      
6 A 3118 2953 40.89      
7 A 3105 2941 50.41      
8 A 3067 2905 57.74      
9 A 3064 2902 16.31      
10 A 3063 2901 24.61      
11 A 3062 2900 17.59      
12 A 3047 2886 13.59      
13 A 3046 2884 22.84      
14 A 1733 1641 40.17      
15 A 1546 1464 2.44      
16 A 1533 1452 18.31      
17 A 1526 1445 7.18      
18 A 1524 1443 2.08      
19 A 1517 1437 1.13      
20 A 1455 1378 4.10      
21 A 1409 1334 1.94      
22 A 1406 1331 1.79      
23 A 1403 1328 4.87      
24 A 1391 1318 0.75      
25 A 1385 1311 0.31      
26 A 1354 1282 0.57      
27 A 1331 1260 1.40      
28 A 1312 1243 2.59      
29 A 1310 1241 2.00      
30 A 1252 1185 3.58      
31 A 1226 1161 2.12      
32 A 1165 1104 4.36      
33 A 1155 1094 7.34      
34 A 1120 1061 1.75      
35 A 1096 1038 0.53      
36 A 1078 1021 6.60      
37 A 1066 1010 0.76      
38 A 1004 951 3.45      
39 A 968 917 85.03      
40 A 936 886 77.62      
41 A 911 863 8.58      
42 A 898 850 32.21      
43 A 871 825 2.10      
44 A 805 763 2.45      
45 A 801 759 0.34      
46 A 561 531 1.53      
47 A 468 443 1.08      
48 A 451 428 2.56      
49 A 408 386 0.09      
50 A 348 329 13.74      
51 A 339 321 1.91      
52 A 258 244 36.08      
53 A 231 219 8.88      
54 A 164 155 0.37      

Unscaled Zero Point Vibrational Energy (zpe) 41901.1 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 39684.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.14292 0.07387 0.05371

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.873 0.010 0.290
C2 1.175 -1.250 -0.220
C3 -0.305 -1.260 0.163
C4 -1.027 -0.010 -0.328
C5 -0.317 1.248 0.181
C6 1.162 1.268 -0.207
N7 -2.443 -0.101 0.031
H8 2.923 0.017 -0.024
H9 1.874 0.005 1.388
H10 1.264 -1.294 -1.313
H11 1.671 -2.146 0.169
H12 -0.815 -2.142 -0.237
H13 -0.397 -1.309 1.258
H14 -0.983 -0.005 -1.427
H15 -0.412 1.276 1.276
H16 -0.830 2.139 -0.203
H17 1.648 2.166 0.190
H18 1.248 1.323 -1.300
H19 -2.935 0.715 -0.318
H20 -2.526 -0.081 1.043

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 N7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20
C11.52822.52432.96552.51811.52804.32531.09571.09792.15472.16883.48342.79813.33262.79203.47602.17002.15474.89784.4639
C21.52821.52822.52912.93662.51763.80402.16782.15611.09781.09552.18042.15812.76783.33703.93733.47272.79104.55654.0811
C32.52431.52821.52452.50762.94552.43563.47602.80122.15422.16501.09441.10012.13542.77173.45833.94333.34963.32422.6641
C42.96552.52911.52451.53152.53761.46363.96163.37062.80493.47652.14392.14451.09952.14652.16163.48702.81012.04142.0326
C52.51812.93662.50761.53151.52902.52213.47182.79323.34553.93283.45112.77522.14451.09961.09742.16882.15592.71822.7180
C61.52802.51762.94552.53761.52903.86322.16762.15542.79243.47173.94113.34872.77672.16212.17421.09561.09794.13584.1207
N74.32533.80402.43561.46362.52213.86325.36734.52634.11974.59592.62422.67392.06542.75202.76984.67964.17391.01481.0155
H81.09572.16783.47603.96163.47182.16765.36731.75932.47632.50674.32163.79774.15013.79404.31492.50792.47745.90695.5531
H91.09792.15612.80123.37062.79322.15544.52631.75933.05882.48033.80492.62664.01072.61773.79422.48163.05855.15204.4140
H102.15471.09782.15422.80493.34552.79244.11972.47633.05881.75732.48973.06092.59284.01484.17173.79192.61694.76034.6245
H112.16881.09552.16503.47653.93283.47174.59592.50672.48031.75732.51862.48213.76444.15594.97494.31203.79055.44394.7579
H123.48342.18041.09442.14393.45113.94112.62424.32163.80492.48972.51861.76142.45103.75954.28054.98034.16983.55842.9684
H132.79812.15811.10012.14452.77523.34872.67393.79772.62663.06092.48211.76143.04132.58523.76914.17094.02163.60862.4669
H143.33262.76782.13541.09952.14452.77672.06544.15014.01072.59283.76442.45103.04133.04542.47383.77492.59952.35792.9130
H152.79203.33702.77172.14651.09962.16212.75203.79402.61774.01484.15593.75952.58523.04541.76222.49253.06463.03742.5233
H163.47603.93733.45832.16161.09742.17422.76984.31493.79424.17174.97494.28053.76912.47381.76222.50902.48772.54413.0585
H172.17003.47273.94333.48702.16881.09564.67962.50792.48163.79194.31204.98034.17093.77492.49252.50901.75794.83414.8163
H182.15472.79103.34962.81012.15591.09794.17392.47743.05852.61693.79054.16984.02162.59953.06462.48771.75794.33984.6585
H194.89784.55653.32422.04142.71824.13581.01485.90695.15204.76035.44393.55843.60862.35793.03742.54414.83414.33981.6289
H204.46394.08112.66412.03262.71804.12071.01555.55314.41404.62454.75792.96842.46692.91302.52333.05854.81634.65851.6289

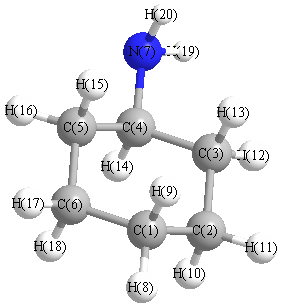
picture of cyclohexanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.362 C1 C2 H10 109.178
C1 C2 H11 110.412 C1 C6 C5 110.917
C1 C6 H17 110.527 C1 C6 H18 109.184
C2 C1 C6 110.928 C2 C1 H8 110.321
C2 C1 H9 109.274 C2 C3 C4 111.883
C2 C3 H12 111.417 C2 C3 H13 109.313
C3 C2 H10 109.137 C3 C2 H11 110.120
C3 C4 C5 110.274 C3 C4 N7 109.176
C3 C4 H14 107.836 C4 C3 H12 108.784
C4 C3 H13 108.503 C4 C5 C6 112.016
C4 C5 H15 108.212 C4 C5 H16 109.507
C4 N7 H19 109.578 C4 N7 H20 108.807
C5 C4 N7 114.700 C5 C4 H14 108.062
C5 C6 H17 110.360 C5 C6 H18 109.207
C6 C1 H8 110.325 C6 C1 H9 109.238
C6 C5 H15 109.595 C6 C5 H16 110.673
N7 C4 H14 106.514 H8 C1 H9 106.648
H10 C2 H11 106.497 H12 C3 H13 106.769
H15 C5 H16 106.662 H17 C6 H18 106.533
H19 N7 H20 106.699
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability