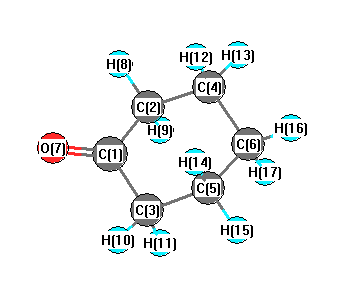
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3113 |
2997 |
20.37 |
|
|
|
| 2 |
A' |
3082 |
2967 |
57.75 |
|
|
|
| 3 |
A' |
3075 |
2960 |
51.27 |
|
|
|
| 4 |
A' |
3033 |
2920 |
49.50 |
|
|
|
| 5 |
A' |
3022 |
2910 |
11.37 |
|
|
|
| 6 |
A' |
3019 |
2907 |
19.00 |
|
|
|
| 7 |
A' |
1818 |
1750 |
214.80 |
|
|
|
| 8 |
A' |
1512 |
1456 |
2.14 |
|
|
|
| 9 |
A' |
1498 |
1442 |
12.63 |
|
|
|
| 10 |
A' |
1477 |
1422 |
10.27 |
|
|
|
| 11 |
A' |
1382 |
1331 |
2.95 |
|
|
|
| 12 |
A' |
1352 |
1301 |
4.20 |
|
|
|
| 13 |
A' |
1283 |
1235 |
0.23 |
|
|
|
| 14 |
A' |
1256 |
1209 |
1.40 |
|
|
|
| 15 |
A' |
1139 |
1097 |
0.60 |
|
|
|
| 16 |
A' |
1035 |
996 |
3.29 |
|
|
|
| 17 |
A' |
1006 |
968 |
1.28 |
|
|
|
| 18 |
A' |
875 |
842 |
4.59 |
|
|
|
| 19 |
A' |
852 |
821 |
0.42 |
|
|
|
| 20 |
A' |
756 |
728 |
3.63 |
|
|
|
| 21 |
A' |
660 |
635 |
1.75 |
|
|
|
| 22 |
A' |
477 |
459 |
0.27 |
|
|
|
| 23 |
A' |
403 |
388 |
1.17 |
|
|
|
| 24 |
A' |
305 |
293 |
0.13 |
|
|
|
| 25 |
A' |
91 |
87 |
2.75 |
|
|
|
| 26 |
A" |
3111 |
2995 |
24.67 |
|
|
|
| 27 |
A" |
3077 |
2962 |
40.47 |
|
|
|
| 28 |
A" |
3032 |
2918 |
21.53 |
|
|
|
| 29 |
A" |
3016 |
2904 |
3.65 |
|
|
|
| 30 |
A" |
1497 |
1441 |
7.56 |
|
|
|
| 31 |
A" |
1467 |
1413 |
9.99 |
|
|
|
| 32 |
A" |
1387 |
1336 |
1.58 |
|
|
|
| 33 |
A" |
1370 |
1319 |
5.80 |
|
|
|
| 34 |
A" |
1345 |
1295 |
17.79 |
|
|
|
| 35 |
A" |
1298 |
1249 |
0.78 |
|
|
|
| 36 |
A" |
1243 |
1197 |
23.86 |
|
|
|
| 37 |
A" |
1148 |
1105 |
21.85 |
|
|
|
| 38 |
A" |
1091 |
1050 |
1.56 |
|
|
|
| 39 |
A" |
1067 |
1027 |
7.59 |
|
|
|
| 40 |
A" |
923 |
888 |
8.60 |
|
|
|
| 41 |
A" |
905 |
871 |
0.48 |
|
|
|
| 42 |
A" |
760 |
731 |
4.76 |
|
|
|
| 43 |
A" |
493 |
474 |
9.62 |
|
|
|
| 44 |
A" |
412 |
397 |
2.57 |
|
|
|
| 45 |
A" |
176 |
170 |
0.73 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33167.5 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 31930.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.353 |
|
|
|
| 2 |
C |
-0.520 |
|
|
|
| 3 |
C |
-0.520 |
|
|
|
| 4 |
C |
-0.442 |
|
|
|
| 5 |
C |
-0.442 |
|
|
|
| 6 |
C |
-0.441 |
|
|
|
| 7 |
O |
-0.305 |
|
|
|
| 8 |
H |
0.247 |
|
|
|
| 9 |
H |
0.232 |
|
|
|
| 10 |
H |
0.247 |
|
|
|
| 11 |
H |
0.232 |
|
|
|
| 12 |
H |
0.227 |
|
|
|
| 13 |
H |
0.231 |
|
|
|
| 14 |
H |
0.227 |
|
|
|
| 15 |
H |
0.231 |
|
|
|
| 16 |
H |
0.230 |
|
|
|
| 17 |
H |
0.215 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.061 |
-2.354 |
0.000 |
3.129 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.210 |
4.739 |
0.000 |
| y |
4.739 |
-48.385 |
0.000 |
| z |
0.000 |
0.000 |
-41.576 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.230 |
4.739 |
0.000 |
| y |
4.739 |
-4.991 |
0.000 |
| z |
0.000 |
0.000 |
5.221 |
|
| Polar |
|---|
| 3z2-r2 | 10.443 |
|---|
| x2-y2 | 3.174 |
|---|
| xy | 4.739 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.939 |
-1.079 |
0.000 |
| y |
-1.079 |
10.133 |
0.000 |
| z |
0.000 |
0.000 |
10.056 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |