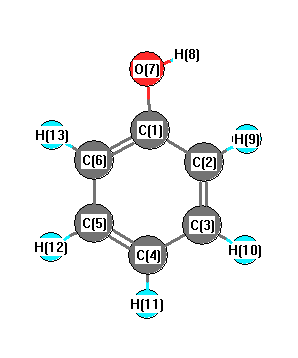
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3823 |
3680 |
38.40 |
|
|
|
| 2 |
A' |
3208 |
3088 |
7.14 |
|
|
|
| 3 |
A' |
3201 |
3081 |
25.40 |
|
|
|
| 4 |
A' |
3186 |
3067 |
23.00 |
|
|
|
| 5 |
A' |
3177 |
3059 |
0.06 |
|
|
|
| 6 |
A' |
3154 |
3037 |
18.28 |
|
|
|
| 7 |
A' |
1671 |
1609 |
45.01 |
|
|
|
| 8 |
A' |
1657 |
1595 |
41.84 |
|
|
|
| 9 |
A' |
1543 |
1485 |
54.62 |
|
|
|
| 10 |
A' |
1513 |
1456 |
37.56 |
|
|
|
| 11 |
A' |
1385 |
1334 |
42.72 |
|
|
|
| 12 |
A' |
1367 |
1316 |
8.55 |
|
|
|
| 13 |
A' |
1303 |
1254 |
76.82 |
|
|
|
| 14 |
A' |
1212 |
1167 |
146.49 |
|
|
|
| 15 |
A' |
1195 |
1150 |
7.73 |
|
|
|
| 16 |
A' |
1180 |
1136 |
7.48 |
|
|
|
| 17 |
A' |
1100 |
1059 |
13.41 |
|
|
|
| 18 |
A' |
1051 |
1012 |
5.74 |
|
|
|
| 19 |
A' |
1013 |
976 |
1.46 |
|
|
|
| 20 |
A' |
835 |
803 |
19.17 |
|
|
|
| 21 |
A' |
630 |
607 |
0.40 |
|
|
|
| 22 |
A' |
535 |
515 |
1.37 |
|
|
|
| 23 |
A' |
405 |
390 |
11.24 |
|
|
|
| 24 |
A" |
972 |
936 |
0.12 |
|
|
|
| 25 |
A" |
948 |
913 |
0.09 |
|
|
|
| 26 |
A" |
882 |
849 |
8.91 |
|
|
|
| 27 |
A" |
816 |
785 |
0.02 |
|
|
|
| 28 |
A" |
762 |
733 |
68.32 |
|
|
|
| 29 |
A" |
698 |
672 |
30.22 |
|
|
|
| 30 |
A" |
516 |
496 |
16.81 |
|
|
|
| 31 |
A" |
420 |
404 |
0.38 |
|
|
|
| 32 |
A" |
370 |
356 |
129.34 |
|
|
|
| 33 |
A" |
231 |
222 |
1.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22977.9 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 22120.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.277 |
|
|
|
| 2 |
C |
-0.278 |
|
|
|
| 3 |
C |
-0.204 |
|
|
|
| 4 |
C |
-0.212 |
|
|
|
| 5 |
C |
-0.201 |
|
|
|
| 6 |
C |
-0.231 |
|
|
|
| 7 |
O |
-0.572 |
|
|
|
| 8 |
H |
0.400 |
|
|
|
| 9 |
H |
0.200 |
|
|
|
| 10 |
H |
0.202 |
|
|
|
| 11 |
H |
0.200 |
|
|
|
| 12 |
H |
0.202 |
|
|
|
| 13 |
H |
0.217 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.431 |
0.078 |
0.000 |
1.434 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.876 |
-4.618 |
0.000 |
| y |
-4.618 |
-36.707 |
0.000 |
| z |
0.000 |
0.000 |
-44.946 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.951 |
-4.618 |
0.000 |
| y |
-4.618 |
3.204 |
0.000 |
| z |
0.000 |
0.000 |
-9.155 |
|
| Polar |
|---|
| 3z2-r2 | -18.310 |
|---|
| x2-y2 | 1.831 |
|---|
| xy | -4.618 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.264 |
-0.250 |
0.000 |
| y |
-0.250 |
12.385 |
-0.001 |
| z |
0.000 |
-0.001 |
4.326 |
<r2> (average value of r
2) Å
2
| <r2> |
185.504 |
| (<r2>)1/2 |
13.620 |