Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -195.952706 |
| Energy at 298.15K | -195.962934 |
| HF Energy | -195.175199 |
| Nuclear repulsion energy | 170.118989 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3277 |
3103 |
16.53 |
|
|
|
| 2 |
A |
3182 |
3014 |
4.69 |
|
|
|
| 3 |
A |
3170 |
3003 |
31.61 |
|
|
|
| 4 |
A |
3166 |
2999 |
31.61 |
|
|
|
| 5 |
A |
3164 |
2997 |
41.14 |
|
|
|
| 6 |
A |
3133 |
2967 |
17.28 |
|
|
|
| 7 |
A |
3111 |
2946 |
3.18 |
|
|
|
| 8 |
A |
3079 |
2917 |
40.63 |
|
|
|
| 9 |
A |
3075 |
2913 |
24.42 |
|
|
|
| 10 |
A |
3061 |
2899 |
14.84 |
|
|
|
| 11 |
A |
1708 |
1618 |
7.27 |
|
|
|
| 12 |
A |
1546 |
1465 |
11.04 |
|
|
|
| 13 |
A |
1541 |
1459 |
9.75 |
|
|
|
| 14 |
A |
1532 |
1451 |
1.30 |
|
|
|
| 15 |
A |
1523 |
1442 |
2.95 |
|
|
|
| 16 |
A |
1475 |
1397 |
2.33 |
|
|
|
| 17 |
A |
1448 |
1372 |
5.46 |
|
|
|
| 18 |
A |
1406 |
1332 |
0.42 |
|
|
|
| 19 |
A |
1359 |
1287 |
0.51 |
|
|
|
| 20 |
A |
1336 |
1265 |
0.36 |
|
|
|
| 21 |
A |
1313 |
1243 |
0.48 |
|
|
|
| 22 |
A |
1295 |
1227 |
0.11 |
|
|
|
| 23 |
A |
1221 |
1157 |
0.39 |
|
|
|
| 24 |
A |
1139 |
1079 |
1.64 |
|
|
|
| 25 |
A |
1092 |
1034 |
1.23 |
|
|
|
| 26 |
A |
1054 |
999 |
4.63 |
|
|
|
| 27 |
A |
1019 |
965 |
20.16 |
|
|
|
| 28 |
A |
977 |
926 |
1.22 |
|
|
|
| 29 |
A |
910 |
862 |
8.63 |
|
|
|
| 30 |
A |
904 |
856 |
40.27 |
|
|
|
| 31 |
A |
891 |
844 |
1.99 |
|
|
|
| 32 |
A |
751 |
712 |
2.80 |
|
|
|
| 33 |
A |
646 |
612 |
11.58 |
|
|
|
| 34 |
A |
444 |
421 |
1.08 |
|
|
|
| 35 |
A |
389 |
368 |
0.26 |
|
|
|
| 36 |
A |
255 |
241 |
0.03 |
|
|
|
| 37 |
A |
238 |
225 |
0.12 |
|
|
|
| 38 |
A |
104 |
98 |
0.01 |
|
|
|
| 39 |
A |
96 |
91 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30515.4 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 28901.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.455 |
-0.238 |
-0.315 |
| C2 |
1.409 |
0.356 |
0.274 |
| C3 |
0.068 |
-0.283 |
0.475 |
| C4 |
-1.038 |
0.430 |
-0.310 |
| C5 |
-2.410 |
-0.197 |
-0.082 |
| H6 |
3.399 |
0.277 |
-0.452 |
| H7 |
2.394 |
-1.262 |
-0.673 |
| H8 |
1.513 |
1.385 |
0.619 |
| H9 |
0.114 |
-1.335 |
0.170 |
| H10 |
-0.190 |
-0.272 |
1.542 |
| H11 |
-1.061 |
1.487 |
-0.021 |
| H12 |
-0.785 |
0.404 |
-1.375 |
| H13 |
-3.186 |
0.318 |
-0.653 |
| H14 |
-2.415 |
-1.248 |
-0.386 |
| H15 |
-2.689 |
-0.155 |
0.975 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3389 | 2.5139 | 3.5562 | 4.8708 | 1.0845 | 1.0862 | 2.0965 | 2.6303 | 3.2311 | 3.9271 | 3.4685 | 5.6784 | 4.9735 | 5.3034 |
C2 | 1.3389 | | 1.4989 | 2.5171 | 3.8758 | 2.1193 | 2.1173 | 1.0906 | 2.1329 | 2.1350 | 2.7327 | 2.7449 | 4.6881 | 4.1989 | 4.1890 | C3 | 2.5139 | 1.4989 | | 1.5320 | 2.5418 | 3.5021 | 2.7716 | 2.2116 | 1.0962 | 1.0979 | 2.1569 | 2.1493 | 3.4964 | 2.7995 | 2.8051 | C4 | 3.5562 | 2.5171 | 1.5320 | | 1.5257 | 4.4421 | 3.8433 | 2.8784 | 2.1615 | 2.1538 | 1.0960 | 1.0951 | 2.1781 | 2.1713 | 2.1717 | C5 | 4.8708 | 3.8758 | 2.5418 | 1.5257 | | 5.8404 | 4.9558 | 4.2883 | 2.7801 | 2.7518 | 2.1589 | 2.1622 | 1.0927 | 1.0934 | 1.0934 | H6 | 1.0845 | 2.1193 | 3.5021 | 4.4421 | 5.8404 | | 1.8518 | 2.4354 | 3.7121 | 4.1416 | 4.6411 | 4.2864 | 6.5884 | 6.0108 | 6.2676 | H7 | 1.0862 | 2.1173 | 2.7716 | 3.8433 | 4.9558 | 1.8518 | | 3.0746 | 2.4319 | 3.5437 | 4.4626 | 3.6565 | 5.7993 | 4.8170 | 5.4562 | H8 | 2.0965 | 1.0906 | 2.2116 | 2.8784 | 4.2883 | 2.4354 | 3.0746 | | 3.0920 | 2.5488 | 2.6544 | 3.1969 | 4.9840 | 4.8344 | 4.4896 | H9 | 2.6303 | 2.1329 | 1.0962 | 2.1615 | 2.7801 | 3.7121 | 2.4319 | 3.0920 | | 1.7614 | 3.0626 | 2.4938 | 3.7817 | 2.5905 | 3.1453 | H10 | 3.2311 | 2.1350 | 1.0979 | 2.1538 | 2.7518 | 4.1416 | 3.5437 | 2.5488 | 1.7614 | | 2.5086 | 3.0522 | 3.7607 | 3.1014 | 2.5653 | H11 | 3.9271 | 2.7327 | 2.1569 | 1.0960 | 2.1589 | 4.6411 | 4.4626 | 2.6544 | 3.0626 | 2.5086 | | 1.7556 | 2.5062 | 3.0729 | 2.5174 | H12 | 3.4685 | 2.7449 | 2.1493 | 1.0951 | 2.1622 | 4.2864 | 3.6565 | 3.1969 | 2.4938 | 3.0522 | 1.7556 | | 2.5090 | 2.5223 | 3.0753 | H13 | 5.6784 | 4.6881 | 3.4964 | 2.1781 | 1.0927 | 6.5884 | 5.7993 | 4.9840 | 3.7817 | 3.7607 | 2.5062 | 2.5090 | | 1.7660 | 1.7664 | H14 | 4.9735 | 4.1989 | 2.7995 | 2.1713 | 1.0934 | 6.0108 | 4.8170 | 4.8344 | 2.5905 | 3.1014 | 3.0729 | 2.5223 | 1.7660 | | 1.7661 | H15 | 5.3034 | 4.1890 | 2.8051 | 2.1717 | 1.0934 | 6.2676 | 5.4562 | 4.4896 | 3.1453 | 2.5653 | 2.5174 | 3.0753 | 1.7664 | 1.7661 | |
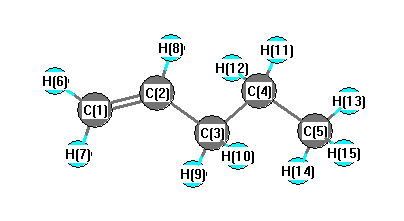 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.622 |
|
C1 |
C2 |
H8 |
118.935 |
| C2 |
C1 |
H6 |
121.621 |
|
C2 |
C1 |
H7 |
121.288 |
| C2 |
C3 |
C4 |
112.293 |
|
C2 |
C3 |
H9 |
109.571 |
| C2 |
C3 |
H10 |
109.639 |
|
C3 |
C2 |
H8 |
116.428 |
| C3 |
C4 |
C5 |
112.457 |
|
C3 |
C4 |
H11 |
109.193 |
| C3 |
C4 |
H12 |
108.648 |
|
C4 |
C3 |
H9 |
109.539 |
| C4 |
C3 |
H10 |
108.847 |
|
C4 |
C5 |
H13 |
111.509 |
| C4 |
C5 |
H14 |
110.917 |
|
C4 |
C5 |
H15 |
110.948 |
| C5 |
C4 |
H11 |
109.777 |
|
C5 |
C4 |
H12 |
110.089 |
| H6 |
C1 |
H7 |
117.091 |
|
H9 |
C3 |
H10 |
106.797 |
| H11 |
C4 |
H12 |
106.490 |
|
H13 |
C5 |
H14 |
107.768 |
| H13 |
C5 |
H15 |
107.807 |
|
H14 |
C5 |
H15 |
107.730 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -195.953102 |
| Energy at 298.15K | -195.963416 |
| HF Energy | -195.174183 |
| Nuclear repulsion energy | 173.884001 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3276 |
3103 |
15.59 |
|
|
|
| 2 |
A |
3181 |
3013 |
4.28 |
|
|
|
| 3 |
A |
3177 |
3009 |
30.02 |
|
|
|
| 4 |
A |
3166 |
2999 |
31.76 |
|
|
|
| 5 |
A |
3162 |
2995 |
28.20 |
|
|
|
| 6 |
A |
3127 |
2961 |
25.20 |
|
|
|
| 7 |
A |
3117 |
2952 |
19.35 |
|
|
|
| 8 |
A |
3078 |
2915 |
19.74 |
|
|
|
| 9 |
A |
3076 |
2913 |
33.04 |
|
|
|
| 10 |
A |
3065 |
2903 |
24.02 |
|
|
|
| 11 |
A |
1708 |
1617 |
5.79 |
|
|
|
| 12 |
A |
1546 |
1464 |
7.81 |
|
|
|
| 13 |
A |
1541 |
1459 |
13.14 |
|
|
|
| 14 |
A |
1531 |
1450 |
1.28 |
|
|
|
| 15 |
A |
1523 |
1442 |
6.19 |
|
|
|
| 16 |
A |
1476 |
1398 |
2.71 |
|
|
|
| 17 |
A |
1451 |
1374 |
9.01 |
|
|
|
| 18 |
A |
1398 |
1324 |
1.06 |
|
|
|
| 19 |
A |
1378 |
1305 |
0.32 |
|
|
|
| 20 |
A |
1336 |
1265 |
1.09 |
|
|
|
| 21 |
A |
1312 |
1242 |
0.63 |
|
|
|
| 22 |
A |
1285 |
1217 |
1.33 |
|
|
|
| 23 |
A |
1216 |
1152 |
0.37 |
|
|
|
| 24 |
A |
1123 |
1064 |
1.64 |
|
|
|
| 25 |
A |
1099 |
1041 |
4.80 |
|
|
|
| 26 |
A |
1037 |
982 |
3.78 |
|
|
|
| 27 |
A |
1019 |
965 |
18.55 |
|
|
|
| 28 |
A |
956 |
905 |
1.85 |
|
|
|
| 29 |
A |
904 |
857 |
46.42 |
|
|
|
| 30 |
A |
901 |
854 |
0.91 |
|
|
|
| 31 |
A |
886 |
840 |
1.26 |
|
|
|
| 32 |
A |
787 |
746 |
3.97 |
|
|
|
| 33 |
A |
647 |
613 |
10.48 |
|
|
|
| 34 |
A |
438 |
415 |
0.35 |
|
|
|
| 35 |
A |
404 |
382 |
2.06 |
|
|
|
| 36 |
A |
302 |
286 |
0.27 |
|
|
|
| 37 |
A |
237 |
224 |
0.05 |
|
|
|
| 38 |
A |
119 |
113 |
0.05 |
|
|
|
| 39 |
A |
91 |
86 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30535.5 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 28920.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.062 |
0.578 |
0.123 |
| C2 |
-1.169 |
-0.272 |
-0.400 |
| C3 |
-0.055 |
-0.922 |
0.364 |
| C4 |
1.329 |
-0.511 |
-0.155 |
| C5 |
1.604 |
0.980 |
0.017 |
| H6 |
-2.845 |
1.023 |
-0.482 |
| H7 |
-2.036 |
0.846 |
1.175 |
| H8 |
-1.231 |
-0.515 |
-1.462 |
| H9 |
-0.154 |
-2.013 |
0.290 |
| H10 |
-0.140 |
-0.666 |
1.426 |
| H11 |
2.092 |
-1.091 |
0.375 |
| H12 |
1.412 |
-0.784 |
-1.214 |
| H13 |
2.605 |
1.241 |
-0.334 |
| H14 |
0.882 |
1.582 |
-0.537 |
| H15 |
1.534 |
1.266 |
1.071 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3390 | 2.5175 | 3.5728 | 3.6896 | 1.0847 | 1.0861 | 2.0963 | 3.2218 | 2.6347 | 4.4843 | 3.9642 | 4.7362 | 3.1796 | 3.7820 |
C2 | 1.3390 | | 1.4991 | 2.5218 | 3.0711 | 2.1196 | 2.1172 | 1.0911 | 2.1298 | 2.1329 | 3.4508 | 2.7549 | 4.0667 | 2.7682 | 3.4402 | C3 | 2.5175 | 1.4991 | | 1.5345 | 2.5473 | 3.5049 | 2.7771 | 2.2097 | 1.0971 | 1.0964 | 2.1538 | 2.1586 | 3.4987 | 2.8213 | 2.7952 | C4 | 3.5728 | 2.5218 | 1.5345 | | 1.5251 | 4.4593 | 3.8648 | 2.8748 | 2.1573 | 2.1642 | 1.0955 | 1.0963 | 2.1741 | 2.1737 | 2.1684 | C5 | 3.6896 | 3.0711 | 2.5473 | 1.5251 | | 4.4774 | 3.8218 | 3.5299 | 3.4809 | 2.7808 | 2.1573 | 2.1596 | 1.0928 | 1.0917 | 1.0939 | H6 | 1.0847 | 2.1196 | 3.5049 | 4.4593 | 4.4774 | | 1.8520 | 2.4349 | 4.1295 | 3.7164 | 5.4389 | 4.6830 | 5.4568 | 3.7691 | 4.6525 | H7 | 1.0861 | 2.1172 | 2.7771 | 3.8648 | 3.8218 | 1.8520 | | 3.0744 | 3.5355 | 2.4386 | 4.6300 | 4.5008 | 4.8962 | 3.4617 | 3.5958 | H8 | 2.0963 | 1.0911 | 2.2097 | 2.8748 | 3.5299 | 2.4349 | 3.0744 | | 2.5446 | 3.0912 | 3.8410 | 2.6692 | 4.3671 | 3.1172 | 4.1511 | H9 | 3.2218 | 2.1298 | 1.0971 | 2.1573 | 3.4809 | 4.1295 | 3.5355 | 2.5446 | | 1.7618 | 2.4292 | 2.4946 | 4.3110 | 3.8310 | 3.7692 | H10 | 2.6347 | 2.1329 | 1.0964 | 2.1642 | 2.7808 | 3.7164 | 2.4386 | 3.0912 | 1.7618 | | 2.5037 | 3.0647 | 3.7774 | 3.1544 | 2.5807 | H11 | 4.4843 | 3.4508 | 2.1538 | 1.0955 | 2.1573 | 5.4389 | 4.6300 | 3.8410 | 2.4292 | 2.5037 | | 1.7548 | 2.4905 | 3.0728 | 2.5204 | H12 | 3.9642 | 2.7549 | 2.1586 | 1.0963 | 2.1596 | 4.6830 | 4.5008 | 2.6692 | 2.4946 | 3.0647 | 1.7548 | | 2.5093 | 2.5178 | 3.0722 | H13 | 4.7362 | 4.0667 | 3.4987 | 2.1741 | 1.0928 | 5.4568 | 4.8962 | 4.3671 | 4.3110 | 3.7774 | 2.4905 | 2.5093 | | 1.7685 | 1.7671 | H14 | 3.1796 | 2.7682 | 2.8213 | 2.1737 | 1.0917 | 3.7691 | 3.4617 | 3.1172 | 3.8310 | 3.1544 | 3.0728 | 2.5178 | 1.7685 | | 1.7635 | H15 | 3.7820 | 3.4402 | 2.7952 | 2.1684 | 1.0939 | 4.6525 | 3.5958 | 4.1511 | 3.7692 | 2.5807 | 2.5204 | 3.0722 | 1.7671 | 1.7635 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.909 |
|
C1 |
C2 |
H8 |
118.874 |
| C2 |
C1 |
H6 |
121.625 |
|
C2 |
C1 |
H7 |
121.267 |
| C2 |
C3 |
C4 |
112.454 |
|
C2 |
C3 |
H9 |
109.261 |
| C2 |
C3 |
H10 |
109.546 |
|
C3 |
C2 |
H8 |
116.212 |
| C3 |
C4 |
C5 |
112.722 |
|
C3 |
C4 |
H11 |
108.809 |
| C3 |
C4 |
H12 |
109.138 |
|
C4 |
C3 |
H9 |
108.984 |
| C4 |
C3 |
H10 |
109.566 |
|
C4 |
C5 |
H13 |
111.219 |
| C4 |
C5 |
H14 |
111.255 |
|
C4 |
C5 |
H15 |
110.697 |
| C5 |
C4 |
H11 |
109.723 |
|
C5 |
C4 |
H12 |
109.860 |
| H6 |
C1 |
H7 |
117.107 |
|
H9 |
C3 |
H10 |
106.870 |
| H11 |
C4 |
H12 |
106.384 |
|
H13 |
C5 |
H14 |
108.113 |
| H13 |
C5 |
H15 |
107.826 |
|
H14 |
C5 |
H15 |
107.578 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
