Jump to
S1C2
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -269.513700 |
| Energy at 298.15K | -269.523325 |
| Nuclear repulsion energy | 198.564426 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3143 |
3026 |
6.81 |
|
|
|
| 2 |
A |
3077 |
2962 |
79.81 |
|
|
|
| 3 |
A |
3016 |
2904 |
80.02 |
|
|
|
| 4 |
A |
3000 |
2888 |
10.69 |
|
|
|
| 5 |
A |
1530 |
1473 |
6.85 |
|
|
|
| 6 |
A |
1511 |
1455 |
0.91 |
|
|
|
| 7 |
A |
1498 |
1442 |
5.52 |
|
|
|
| 8 |
A |
1475 |
1420 |
0.08 |
|
|
|
| 9 |
A |
1353 |
1303 |
0.81 |
|
|
|
| 10 |
A |
1223 |
1178 |
35.58 |
|
|
|
| 11 |
A |
1185 |
1141 |
10.68 |
|
|
|
| 12 |
A |
1156 |
1113 |
76.01 |
|
|
|
| 13 |
A |
951 |
916 |
5.23 |
|
|
|
| 14 |
A |
609 |
586 |
6.68 |
|
|
|
| 15 |
A |
320 |
308 |
4.95 |
|
|
|
| 16 |
A |
155 |
149 |
0.28 |
|
|
|
| 17 |
A |
98 |
95 |
2.57 |
|
|
|
| 18 |
B |
3143 |
3026 |
53.04 |
|
|
|
| 19 |
B |
3076 |
2961 |
10.88 |
|
|
|
| 20 |
B |
3068 |
2954 |
54.98 |
|
|
|
| 21 |
B |
3001 |
2889 |
120.69 |
|
|
|
| 22 |
B |
1523 |
1466 |
9.21 |
|
|
|
| 23 |
B |
1499 |
1444 |
0.27 |
|
|
|
| 24 |
B |
1493 |
1438 |
16.64 |
|
|
|
| 25 |
B |
1439 |
1386 |
13.43 |
|
|
|
| 26 |
B |
1263 |
1216 |
16.31 |
|
|
|
| 27 |
B |
1183 |
1139 |
7.99 |
|
|
|
| 28 |
B |
1173 |
1129 |
183.51 |
|
|
|
| 29 |
B |
1085 |
1044 |
176.00 |
|
|
|
| 30 |
B |
966 |
930 |
74.79 |
|
|
|
| 31 |
B |
456 |
439 |
4.98 |
|
|
|
| 32 |
B |
215 |
207 |
17.02 |
|
|
|
| 33 |
B |
139 |
134 |
2.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25009.8 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 24076.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.915 |
| H2 |
-0.717 |
0.545 |
1.542 |
| H3 |
0.717 |
-0.545 |
1.542 |
| O4 |
0.773 |
0.881 |
0.155 |
| O5 |
-0.773 |
-0.881 |
0.155 |
| C6 |
0.000 |
1.832 |
-0.552 |
| C7 |
0.000 |
-1.832 |
-0.552 |
| H8 |
0.699 |
2.464 |
-1.100 |
| H9 |
-0.699 |
-2.464 |
-1.100 |
| H10 |
-0.687 |
1.352 |
-1.256 |
| H11 |
-0.583 |
2.458 |
0.139 |
| H12 |
0.687 |
-1.352 |
-1.256 |
| H13 |
0.583 |
-2.458 |
0.139 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
O4 |
O5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.0976 | 1.0976 | 1.3970 | 1.3970 | 2.3468 | 2.3468 | 3.2588 | 3.2588 | 2.6485 | 2.6428 | 2.6485 | 2.6428 |
H2 | 1.0976 | | 1.8020 | 2.0630 | 1.9906 | 2.5598 | 3.2480 | 3.5588 | 4.0045 | 2.9123 | 2.3759 | 3.6608 | 3.5608 | H3 | 1.0976 | 1.8020 | | 1.9906 | 2.0630 | 3.2480 | 2.5598 | 4.0045 | 3.5588 | 3.6608 | 3.5608 | 2.9123 | 2.3759 | O4 | 1.3970 | 2.0630 | 1.9906 | | 2.3443 | 1.4142 | 2.9084 | 2.0211 | 3.8641 | 2.0841 | 2.0793 | 2.6435 | 3.3448 | O5 | 1.3970 | 1.9906 | 2.0630 | 2.3443 | | 2.9084 | 1.4142 | 3.8641 | 2.0211 | 2.6435 | 3.3448 | 2.0841 | 2.0793 | C6 | 2.3468 | 2.5598 | 3.2480 | 1.4142 | 2.9084 | | 3.6640 | 1.0901 | 4.3868 | 1.0947 | 1.0991 | 3.3328 | 4.3841 | C7 | 2.3468 | 3.2480 | 2.5598 | 2.9084 | 1.4142 | 3.6640 | | 4.3868 | 1.0901 | 3.3328 | 4.3841 | 1.0947 | 1.0991 | H8 | 3.2588 | 3.5588 | 4.0045 | 2.0211 | 3.8641 | 1.0901 | 4.3868 | | 5.1222 | 1.7833 | 1.7824 | 3.8194 | 5.0767 | H9 | 3.2588 | 4.0045 | 3.5588 | 3.8641 | 2.0211 | 4.3868 | 1.0901 | 5.1222 | | 3.8194 | 5.0767 | 1.7833 | 1.7824 | H10 | 2.6485 | 2.9123 | 3.6608 | 2.0841 | 2.6435 | 1.0947 | 3.3328 | 1.7833 | 3.8194 | | 1.7830 | 3.0333 | 4.2515 | H11 | 2.6428 | 2.3759 | 3.5608 | 2.0793 | 3.3448 | 1.0991 | 4.3841 | 1.7824 | 5.0767 | 1.7830 | | 4.2515 | 5.0523 | H12 | 2.6485 | 3.6608 | 2.9123 | 2.6435 | 2.0841 | 3.3328 | 1.0947 | 3.8194 | 1.7833 | 3.0333 | 4.2515 | | 1.7830 | H13 | 2.6428 | 3.5608 | 2.3759 | 3.3448 | 2.0793 | 4.3841 | 1.0991 | 5.0767 | 1.7824 | 4.2515 | 5.0523 | 1.7830 | |
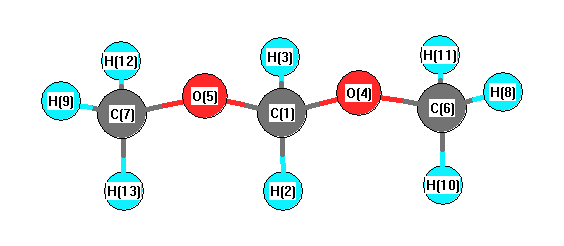 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O4 |
C6 |
113.187 |
|
C1 |
O5 |
C7 |
113.187 |
| H2 |
C1 |
H3 |
110.348 |
|
H2 |
C1 |
O4 |
111.014 |
| H2 |
C1 |
O5 |
105.239 |
|
H3 |
C1 |
O4 |
105.239 |
| H3 |
C1 |
O5 |
111.014 |
|
O4 |
C1 |
O5 |
114.076 |
| O4 |
C6 |
H8 |
106.902 |
|
O4 |
C6 |
H10 |
111.704 |
| O4 |
C6 |
H11 |
111.024 |
|
O5 |
C7 |
H9 |
106.902 |
| O5 |
C7 |
H12 |
111.704 |
|
O5 |
C7 |
H13 |
111.024 |
| H8 |
C6 |
H10 |
109.420 |
|
H8 |
C6 |
H11 |
109.006 |
| H9 |
C7 |
H12 |
109.420 |
|
H9 |
C7 |
H13 |
109.006 |
| H10 |
C6 |
H11 |
108.733 |
|
H12 |
C7 |
H13 |
108.733 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.078 |
|
|
|
| 2 |
H |
0.200 |
|
|
|
| 3 |
H |
0.200 |
|
|
|
| 4 |
O |
-0.330 |
|
|
|
| 5 |
O |
-0.330 |
|
|
|
| 6 |
C |
-0.475 |
|
|
|
| 7 |
C |
-0.475 |
|
|
|
| 8 |
H |
0.222 |
|
|
|
| 9 |
H |
0.222 |
|
|
|
| 10 |
H |
0.221 |
|
|
|
| 11 |
H |
0.201 |
|
|
|
| 12 |
H |
0.221 |
|
|
|
| 13 |
H |
0.201 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.234 |
0.234 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.598 |
-2.463 |
0.000 |
| y |
-2.463 |
-27.803 |
0.000 |
| z |
0.000 |
0.000 |
-29.432 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.980 |
-2.463 |
0.000 |
| y |
-2.463 |
4.212 |
0.000 |
| z |
0.000 |
0.000 |
1.769 |
|
| Polar |
|---|
| 3z2-r2 | 3.537 |
|---|
| x2-y2 | -6.795 |
|---|
| xy | -2.463 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.397 |
-0.083 |
0.000 |
| y |
-0.083 |
7.126 |
0.000 |
| z |
0.000 |
0.000 |
6.064 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
Jump to
S1C1
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -269.503704 |
| Energy at 298.15K | -269.513107 |
| Nuclear repulsion energy | 193.656762 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3142 |
3025 |
27.87 |
|
|
|
| 2 |
A1 |
2981 |
2870 |
73.33 |
|
|
|
| 3 |
A1 |
2882 |
2774 |
64.93 |
|
|
|
| 4 |
A1 |
1562 |
1503 |
0.55 |
|
|
|
| 5 |
A1 |
1520 |
1463 |
6.05 |
|
|
|
| 6 |
A1 |
1488 |
1432 |
0.18 |
|
|
|
| 7 |
A1 |
1282 |
1234 |
14.65 |
|
|
|
| 8 |
A1 |
1182 |
1138 |
9.39 |
|
|
|
| 9 |
A1 |
1061 |
1021 |
64.65 |
|
|
|
| 10 |
A1 |
483 |
465 |
2.04 |
|
|
|
| 11 |
A1 |
218 |
210 |
1.82 |
|
|
|
| 12 |
A2 |
3026 |
2913 |
0.00 |
|
|
|
| 13 |
A2 |
1500 |
1444 |
0.00 |
|
|
|
| 14 |
A2 |
1265 |
1218 |
0.00 |
|
|
|
| 15 |
A2 |
1185 |
1141 |
0.00 |
|
|
|
| 16 |
A2 |
223 |
214 |
0.00 |
|
|
|
| 17 |
A2 |
74 |
71 |
0.00 |
|
|
|
| 18 |
B1 |
3026 |
2914 |
155.59 |
|
|
|
| 19 |
B1 |
2898 |
2790 |
151.66 |
|
|
|
| 20 |
B1 |
1500 |
1444 |
18.52 |
|
|
|
| 21 |
B1 |
1204 |
1159 |
22.06 |
|
|
|
| 22 |
B1 |
1156 |
1113 |
8.94 |
|
|
|
| 23 |
B1 |
231 |
223 |
6.05 |
|
|
|
| 24 |
B1 |
79 |
77 |
0.69 |
|
|
|
| 25 |
B2 |
3142 |
3025 |
28.90 |
|
|
|
| 26 |
B2 |
2979 |
2867 |
74.35 |
|
|
|
| 27 |
B2 |
1532 |
1475 |
0.15 |
|
|
|
| 28 |
B2 |
1502 |
1446 |
35.74 |
|
|
|
| 29 |
B2 |
1456 |
1402 |
69.48 |
|
|
|
| 30 |
B2 |
1233 |
1187 |
211.47 |
|
|
|
| 31 |
B2 |
1177 |
1133 |
249.70 |
|
|
|
| 32 |
B2 |
1024 |
986 |
1.24 |
|
|
|
| 33 |
B2 |
426 |
410 |
3.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24819.6 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 23893.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.308 |
| H2 |
-0.899 |
0.000 |
0.960 |
| H3 |
0.899 |
0.000 |
0.960 |
| O4 |
0.000 |
1.111 |
-0.525 |
| O5 |
0.000 |
-1.111 |
-0.525 |
| C6 |
0.000 |
2.315 |
0.197 |
| C7 |
0.000 |
-2.315 |
0.197 |
| H8 |
0.000 |
3.124 |
-0.533 |
| H9 |
0.000 |
-3.124 |
-0.533 |
| H10 |
-0.893 |
2.412 |
0.833 |
| H11 |
0.893 |
2.412 |
0.833 |
| H12 |
0.893 |
-2.412 |
0.833 |
| H13 |
-0.893 |
-2.412 |
0.833 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
O4 |
O5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.1101 | 1.1101 | 1.3887 | 1.3887 | 2.3180 | 2.3180 | 3.2354 | 3.2354 | 2.6246 | 2.6246 | 2.6246 | 2.6246 |
H2 | 1.1101 | | 1.7971 | 2.0609 | 2.0609 | 2.5981 | 2.5981 | 3.5774 | 3.5774 | 2.4151 | 3.0068 | 3.0068 | 2.4151 | H3 | 1.1101 | 1.7971 | | 2.0609 | 2.0609 | 2.5981 | 2.5981 | 3.5774 | 3.5774 | 3.0068 | 2.4151 | 2.4151 | 3.0068 | O4 | 1.3887 | 2.0609 | 2.0609 | | 2.2214 | 1.4046 | 3.5013 | 2.0133 | 4.2347 | 2.0819 | 2.0819 | 3.8793 | 3.8793 | O5 | 1.3887 | 2.0609 | 2.0609 | 2.2214 | | 3.5013 | 1.4046 | 4.2347 | 2.0133 | 3.8793 | 3.8793 | 2.0819 | 2.0819 | C6 | 2.3180 | 2.5981 | 2.5981 | 1.4046 | 3.5013 | | 4.6306 | 1.0898 | 5.4881 | 1.1002 | 1.1002 | 4.8524 | 4.8524 | C7 | 2.3180 | 2.5981 | 2.5981 | 3.5013 | 1.4046 | 4.6306 | | 5.4881 | 1.0898 | 4.8524 | 4.8524 | 1.1002 | 1.1002 | H8 | 3.2354 | 3.5774 | 3.5774 | 2.0133 | 4.2347 | 1.0898 | 5.4881 | | 6.2480 | 1.7808 | 1.7808 | 5.7714 | 5.7714 | H9 | 3.2354 | 3.5774 | 3.5774 | 4.2347 | 2.0133 | 5.4881 | 1.0898 | 6.2480 | | 5.7714 | 5.7714 | 1.7808 | 1.7808 | H10 | 2.6246 | 2.4151 | 3.0068 | 2.0819 | 3.8793 | 1.1002 | 4.8524 | 1.7808 | 5.7714 | | 1.7852 | 5.1433 | 4.8235 | H11 | 2.6246 | 3.0068 | 2.4151 | 2.0819 | 3.8793 | 1.1002 | 4.8524 | 1.7808 | 5.7714 | 1.7852 | | 4.8235 | 5.1433 | H12 | 2.6246 | 3.0068 | 2.4151 | 3.8793 | 2.0819 | 4.8524 | 1.1002 | 5.7714 | 1.7808 | 5.1433 | 4.8235 | | 1.7852 | H13 | 2.6246 | 2.4151 | 3.0068 | 3.8793 | 2.0819 | 4.8524 | 1.1002 | 5.7714 | 1.7808 | 4.8235 | 5.1433 | 1.7852 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O4 |
C6 |
112.163 |
|
C1 |
O5 |
C7 |
112.163 |
| H2 |
C1 |
H3 |
108.087 |
|
H2 |
C1 |
O4 |
110.637 |
| H2 |
C1 |
O5 |
110.637 |
|
H3 |
C1 |
O4 |
110.637 |
| H3 |
C1 |
O5 |
110.637 |
|
O4 |
C1 |
O5 |
106.224 |
| O4 |
C6 |
H8 |
106.958 |
|
O4 |
C6 |
H10 |
111.865 |
| O4 |
C6 |
H11 |
111.865 |
|
O5 |
C7 |
H9 |
106.958 |
| O5 |
C7 |
H12 |
111.865 |
|
O5 |
C7 |
H13 |
111.865 |
| H8 |
C6 |
H10 |
108.806 |
|
H8 |
C6 |
H11 |
108.806 |
| H9 |
C7 |
H12 |
108.806 |
|
H9 |
C7 |
H13 |
108.806 |
| H10 |
C6 |
H11 |
108.454 |
|
H12 |
C7 |
H13 |
108.454 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.023 |
|
|
|
| 2 |
H |
0.162 |
|
|
|
| 3 |
H |
0.162 |
|
|
|
| 4 |
O |
-0.310 |
|
|
|
| 5 |
O |
-0.310 |
|
|
|
| 6 |
C |
-0.473 |
|
|
|
| 7 |
C |
-0.473 |
|
|
|
| 8 |
H |
0.231 |
|
|
|
| 9 |
H |
0.231 |
|
|
|
| 10 |
H |
0.201 |
|
|
|
| 11 |
H |
0.201 |
|
|
|
| 12 |
H |
0.201 |
|
|
|
| 13 |
H |
0.201 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.437 |
2.437 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.839 |
0.000 |
0.000 |
| y |
0.000 |
-24.776 |
0.000 |
| z |
0.000 |
0.000 |
-33.430 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.736 |
0.000 |
0.000 |
| y |
0.000 |
7.858 |
0.000 |
| z |
0.000 |
0.000 |
-5.122 |
|
| Polar |
|---|
| 3z2-r2 | -10.244 |
|---|
| x2-y2 | -7.063 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.635 |
0.000 |
0.000 |
| y |
0.000 |
7.979 |
0.000 |
| z |
0.000 |
0.000 |
5.568 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
