return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH(C2H5)2 (diethylamine)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-213.177470
Energy at 298.15K-213.190244
HF Energy-212.354039
Nuclear repulsion energy188.742865
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3534 3347 0.42      
2 A' 3186 3018 39.31      
3 A' 3165 2997 62.85      
4 A' 3112 2948 44.80      
5 A' 3080 2917 12.41      
6 A' 2991 2833 163.77      
7 A' 1566 1483 3.46      
8 A' 1540 1459 4.07      
9 A' 1530 1449 18.75      
10 A' 1458 1381 5.40      
11 A' 1425 1350 2.00      
12 A' 1354 1282 4.76      
13 A' 1266 1199 1.58      
14 A' 1190 1127 13.22      
15 A' 1094 1036 7.64      
16 A' 927 878 11.15      
17 A' 845 800 7.35      
18 A' 804 762 67.47      
19 A' 434 411 0.27      
20 A' 268 254 1.08      
21 A' 191 181 0.76      
22 A' 115 109 1.56      
23 A" 3186 3018 7.87      
24 A" 3165 2997 14.43      
25 A" 3111 2946 0.63      
26 A" 3080 2917 31.29      
27 A" 2989 2831 9.54      
28 A" 1555 1473 1.73      
29 A" 1544 1463 3.22      
30 A" 1534 1453 11.24      
31 A" 1510 1431 23.09      
32 A" 1443 1367 19.75      
33 A" 1384 1311 18.03      
34 A" 1313 1243 4.75      
35 A" 1192 1129 46.96      
36 A" 1137 1077 6.66      
37 A" 1092 1034 5.11      
38 A" 967 916 0.11      
39 A" 820 777 0.44      
40 A" 436 413 0.73      
41 A" 270 255 0.85      
42 A" 103 98 1.42      

Unscaled Zero Point Vibrational Energy (zpe) 33452.3 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 31682.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.59373 0.07042 0.06640

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.017 -0.293 0.000
C2 0.017 0.518 1.213
C3 0.017 0.518 -1.213
C4 0.017 -0.368 2.449
C5 0.017 -0.368 -2.449
H6 -0.820 -0.869 0.000
H7 -0.835 1.219 1.251
H8 0.926 1.129 1.198
H9 -0.835 1.219 -1.251
H10 0.926 1.129 -1.198
H11 0.081 0.230 3.361
H12 -0.898 -0.963 2.505
H13 0.866 -1.053 2.415
H14 0.081 0.230 -3.361
H15 -0.898 -0.963 -2.505
H16 0.866 -1.053 -2.415

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.45871.45872.45002.45001.01652.14002.06942.14002.06943.40222.75002.67003.40222.75002.6700
C21.45872.42521.52063.76702.02341.10451.09492.69992.64772.16862.16842.15204.58324.10514.0429
C31.45872.42523.76701.52062.02342.69992.64771.10451.09494.58324.10514.04292.16862.16842.1520
C42.45001.52063.76704.89772.63622.16322.15154.11544.04541.09261.09381.09105.84105.07284.9845
C52.45003.76701.52064.89772.63624.11544.04542.16322.15155.84105.07284.98451.09261.09381.0910
H61.01652.02342.02342.63622.63622.43442.91112.43442.91113.64932.50792.95133.64932.50792.9513
H72.14001.10452.69992.16324.11542.43441.76352.50263.01782.50382.51803.06734.80544.34494.6364
H82.06941.09492.64772.15154.04542.91111.76353.01782.39642.48993.06792.49844.72324.62794.2209
H92.14002.69991.10454.11542.16322.43442.50263.01781.76354.80544.34494.63642.50382.51803.0673
H102.06942.64771.09494.04542.15152.91113.01782.39641.76354.72324.62794.22092.48993.06792.4984
H113.40222.16864.58321.09265.84103.64932.50382.48994.80544.72321.76541.77666.72236.06585.9686
H122.75002.16844.10511.09385.07282.50792.51803.06794.34494.62791.76541.76926.06585.00995.2276
H132.67002.15204.04291.09104.98452.95133.06732.49844.63644.22091.77661.76925.96865.22764.8299
H143.40224.58322.16865.84101.09263.64934.80544.72322.50382.48996.72236.06585.96861.76541.7766
H152.75004.10512.16845.07281.09382.50794.34494.62792.51803.06796.06585.00995.22761.76541.7692
H162.67004.04292.15204.98451.09102.95134.63644.22093.06732.49845.96865.22764.82991.77661.7692

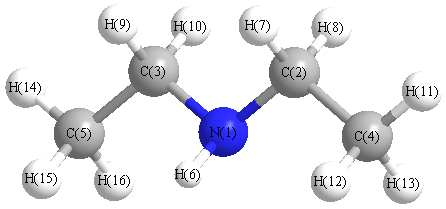
picture of diethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C4 110.622 N1 C2 H7 112.475
N1 C2 H8 107.413 N1 C3 C5 110.622
N1 C3 H9 112.475 N1 C3 H10 107.413
C2 N1 C3 112.469 C2 N1 H6 108.345
C2 C4 H11 111.114 C2 C4 H12 111.019
C2 C4 H13 109.886 C3 N1 H6 108.345
C3 C5 H14 111.114 C3 C5 H15 111.019
C3 C5 H16 109.886 C4 C2 H7 109.972
C4 C2 H8 109.617 C5 C3 H8 151.000
C5 C3 H10 109.617 H7 C2 H8 106.602
H9 C3 H10 106.602 H11 C4 H12 107.691
H11 C4 H13 108.897 H12 C4 H13 108.143
H14 C5 H15 107.691 H14 C5 H16 108.897
H15 C5 H16 108.143
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability