Jump to
S1C2
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -231.156568 |
| Energy at 298.15K | -231.162321 |
| Nuclear repulsion energy | 159.090079 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3269 |
3148 |
9.39 |
|
|
|
| 2 |
A1 |
3176 |
3058 |
1.80 |
|
|
|
| 3 |
A1 |
3154 |
3036 |
33.89 |
|
|
|
| 4 |
A1 |
1752 |
1686 |
4.91 |
|
|
|
| 5 |
A1 |
1438 |
1384 |
0.50 |
|
|
|
| 6 |
A1 |
1362 |
1311 |
6.00 |
|
|
|
| 7 |
A1 |
1219 |
1174 |
21.34 |
|
|
|
| 8 |
A1 |
870 |
838 |
2.34 |
|
|
|
| 9 |
A1 |
524 |
504 |
0.26 |
|
|
|
| 10 |
A1 |
231 |
222 |
0.00 |
|
|
|
| 11 |
A2 |
961 |
925 |
0.00 |
|
|
|
| 12 |
A2 |
840 |
809 |
0.00 |
|
|
|
| 13 |
A2 |
716 |
690 |
0.00 |
|
|
|
| 14 |
A2 |
112 |
107 |
0.00 |
|
|
|
| 15 |
B1 |
974 |
938 |
54.28 |
|
|
|
| 16 |
B1 |
845 |
813 |
126.05 |
|
|
|
| 17 |
B1 |
688 |
662 |
10.85 |
|
|
|
| 18 |
B1 |
84 |
81 |
1.71 |
|
|
|
| 19 |
B2 |
3269 |
3148 |
2.49 |
|
|
|
| 20 |
B2 |
3176 |
3057 |
0.05 |
|
|
|
| 21 |
B2 |
3147 |
3030 |
4.42 |
|
|
|
| 22 |
B2 |
1702 |
1638 |
471.84 |
|
|
|
| 23 |
B2 |
1427 |
1374 |
59.64 |
|
|
|
| 24 |
B2 |
1336 |
1286 |
18.24 |
|
|
|
| 25 |
B2 |
1233 |
1187 |
455.53 |
|
|
|
| 26 |
B2 |
1037 |
999 |
4.09 |
|
|
|
| 27 |
B2 |
488 |
470 |
7.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19514.9 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 18787.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.327 |
| C2 |
0.000 |
1.169 |
-0.370 |
| C3 |
0.000 |
-1.169 |
-0.370 |
| C4 |
0.000 |
2.350 |
0.235 |
| C5 |
0.000 |
-2.350 |
0.235 |
| H6 |
0.000 |
1.061 |
-1.454 |
| H7 |
0.000 |
-1.061 |
-1.454 |
| H8 |
0.000 |
3.257 |
-0.355 |
| H9 |
0.000 |
2.431 |
1.316 |
| H10 |
0.000 |
-3.257 |
-0.355 |
| H11 |
0.000 |
-2.431 |
1.316 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3613 | 1.3613 | 2.3514 | 2.3514 | 2.0731 | 2.0731 | 3.3275 | 2.6247 | 3.3275 | 2.6247 |
C2 | 1.3613 | | 2.3382 | 1.3267 | 3.5704 | 1.0890 | 2.4795 | 2.0878 | 2.1063 | 4.4259 | 3.9757 | C3 | 1.3613 | 2.3382 | | 3.5704 | 1.3267 | 2.4795 | 1.0890 | 4.4259 | 3.9757 | 2.0878 | 2.1063 | C4 | 2.3514 | 1.3267 | 3.5704 | | 4.6993 | 2.1243 | 3.8060 | 1.0822 | 1.0839 | 5.6374 | 4.9016 | C5 | 2.3514 | 3.5704 | 1.3267 | 4.6993 | | 3.8060 | 2.1243 | 5.6374 | 4.9016 | 1.0822 | 1.0839 | H6 | 2.0731 | 1.0890 | 2.4795 | 2.1243 | 3.8060 | | 2.1223 | 2.4553 | 3.0901 | 4.4556 | 4.4574 | H7 | 2.0731 | 2.4795 | 1.0890 | 3.8060 | 2.1243 | 2.1223 | | 4.4556 | 4.4574 | 2.4553 | 3.0901 | H8 | 3.3275 | 2.0878 | 4.4259 | 1.0822 | 5.6374 | 2.4553 | 4.4556 | | 1.8637 | 6.5136 | 5.9285 | H9 | 2.6247 | 2.1063 | 3.9757 | 1.0839 | 4.9016 | 3.0901 | 4.4574 | 1.8637 | | 5.9285 | 4.8627 | H10 | 3.3275 | 4.4259 | 2.0878 | 5.6374 | 1.0822 | 4.4556 | 2.4553 | 6.5136 | 5.9285 | | 1.8637 | H11 | 2.6247 | 3.9757 | 2.1063 | 4.9016 | 1.0839 | 4.4574 | 3.0901 | 5.9285 | 4.8627 | 1.8637 | |
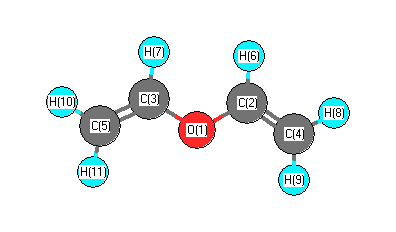 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
122.037 |
|
O1 |
C2 |
H6 |
115.126 |
| O1 |
C3 |
C5 |
122.037 |
|
O1 |
C3 |
H7 |
115.126 |
| C2 |
O1 |
C3 |
118.369 |
|
C2 |
C4 |
H8 |
119.808 |
| C2 |
C4 |
H9 |
121.473 |
|
C3 |
C5 |
H10 |
119.808 |
| C3 |
C5 |
H11 |
121.473 |
|
C4 |
C2 |
H6 |
122.837 |
| C5 |
C3 |
H7 |
122.837 |
|
H8 |
C4 |
H9 |
118.719 |
| H10 |
C5 |
H11 |
118.719 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.269 |
|
|
|
| 2 |
C |
0.001 |
|
|
|
| 3 |
C |
0.001 |
|
|
|
| 4 |
C |
-0.523 |
|
|
|
| 5 |
C |
-0.523 |
|
|
|
| 6 |
H |
0.202 |
|
|
|
| 7 |
H |
0.202 |
|
|
|
| 8 |
H |
0.224 |
|
|
|
| 9 |
H |
0.231 |
|
|
|
| 10 |
H |
0.224 |
|
|
|
| 11 |
H |
0.231 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.368 |
1.368 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.123 |
0.000 |
0.000 |
| y |
0.000 |
-27.354 |
0.000 |
| z |
0.000 |
0.000 |
-27.306 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.793 |
0.000 |
0.000 |
| y |
0.000 |
3.360 |
0.000 |
| z |
0.000 |
0.000 |
3.433 |
|
| Polar |
|---|
| 3z2-r2 | 6.866 |
|---|
| x2-y2 | -6.768 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
147.529 |
| (<r2>)1/2 |
12.146 |
Jump to
S1C1
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -231.156724 |
| Energy at 298.15K | -231.162776 |
| HF Energy | -231.156724 |
| Nuclear repulsion energy | 161.693039 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3265 |
3144 |
7.31 |
|
|
|
| 2 |
A |
3262 |
3141 |
10.29 |
|
|
|
| 3 |
A |
3208 |
3089 |
12.25 |
|
|
|
| 4 |
A |
3175 |
3056 |
4.43 |
|
|
|
| 5 |
A |
3172 |
3053 |
1.19 |
|
|
|
| 6 |
A |
3156 |
3039 |
10.07 |
|
|
|
| 7 |
A |
1723 |
1659 |
92.04 |
|
|
|
| 8 |
A |
1701 |
1638 |
197.66 |
|
|
|
| 9 |
A |
1444 |
1390 |
1.70 |
|
|
|
| 10 |
A |
1424 |
1371 |
18.14 |
|
|
|
| 11 |
A |
1370 |
1319 |
69.37 |
|
|
|
| 12 |
A |
1332 |
1282 |
5.28 |
|
|
|
| 13 |
A |
1253 |
1206 |
290.78 |
|
|
|
| 14 |
A |
1148 |
1105 |
16.70 |
|
|
|
| 15 |
A |
1020 |
982 |
35.11 |
|
|
|
| 16 |
A |
996 |
959 |
8.83 |
|
|
|
| 17 |
A |
984 |
947 |
42.46 |
|
|
|
| 18 |
A |
887 |
854 |
22.19 |
|
|
|
| 19 |
A |
866 |
834 |
52.06 |
|
|
|
| 20 |
A |
849 |
817 |
65.06 |
|
|
|
| 21 |
A |
736 |
709 |
7.52 |
|
|
|
| 22 |
A |
722 |
695 |
3.28 |
|
|
|
| 23 |
A |
591 |
569 |
2.42 |
|
|
|
| 24 |
A |
456 |
439 |
3.07 |
|
|
|
| 25 |
A |
281 |
271 |
1.52 |
|
|
|
| 26 |
A |
210 |
203 |
9.46 |
|
|
|
| 27 |
A |
96 |
92 |
0.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19663.0 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 18929.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.044 |
-0.791 |
0.046 |
| C2 |
1.283 |
-0.491 |
0.006 |
| C3 |
-0.930 |
0.218 |
0.305 |
| C4 |
1.838 |
0.708 |
-0.149 |
| C5 |
-2.156 |
0.218 |
-0.198 |
| H6 |
1.866 |
-1.401 |
0.105 |
| H7 |
-0.561 |
0.974 |
0.996 |
| H8 |
2.918 |
0.792 |
-0.152 |
| H9 |
1.263 |
1.614 |
-0.300 |
| H10 |
-2.860 |
0.988 |
0.092 |
| H11 |
-2.484 |
-0.551 |
-0.888 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3617 | 1.3671 | 2.4142 | 2.3531 | 2.0061 | 2.0697 | 3.3643 | 2.7593 | 3.3310 | 2.6228 |
C2 | 1.3617 | | 2.3429 | 1.3300 | 3.5176 | 1.0849 | 2.5546 | 2.0837 | 2.1271 | 4.4003 | 3.8719 | C3 | 1.3671 | 2.3429 | | 2.8472 | 1.3257 | 3.2365 | 1.0883 | 3.9166 | 2.6690 | 2.0893 | 2.1048 | C4 | 2.4142 | 1.3300 | 2.8472 | | 4.0245 | 2.1239 | 2.6713 | 1.0829 | 1.0838 | 4.7127 | 4.5617 | C5 | 2.3531 | 3.5176 | 1.3257 | 4.0245 | | 4.3462 | 2.1311 | 5.1064 | 3.6948 | 1.0828 | 1.0841 | H6 | 2.0061 | 1.0849 | 3.2365 | 2.1239 | 4.3462 | | 3.5098 | 2.4454 | 3.1009 | 5.2954 | 4.5417 | H7 | 2.0697 | 2.5546 | 1.0883 | 2.6713 | 2.1311 | 3.5098 | | 3.6674 | 2.3272 | 2.4707 | 3.0939 | H8 | 3.3643 | 2.0837 | 3.9166 | 1.0829 | 5.1064 | 2.4454 | 3.6674 | | 1.8534 | 5.7863 | 5.6144 | H9 | 2.7593 | 2.1271 | 2.6690 | 1.0838 | 3.6948 | 3.1009 | 2.3272 | 1.8534 | | 4.1890 | 4.3674 | H10 | 3.3310 | 4.4003 | 2.0893 | 4.7127 | 1.0828 | 5.2954 | 2.4707 | 5.7863 | 4.1890 | | 1.8633 | H11 | 2.6228 | 3.8719 | 2.1048 | 4.5617 | 1.0841 | 4.5417 | 3.0939 | 5.6144 | 4.3674 | 1.8633 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
127.504 |
|
O1 |
C2 |
H6 |
109.641 |
| O1 |
C3 |
C5 |
121.810 |
|
O1 |
C3 |
H7 |
114.425 |
| C2 |
O1 |
C3 |
118.309 |
|
C2 |
C4 |
H8 |
119.094 |
| C2 |
C4 |
H9 |
123.271 |
|
C3 |
C5 |
H10 |
119.994 |
| C3 |
C5 |
H11 |
121.397 |
|
C4 |
C2 |
H6 |
122.849 |
| C5 |
C3 |
H7 |
123.670 |
|
H8 |
C4 |
H9 |
117.616 |
| H10 |
C5 |
H11 |
118.601 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.277 |
|
|
|
| 2 |
C |
0.004 |
|
|
|
| 3 |
C |
-0.027 |
|
|
|
| 4 |
C |
-0.554 |
|
|
|
| 5 |
C |
-0.502 |
|
|
|
| 6 |
H |
0.217 |
|
|
|
| 7 |
H |
0.222 |
|
|
|
| 8 |
H |
0.226 |
|
|
|
| 9 |
H |
0.235 |
|
|
|
| 10 |
H |
0.224 |
|
|
|
| 11 |
H |
0.231 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.341 |
0.673 |
0.286 |
0.807 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.949 |
-1.803 |
0.574 |
| y |
-1.803 |
-29.355 |
1.061 |
| z |
0.574 |
1.061 |
-32.526 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.991 |
-1.803 |
0.574 |
| y |
-1.803 |
0.382 |
1.061 |
| z |
0.574 |
1.061 |
-4.373 |
|
| Polar |
|---|
| 3z2-r2 | -8.747 |
|---|
| x2-y2 | 2.406 |
|---|
| xy | -1.803 |
|---|
| xz | 0.574 |
|---|
| yz | 1.061 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.669 |
0.428 |
0.571 |
| y |
0.428 |
6.541 |
0.185 |
| z |
0.571 |
0.185 |
4.349 |
<r2> (average value of r
2) Å
2
| <r2> |
130.302 |
| (<r2>)1/2 |
11.415 |
