return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HCOOC2H5 (Ethyl formate)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS trans 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-267.776646
Energy at 298.15K-267.783930
HF Energy-266.888277
Nuclear repulsion energy177.870053
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3186 3017 18.01      
2 A' 3113 2949 5.76      
3 A' 3109 2945 74.15      
4 A' 3095 2931 13.52      
5 A' 1807 1711 267.99      
6 A' 1561 1478 9.69      
7 A' 1539 1458 3.70      
8 A' 1464 1386 24.31      
9 A' 1426 1351 1.13      
10 A' 1421 1346 1.44      
11 A' 1240 1174 418.28      
12 A' 1163 1102 9.52      
13 A' 1069 1012 16.01      
14 A' 886 839 11.25      
15 A' 801 758 2.93      
16 A' 391 370 6.08      
17 A' 237 225 6.49      
18 A" 3198 3029 29.40      
19 A" 3166 2999 8.09      
20 A" 1527 1446 8.86      
21 A" 1335 1264 1.17      
22 A" 1213 1149 4.74      
23 A" 1047 992 0.00      
24 A" 829 785 0.61      
25 A" 353 334 21.23      
26 A" 251 237 2.80      
27 A" 79 75 0.41      

Unscaled Zero Point Vibrational Energy (zpe) 20251.9 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 19180.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.59286 0.09735 0.08632

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.172 -0.255 0.000
C2 -0.692 -0.559 0.000
O3 0.000 0.708 0.000
C4 1.337 0.613 0.000
O5 1.975 -0.410 0.000
H6 -2.743 -1.187 0.000
H7 -2.447 0.319 0.886
H8 -2.447 0.319 -0.886
H9 -0.387 -1.126 -0.883
H10 -0.387 -1.126 0.883
H11 1.768 1.623 0.000

Atom - Atom Distances (Å)
  C1 C2 O3 C4 O5 H6 H7 H8 H9 H10 H11
C11.51062.37593.61494.15001.09241.09091.09092.17312.17314.3645
C21.51061.44382.34362.67152.14472.15282.15281.09231.09233.2884
O32.37591.44381.34062.26973.33362.63112.63112.07142.07141.9906
C43.61492.34361.34061.20564.45953.89753.89752.60272.60271.0980
O54.15002.67152.26971.20564.78174.56854.56852.62122.62122.0437
H61.09242.14473.33364.45954.78171.77171.77172.51682.51685.3142
H71.09092.15282.63113.89754.56851.77171.77133.07532.51614.4997
H81.09092.15282.63113.89754.56851.77171.77132.51613.07534.4997
H92.17311.09232.07142.60272.62122.51683.07532.51611.76523.6023
H102.17311.09232.07142.60272.62122.51682.51613.07531.76523.6023
H114.36453.28841.99061.09802.04375.31424.49974.49973.60233.6023

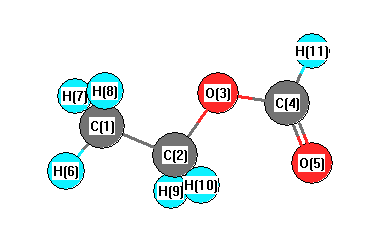
picture of Ethyl formate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 107.042 C1 C2 H9 112.213
C1 C2 H10 112.213 C2 C1 H6 109.924
C2 C1 H7 110.663 C2 C1 H8 110.663
C2 O3 C4 114.581 O3 C2 H9 108.732
O3 C2 H10 108.732 O3 C4 O5 126.012
O3 C4 H11 109.019 O5 C4 H11 124.969
H6 C1 H7 108.487 H6 C1 H8 108.487
H7 C1 H8 108.554 H9 C2 H10 107.814
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability