return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H9N (Pentanenitrile)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-250.038397
Energy at 298.15K-250.047907
HF Energy-249.075938
Nuclear repulsion energy210.923112
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3171 3004 27.43      
2 A' 3098 2935 23.27      
3 A' 3088 2924 13.75      
4 A' 3080 2917 20.62      
5 A' 3073 2910 15.94      
6 A' 2208 2091 0.02      
7 A' 1548 1466 13.13      
8 A' 1535 1454 2.23      
9 A' 1528 1447 0.95      
10 A' 1515 1435 3.50      
11 A' 1452 1375 6.29      
12 A' 1430 1354 0.59      
13 A' 1379 1306 1.71      
14 A' 1294 1225 0.33      
15 A' 1150 1089 4.94      
16 A' 1104 1046 0.13      
17 A' 1058 1002 0.94      
18 A' 956 906 0.04      
19 A' 927 878 3.50      
20 A' 541 512 0.89      
21 A' 392 372 0.62      
22 A' 282 268 0.79      
23 A' 125 118 5.32      
24 A" 3169 3002 49.34      
25 A" 3157 2990 11.08      
26 A" 3138 2972 4.45      
27 A" 3117 2952 4.81      
28 A" 1542 1460 11.02      
29 A" 1365 1293 0.12      
30 A" 1348 1277 0.74      
31 A" 1271 1204 0.05      
32 A" 1165 1103 0.02      
33 A" 955 904 0.66      
34 A" 807 764 0.06      
35 A" 736 697 4.75      
36 A" 376 356 0.16      
37 A" 252 238 0.14      
38 A" 115 109 0.03      
39 A" 89 84 3.70      

Unscaled Zero Point Vibrational Energy (zpe) 29267.0 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 27718.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.49996 0.04448 0.04212

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.402 0.000
C2 1.414 0.974 0.000
C3 1.421 2.500 0.000
C4 0.006 -1.132 0.000
N5 -2.444 -2.104 0.000
C6 -1.348 -1.687 0.000
H7 2.440 2.893 0.000
H8 -0.547 0.759 0.879
H9 -0.547 0.759 -0.879
H10 1.954 0.602 0.878
H11 1.954 0.602 -0.878
H12 0.911 2.893 -0.883
H13 0.911 2.893 0.883
H14 0.536 -1.509 -0.880
H15 0.536 -1.509 0.880

Atom - Atom Distances (Å)
  C1 C2 C3 C4 N5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.52562.53381.53363.50042.48563.48711.09491.09492.15192.15192.79542.79542.17122.1712
C21.52561.52572.53344.93573.83502.17602.15962.15961.09611.09612.17132.17132.77712.7771
C32.53381.52573.89756.01105.01911.09232.77022.77022.15792.15791.09301.09304.19874.1987
C41.53362.53343.89752.63581.46274.70382.15702.15702.75222.75224.21844.21841.09451.0945
N53.50044.93576.01102.63581.17336.98753.54543.54545.23835.23836.08296.08293.16403.1640
C62.48563.83505.01911.46271.17335.94312.71952.71954.11254.11255.18185.18182.08672.0867
H73.48712.17601.09234.70386.98755.94313.77453.77452.50132.50131.76581.76584.87654.8765
H81.09492.15962.77022.15703.54542.71953.77451.75782.50583.06053.12752.58393.06782.5137
H91.09492.15962.77022.15703.54542.71953.77451.75783.06052.50582.58393.12752.51373.0678
H102.15191.09612.15792.75225.23834.11252.50132.50583.06051.75683.07222.51703.09192.5434
H112.15191.09612.15792.75225.23834.11252.50133.06052.50581.75682.51703.07222.54343.0919
H122.79542.17131.09304.21846.08295.18181.76583.12752.58393.07222.51701.76634.41794.7567
H132.79542.17131.09304.21846.08295.18181.76582.58393.12752.51703.07221.76634.75674.4179
H142.17122.77714.19871.09453.16402.08674.87653.06782.51373.09192.54344.41794.75671.7594
H152.17122.77714.19871.09453.16402.08674.87652.51373.06782.54343.09194.75674.41791.7594

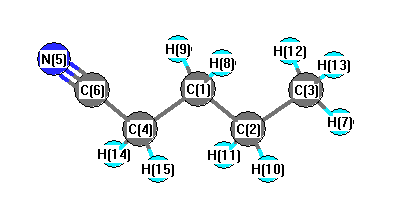
picture of Pentanenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.284 C1 C2 H10 109.230
C1 C2 H11 109.230 C1 C4 C6 112.083
C1 C4 H14 110.292 C1 C4 H15 110.292
C2 C1 C4 111.817 C2 C1 H8 109.902
C2 C1 H9 109.902 C2 C3 H7 111.367
C2 C3 H12 110.945 C2 C3 H13 110.945
C3 C2 H10 109.703 C3 C2 H11 109.703
C4 C1 H8 109.157 C4 C1 H9 109.157
C4 C6 N5 178.529 C6 C4 H14 108.518
C6 C4 H15 108.518 H7 C3 H12 107.811
H7 C3 H13 107.811 H8 C1 H9 106.771
H10 C2 H11 106.522 H12 C3 H13 107.807
H14 C4 H15 106.978
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability