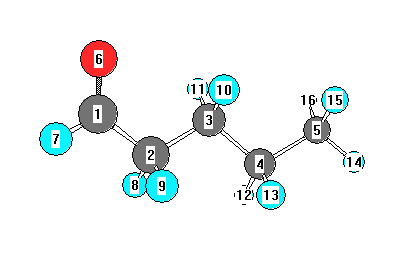
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3107 |
2991 |
43.68 |
|
|
|
| 2 |
A' |
3048 |
2935 |
37.63 |
|
|
|
| 3 |
A' |
3034 |
2920 |
25.98 |
|
|
|
| 4 |
A' |
3019 |
2906 |
35.56 |
|
|
|
| 5 |
A' |
3008 |
2896 |
20.24 |
|
|
|
| 6 |
A' |
2871 |
2764 |
179.89 |
|
|
|
| 7 |
A' |
1835 |
1766 |
161.43 |
|
|
|
| 8 |
A' |
1519 |
1463 |
11.32 |
|
|
|
| 9 |
A' |
1504 |
1448 |
1.82 |
|
|
|
| 10 |
A' |
1495 |
1439 |
0.94 |
|
|
|
| 11 |
A' |
1458 |
1404 |
17.61 |
|
|
|
| 12 |
A' |
1424 |
1371 |
14.28 |
|
|
|
| 13 |
A' |
1421 |
1368 |
2.97 |
|
|
|
| 14 |
A' |
1414 |
1361 |
6.13 |
|
|
|
| 15 |
A' |
1374 |
1323 |
14.60 |
|
|
|
| 16 |
A' |
1285 |
1237 |
7.27 |
|
|
|
| 17 |
A' |
1142 |
1100 |
11.87 |
|
|
|
| 18 |
A' |
1083 |
1043 |
0.43 |
|
|
|
| 19 |
A' |
1050 |
1011 |
1.45 |
|
|
|
| 20 |
A' |
926 |
891 |
0.52 |
|
|
|
| 21 |
A' |
898 |
865 |
14.16 |
|
|
|
| 22 |
A' |
697 |
671 |
13.25 |
|
|
|
| 23 |
A' |
395 |
381 |
1.92 |
|
|
|
| 24 |
A' |
290 |
279 |
4.21 |
|
|
|
| 25 |
A' |
136 |
131 |
5.18 |
|
|
|
| 26 |
A" |
3104 |
2988 |
78.60 |
|
|
|
| 27 |
A" |
3087 |
2972 |
18.17 |
|
|
|
| 28 |
A" |
3045 |
2932 |
7.63 |
|
|
|
| 29 |
A" |
3031 |
2918 |
10.65 |
|
|
|
| 30 |
A" |
1511 |
1454 |
9.95 |
|
|
|
| 31 |
A" |
1343 |
1293 |
0.19 |
|
|
|
| 32 |
A" |
1322 |
1272 |
0.10 |
|
|
|
| 33 |
A" |
1242 |
1195 |
0.01 |
|
|
|
| 34 |
A" |
1160 |
1117 |
0.16 |
|
|
|
| 35 |
A" |
984 |
947 |
0.91 |
|
|
|
| 36 |
A" |
858 |
826 |
0.17 |
|
|
|
| 37 |
A" |
743 |
715 |
2.38 |
|
|
|
| 38 |
A" |
665 |
640 |
4.40 |
|
|
|
| 39 |
A" |
236 |
227 |
0.00 |
|
|
|
| 40 |
A" |
185 |
178 |
1.22 |
|
|
|
| 41 |
A" |
111 |
107 |
1.03 |
|
|
|
| 42 |
A" |
65 |
63 |
1.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31061.5 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 29902.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.134 |
|
|
|
| 2 |
C |
-0.529 |
|
|
|
| 3 |
C |
-0.430 |
|
|
|
| 4 |
C |
-0.430 |
|
|
|
| 5 |
C |
-0.649 |
|
|
|
| 6 |
O |
-0.287 |
|
|
|
| 7 |
H |
0.163 |
|
|
|
| 8 |
H |
0.240 |
|
|
|
| 9 |
H |
0.240 |
|
|
|
| 10 |
H |
0.230 |
|
|
|
| 11 |
H |
0.230 |
|
|
|
| 12 |
H |
0.213 |
|
|
|
| 13 |
H |
0.213 |
|
|
|
| 14 |
H |
0.222 |
|
|
|
| 15 |
H |
0.220 |
|
|
|
| 16 |
H |
0.220 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.600 |
-0.109 |
0.000 |
2.602 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.952 |
3.963 |
0.000 |
| y |
3.963 |
-36.996 |
0.000 |
| z |
0.000 |
0.000 |
-37.223 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.842 |
3.963 |
0.000 |
| y |
3.963 |
4.092 |
0.000 |
| z |
0.000 |
0.000 |
3.751 |
|
| Polar |
|---|
| 3z2-r2 | 7.501 |
|---|
| x2-y2 | -7.956 |
|---|
| xy | 3.963 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.040 |
-0.404 |
0.000 |
| y |
-0.404 |
10.091 |
0.000 |
| z |
0.000 |
0.000 |
7.043 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |