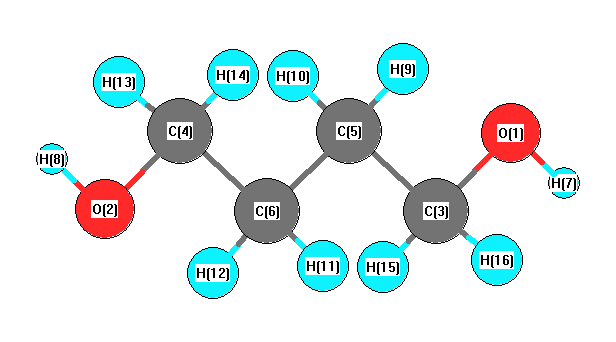
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3826 |
3683 |
0.00 |
|
|
|
| 2 |
Ag |
3033 |
2920 |
0.00 |
|
|
|
| 3 |
Ag |
2973 |
2863 |
0.00 |
|
|
|
| 4 |
Ag |
1537 |
1480 |
0.00 |
|
|
|
| 5 |
Ag |
1501 |
1445 |
0.00 |
|
|
|
| 6 |
Ag |
1475 |
1420 |
0.00 |
|
|
|
| 7 |
Ag |
1394 |
1342 |
0.00 |
|
|
|
| 8 |
Ag |
1281 |
1233 |
0.00 |
|
|
|
| 9 |
Ag |
1096 |
1055 |
0.00 |
|
|
|
| 10 |
Ag |
1077 |
1037 |
0.00 |
|
|
|
| 11 |
Ag |
1047 |
1008 |
0.00 |
|
|
|
| 12 |
Ag |
361 |
348 |
0.00 |
|
|
|
| 13 |
Ag |
339 |
327 |
0.00 |
|
|
|
| 14 |
Au |
3092 |
2976 |
98.20 |
|
|
|
| 15 |
Au |
3001 |
2889 |
110.78 |
|
|
|
| 16 |
Au |
1337 |
1287 |
1.83 |
|
|
|
| 17 |
Au |
1228 |
1182 |
3.27 |
|
|
|
| 18 |
Au |
955 |
919 |
3.39 |
|
|
|
| 19 |
Au |
760 |
731 |
0.86 |
|
|
|
| 20 |
Au |
302 |
291 |
281.57 |
|
|
|
| 21 |
Au |
97 |
93 |
18.67 |
|
|
|
| 22 |
Au |
79 |
76 |
2.20 |
|
|
|
| 23 |
Bg |
3067 |
2952 |
0.00 |
|
|
|
| 24 |
Bg |
3002 |
2890 |
0.00 |
|
|
|
| 25 |
Bg |
1331 |
1282 |
0.00 |
|
|
|
| 26 |
Bg |
1298 |
1249 |
0.00 |
|
|
|
| 27 |
Bg |
1181 |
1137 |
0.00 |
|
|
|
| 28 |
Bg |
821 |
791 |
0.00 |
|
|
|
| 29 |
Bg |
298 |
287 |
0.00 |
|
|
|
| 30 |
Bg |
156 |
150 |
0.00 |
|
|
|
| 31 |
Bu |
3826 |
3683 |
21.34 |
|
|
|
| 32 |
Bu |
3042 |
2928 |
78.52 |
|
|
|
| 33 |
Bu |
2973 |
2862 |
127.77 |
|
|
|
| 34 |
Bu |
1541 |
1483 |
6.29 |
|
|
|
| 35 |
Bu |
1510 |
1454 |
4.03 |
|
|
|
| 36 |
Bu |
1474 |
1419 |
7.76 |
|
|
|
| 37 |
Bu |
1321 |
1271 |
106.14 |
|
|
|
| 38 |
Bu |
1219 |
1174 |
47.02 |
|
|
|
| 39 |
Bu |
1090 |
1050 |
206.04 |
|
|
|
| 40 |
Bu |
1006 |
969 |
1.82 |
|
|
|
| 41 |
Bu |
524 |
504 |
43.16 |
|
|
|
| 42 |
Bu |
136 |
131 |
7.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31302.4 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 30134.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.577 |
|
|
|
| 2 |
O |
-0.577 |
|
|
|
| 3 |
C |
-0.172 |
|
|
|
| 4 |
C |
-0.172 |
|
|
|
| 5 |
C |
-0.467 |
|
|
|
| 6 |
C |
-0.467 |
|
|
|
| 7 |
H |
0.390 |
|
|
|
| 8 |
H |
0.390 |
|
|
|
| 9 |
H |
0.223 |
|
|
|
| 10 |
H |
0.223 |
|
|
|
| 11 |
H |
0.223 |
|
|
|
| 12 |
H |
0.223 |
|
|
|
| 13 |
H |
0.190 |
|
|
|
| 14 |
H |
0.190 |
|
|
|
| 15 |
H |
0.190 |
|
|
|
| 16 |
H |
0.190 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.324 |
8.809 |
0.000 |
| y |
8.809 |
-43.196 |
0.000 |
| z |
0.000 |
0.000 |
-38.233 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
13.391 |
8.809 |
0.000 |
| y |
8.809 |
-10.418 |
0.000 |
| z |
0.000 |
0.000 |
-2.973 |
|
| Polar |
|---|
| 3z2-r2 | -5.946 |
|---|
| x2-y2 | 15.872 |
|---|
| xy | 8.809 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
283.826 |
| (<r2>)1/2 |
16.847 |