return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12S (1-Pentanethiol)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-594.934293
Energy at 298.15K-594.947206
HF Energy-593.895735
Nuclear repulsion energy288.012211
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3166 2999 32.31      
2 A' 3099 2935 37.60      
3 A' 3079 2916 44.31      
4 A' 3076 2913 28.12      
5 A' 3073 2911 5.38      
6 A' 3059 2898 8.06      
7 A' 2682 2540 13.26      
8 A' 1550 1468 12.56      
9 A' 1540 1459 0.45      
10 A' 1530 1449 2.00      
11 A' 1526 1445 0.58      
12 A' 1521 1441 1.98      
13 A' 1450 1373 7.03      
14 A' 1433 1357 0.46      
15 A' 1404 1330 14.70      
16 A' 1344 1273 17.75      
17 A' 1275 1208 6.95      
18 A' 1165 1104 5.17      
19 A' 1110 1051 0.05      
20 A' 1100 1042 1.30      
21 A' 1062 1006 0.66      
22 A' 934 884 0.79      
23 A' 890 843 2.51      
24 A' 784 743 1.62      
25 A' 449 425 1.08      
26 A' 356 337 0.41      
27 A' 254 241 1.01      
28 A' 117 111 1.09      
29 A" 3166 2998 60.39      
30 A" 3164 2997 14.96      
31 A" 3136 2970 27.83      
32 A" 3123 2957 3.73      
33 A" 3102 2938 1.71      
34 A" 1542 1460 10.23      
35 A" 1367 1294 1.45      
36 A" 1364 1292 0.08      
37 A" 1321 1252 0.60      
38 A" 1254 1188 0.53      
39 A" 1123 1064 5.06      
40 A" 993 941 0.34      
41 A" 867 821 2.55      
42 A" 777 736 0.45      
43 A" 739 700 5.07      
44 A" 252 239 0.04      
45 A" 171 162 15.52      
46 A" 141 134 6.97      
47 A" 98 92 4.29      
48 A" 64 61 2.63      

Unscaled Zero Point Vibrational Energy (zpe) 35896.0 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 33997.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.47761 0.02689 0.02608

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 -2.383 1.688 0.000
H2 -3.627 1.180 0.000
C3 -1.500 0.095 0.000
H4 -1.774 -0.478 0.888
H5 -1.774 -0.478 -0.888
C6 0.000 0.358 0.000
H7 0.264 0.955 -0.880
H8 0.264 0.955 0.880
C9 0.812 -0.934 0.000
H10 0.543 -1.534 0.879
H11 0.543 -1.534 -0.879
C12 2.318 -0.690 0.000
H13 2.583 -0.089 0.878
H14 2.583 -0.089 -0.878
C15 3.124 -1.985 0.000
H16 2.896 -2.587 0.883
H17 2.896 -2.587 -0.883
H18 4.199 -1.788 0.000

Atom - Atom Distances (Å)
  S1 H2 C3 H4 H5 C6 H7 H8 C9 H10 H11 C12 H13 H14 C15 H16 H17 H18
S11.34431.82122.41902.41902.72882.88442.88444.13314.44034.44035.26805.34705.34706.61966.84986.84987.4435
H21.34432.38842.64062.64063.71953.99653.99654.91715.05335.05336.23276.39976.39977.45697.58447.58448.3706
C31.82122.38841.09251.09251.52262.15082.15082.53042.75712.75713.89774.18044.18045.07045.22445.22446.0020
H42.41902.64061.09251.77702.15343.05592.49202.77232.54713.10034.19334.37524.71825.20185.12435.42186.1798
H52.41902.64061.09251.77702.15342.49203.05592.77233.10032.54714.19334.71824.37525.20185.42185.12436.1798
C62.72883.71951.52262.15342.15341.09591.09591.52632.15592.15592.54392.76482.76483.90524.22374.22374.7158
H72.88443.99652.15083.05592.49201.09591.76062.15503.06062.50432.77443.09152.54314.19484.75184.41264.8766
H82.88443.99652.15082.49203.05591.09591.76062.15502.50433.06062.77442.54313.09154.19484.41264.75184.8766
C94.13314.91712.53042.77232.77231.52632.15502.15501.09741.09741.52632.15042.15042.54002.80262.80263.4934
H104.44035.05332.75712.54713.10032.15593.06062.50431.09741.75832.15302.49993.05542.76362.57723.12213.7686
H114.44035.05332.75713.10032.54712.15592.50433.06061.09741.75832.15303.05542.49992.76363.12212.57723.7686
C125.26806.23273.89774.19334.19332.54392.77442.77441.52632.15302.15301.09601.09601.52572.17142.17142.1783
H135.34706.39974.18044.37524.71822.76483.09152.54312.15042.49993.05541.09601.75522.15812.51803.07252.5036
H145.34706.39974.18044.71824.37522.76482.54313.09152.15043.05542.49991.09601.75522.15813.07252.51802.5036
C156.61967.45695.07045.20185.20183.90524.19484.19482.54002.76362.76361.52572.15812.15811.09331.09331.0927
H166.84987.58445.22445.12435.42184.22374.75184.41262.80262.57723.12212.17142.51803.07251.09331.76611.7659
H176.84987.58445.22445.42185.12434.22374.41264.75182.80263.12212.57722.17143.07252.51801.09331.76611.7659
H187.44358.37066.00206.17986.17984.71584.87664.87663.49343.76863.76862.17832.50362.50361.09271.76591.7659

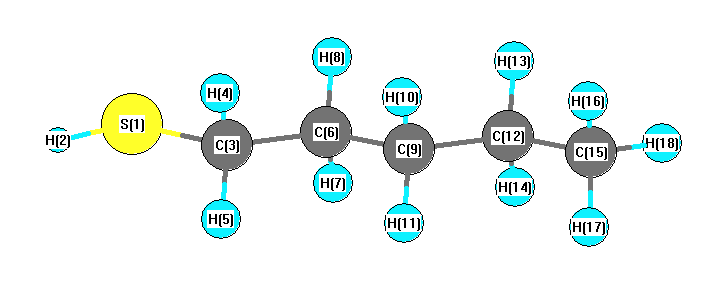
picture of 1-Pentanethiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C3 H4 109.696 S1 C3 H5 109.696
S1 C3 C6 109.065 H2 S1 C3 96.806
C3 C6 H7 109.366 C3 C6 H8 109.366
C3 C6 C9 112.190 H4 C3 H5 108.833
H4 C3 C6 109.770 H5 C3 C6 109.770
C6 C9 H10 109.427 C6 C9 H11 109.427
C6 C9 C12 112.895 H7 C6 H8 106.888
H7 C6 C9 109.443 H8 C6 C9 109.443
C9 C12 H13 109.073 C9 C12 H14 109.073
C9 C12 C15 112.665 H10 C9 H11 106.479
H10 C9 C12 109.205 H11 C9 C12 109.205
C12 C15 H16 110.936 C12 C15 H17 110.936
C12 C15 H18 111.522 H13 C12 H14 106.396
H13 C12 C15 109.717 H14 C12 C15 109.717
H16 C15 H17 107.741 H16 C15 H18 107.771
H17 C15 H18 107.771
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability