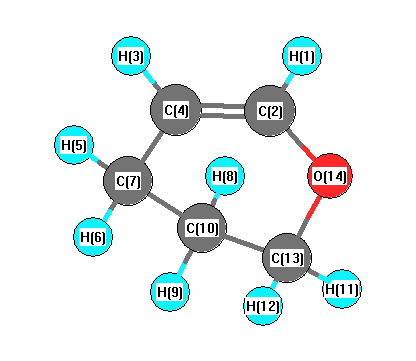
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3206 |
3087 |
30.75 |
|
|
|
| 2 |
A |
3183 |
3064 |
3.52 |
|
|
|
| 3 |
A |
3120 |
3003 |
40.54 |
|
|
|
| 4 |
A |
3096 |
2980 |
44.78 |
|
|
|
| 5 |
A |
3058 |
2944 |
49.12 |
|
|
|
| 6 |
A |
3045 |
2931 |
12.02 |
|
|
|
| 7 |
A |
3019 |
2906 |
54.00 |
|
|
|
| 8 |
A |
3011 |
2898 |
36.46 |
|
|
|
| 9 |
A |
1722 |
1658 |
78.59 |
|
|
|
| 10 |
A |
1514 |
1457 |
3.79 |
|
|
|
| 11 |
A |
1498 |
1442 |
4.50 |
|
|
|
| 12 |
A |
1485 |
1429 |
3.60 |
|
|
|
| 13 |
A |
1437 |
1383 |
2.81 |
|
|
|
| 14 |
A |
1404 |
1352 |
3.34 |
|
|
|
| 15 |
A |
1378 |
1326 |
1.61 |
|
|
|
| 16 |
A |
1356 |
1305 |
3.90 |
|
|
|
| 17 |
A |
1313 |
1264 |
3.82 |
|
|
|
| 18 |
A |
1283 |
1235 |
100.37 |
|
|
|
| 19 |
A |
1263 |
1216 |
6.32 |
|
|
|
| 20 |
A |
1220 |
1174 |
4.18 |
|
|
|
| 21 |
A |
1111 |
1069 |
25.61 |
|
|
|
| 22 |
A |
1105 |
1064 |
57.28 |
|
|
|
| 23 |
A |
1073 |
1033 |
9.45 |
|
|
|
| 24 |
A |
1041 |
1002 |
3.06 |
|
|
|
| 25 |
A |
955 |
919 |
12.66 |
|
|
|
| 26 |
A |
941 |
906 |
0.67 |
|
|
|
| 27 |
A |
916 |
881 |
24.40 |
|
|
|
| 28 |
A |
891 |
858 |
1.40 |
|
|
|
| 29 |
A |
856 |
824 |
4.48 |
|
|
|
| 30 |
A |
765 |
736 |
7.79 |
|
|
|
| 31 |
A |
742 |
714 |
26.46 |
|
|
|
| 32 |
A |
512 |
492 |
6.87 |
|
|
|
| 33 |
A |
495 |
477 |
2.01 |
|
|
|
| 34 |
A |
450 |
433 |
15.71 |
|
|
|
| 35 |
A |
280 |
270 |
0.57 |
|
|
|
| 36 |
A |
183 |
176 |
3.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26960.0 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 25954.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.211 |
|
|
|
| 2 |
C |
0.012 |
|
|
|
| 3 |
H |
0.202 |
|
|
|
| 4 |
C |
-0.305 |
|
|
|
| 5 |
H |
0.229 |
|
|
|
| 6 |
H |
0.225 |
|
|
|
| 7 |
C |
-0.433 |
|
|
|
| 8 |
H |
0.236 |
|
|
|
| 9 |
H |
0.223 |
|
|
|
| 10 |
C |
-0.498 |
|
|
|
| 11 |
H |
0.221 |
|
|
|
| 12 |
H |
0.216 |
|
|
|
| 13 |
C |
-0.229 |
|
|
|
| 14 |
O |
-0.311 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.126 |
-0.569 |
0.270 |
1.290 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.877 |
0.825 |
0.128 |
| y |
0.825 |
-33.441 |
-0.577 |
| z |
0.128 |
-0.577 |
-37.792 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.260 |
0.825 |
0.128 |
| y |
0.825 |
4.393 |
-0.577 |
| z |
0.128 |
-0.577 |
-2.133 |
|
| Polar |
|---|
| 3z2-r2 | -4.266 |
|---|
| x2-y2 | -4.436 |
|---|
| xy | 0.825 |
|---|
| xz | 0.128 |
|---|
| yz | -0.577 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
140.711 |
| (<r2>)1/2 |
11.862 |