return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8O (2H-Pyran, 3,4-dihydro-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-269.866360
Energy at 298.15K-269.876348
HF Energy-268.885039
Nuclear repulsion energy241.696987
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3243 3072 24.94      
2 A 3221 3051 1.20      
3 A 3173 3006 30.24      
4 A 3153 2986 30.54      
5 A 3117 2952 24.45      
6 A 3092 2928 18.80      
7 A 3068 2905 41.29      
8 A 3063 2901 35.81      
9 A 1710 1620 66.68      
10 A 1544 1463 3.82      
11 A 1529 1448 4.55      
12 A 1517 1437 4.23      
13 A 1450 1374 2.08      
14 A 1419 1344 4.24      
15 A 1398 1324 1.66      
16 A 1370 1297 3.88      
17 A 1329 1259 4.09      
18 A 1301 1232 99.74      
19 A 1279 1212 6.99      
20 A 1236 1171 3.84      
21 A 1129 1070 14.23      
22 A 1122 1063 60.07      
23 A 1092 1035 9.56      
24 A 1050 994 3.28      
25 A 967 916 14.30      
26 A 933 884 17.78      
27 A 924 875 3.32      
28 A 906 858 1.03      
29 A 870 824 5.28      
30 A 772 732 9.22      
31 A 747 707 25.04      
32 A 515 487 6.31      
33 A 495 469 2.30      
34 A 452 428 16.52      
35 A 294 278 0.67      
36 A 186 176 3.53      

Unscaled Zero Point Vibrational Energy (zpe) 27332.7 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 25886.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.17550 0.15817 0.09103

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 1.659 1.832 -0.199
C2 0.896 1.068 -0.083
H3 -0.698 2.396 0.113
C4 -0.409 1.351 0.071
H5 -2.298 0.539 -0.530
H6 -1.871 0.172 1.132
C7 -1.457 0.274 0.120
H8 -0.702 -1.044 -1.419
H9 -1.463 -1.901 -0.072
C10 -0.832 -1.045 -0.332
H11 1.016 -2.140 0.052
H12 0.446 -1.130 1.405
C13 0.535 -1.196 0.312
O14 1.439 -0.179 -0.127

Atom - Atom Distances (Å)
  H1 C2 H3 C4 H5 H6 C7 H8 H9 C10 H11 H12 C13 O14
H11.08562.44272.13974.17574.12103.49843.91564.86813.80824.03173.58013.27002.0244
C21.08562.08361.34393.26823.15202.49212.96603.79242.74153.21322.69262.32661.3607
H32.44272.08361.08532.53422.71282.25403.76514.36843.47254.84903.92553.80303.3542
C42.13971.34391.08532.14242.15651.50382.83543.42132.46643.77002.94302.72672.4065
H54.17573.26822.53422.14241.75421.09532.41702.61882.16724.30043.74913.42743.8262
H64.12103.15202.71282.15651.75421.09733.05792.43202.16913.85292.67162.88703.5582
C73.49842.49212.25401.50381.09531.09732.16232.18321.52833.45622.69102.48362.9415
H83.91562.96603.76512.83542.41703.05792.16231.76851.09452.51313.04942.13362.6458
H94.86813.79244.36843.42132.61882.43202.18321.76851.09442.49352.53402.15413.3749
C103.80822.74153.47252.46642.16722.16911.52831.09451.09442.18182.15821.51962.4393
H114.03173.21324.84903.77004.30043.85293.45622.51312.49352.18181.78191.09062.0142
H123.58012.69263.92552.94303.74912.67162.69103.04942.53402.15821.78191.09812.0583
C133.27002.32663.80302.72673.42742.88702.48362.13362.15411.51961.09061.09811.4294
O142.02441.36073.35422.40653.82623.55822.94152.64583.37492.43932.01422.05831.4294

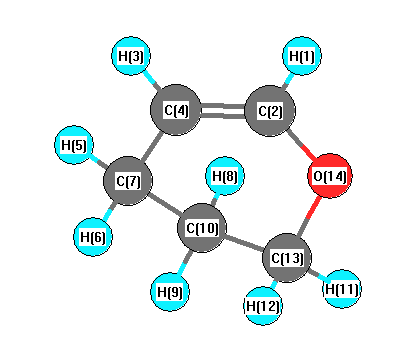
picture of 2H-Pyran, 3,4-dihydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 123.109 H1 C2 O14 111.194
C2 C4 H3 117.734 C2 C4 C7 122.020
C2 O14 C13 112.979 H3 C4 C7 120.179
C4 C2 O14 125.695 C4 C7 H5 110.035
C4 C7 H6 111.043 C4 C7 C10 108.862
H5 C7 H6 106.262 H5 C7 C10 110.299
H6 C7 C10 110.326 C7 C10 H8 109.962
C7 C10 H9 111.634 C7 C10 C13 109.148
H8 C10 H9 107.794 H8 C10 C13 108.317
H9 C10 C13 109.923 C10 C13 H11 112.381
C10 C13 H12 110.030 C10 C13 O14 111.583
H11 C13 H12 109.002 H11 C13 O14 105.330
H12 C13 O14 108.332
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability