Jump to
S1C2
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -250.235273 |
| Energy at 298.15K | -250.250009 |
| Nuclear repulsion energy | 260.068012 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3782 |
3420 |
0.60 |
|
|
|
| 2 |
A' |
3225 |
2916 |
148.49 |
|
|
|
| 3 |
A' |
3218 |
2911 |
23.22 |
|
|
|
| 4 |
A' |
3205 |
2899 |
107.49 |
|
|
|
| 5 |
A' |
3177 |
2873 |
9.55 |
|
|
|
| 6 |
A' |
3157 |
2856 |
36.91 |
|
|
|
| 7 |
A' |
3083 |
2788 |
176.36 |
|
|
|
| 8 |
A' |
1655 |
1497 |
1.83 |
|
|
|
| 9 |
A' |
1633 |
1477 |
4.38 |
|
|
|
| 10 |
A' |
1620 |
1465 |
5.24 |
|
|
|
| 11 |
A' |
1570 |
1420 |
4.61 |
|
|
|
| 12 |
A' |
1516 |
1371 |
0.16 |
|
|
|
| 13 |
A' |
1433 |
1296 |
2.15 |
|
|
|
| 14 |
A' |
1401 |
1267 |
4.36 |
|
|
|
| 15 |
A' |
1281 |
1158 |
8.05 |
|
|
|
| 16 |
A' |
1155 |
1044 |
16.20 |
|
|
|
| 17 |
A' |
1123 |
1016 |
0.65 |
|
|
|
| 18 |
A' |
973 |
880 |
7.94 |
|
|
|
| 19 |
A' |
948 |
857 |
18.79 |
|
|
|
| 20 |
A' |
871 |
788 |
1.24 |
|
|
|
| 21 |
A' |
856 |
774 |
76.79 |
|
|
|
| 22 |
A' |
587 |
531 |
22.61 |
|
|
|
| 23 |
A' |
463 |
419 |
8.12 |
|
|
|
| 24 |
A' |
424 |
384 |
1.75 |
|
|
|
| 25 |
A' |
259 |
234 |
2.33 |
|
|
|
| 26 |
A" |
3218 |
2911 |
77.51 |
|
|
|
| 27 |
A" |
3212 |
2905 |
92.75 |
|
|
|
| 28 |
A" |
3177 |
2874 |
43.59 |
|
|
|
| 29 |
A" |
3074 |
2780 |
22.96 |
|
|
|
| 30 |
A" |
1653 |
1495 |
4.06 |
|
|
|
| 31 |
A" |
1631 |
1475 |
1.11 |
|
|
|
| 32 |
A" |
1612 |
1458 |
3.24 |
|
|
|
| 33 |
A" |
1518 |
1373 |
0.79 |
|
|
|
| 34 |
A" |
1500 |
1356 |
31.48 |
|
|
|
| 35 |
A" |
1478 |
1337 |
7.37 |
|
|
|
| 36 |
A" |
1408 |
1273 |
2.36 |
|
|
|
| 37 |
A" |
1288 |
1165 |
4.67 |
|
|
|
| 38 |
A" |
1273 |
1152 |
5.66 |
|
|
|
| 39 |
A" |
1242 |
1123 |
17.11 |
|
|
|
| 40 |
A" |
1133 |
1024 |
1.64 |
|
|
|
| 41 |
A" |
1062 |
960 |
0.84 |
|
|
|
| 42 |
A" |
938 |
849 |
0.04 |
|
|
|
| 43 |
A" |
890 |
805 |
0.63 |
|
|
|
| 44 |
A" |
482 |
436 |
1.00 |
|
|
|
| 45 |
A" |
263 |
238 |
0.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37332.5 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 33763.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.628 |
1.331 |
0.000 |
| H2 |
0.547 |
2.414 |
0.000 |
| H3 |
1.693 |
1.101 |
0.000 |
| C4 |
-0.015 |
0.747 |
1.260 |
| C5 |
-0.015 |
0.747 |
-1.260 |
| C6 |
-0.015 |
-0.778 |
-1.210 |
| C7 |
-0.015 |
-0.778 |
1.210 |
| N8 |
-0.680 |
-1.231 |
0.000 |
| H9 |
-0.750 |
-2.226 |
0.000 |
| H10 |
0.508 |
1.088 |
2.148 |
| H11 |
0.508 |
1.088 |
-2.148 |
| H12 |
-1.043 |
1.091 |
1.339 |
| H13 |
-1.043 |
1.091 |
-1.339 |
| H14 |
1.017 |
-1.140 |
-1.273 |
| H15 |
1.017 |
-1.140 |
1.273 |
| H16 |
-0.546 |
-1.180 |
-2.066 |
| H17 |
-0.546 |
-1.180 |
2.066 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0861 | 1.0892 | 1.5303 | 1.5303 | 2.5149 | 2.5149 | 2.8764 | 3.8146 | 2.1653 | 2.1653 | 2.1549 | 2.1549 | 2.8067 | 2.8067 | 3.4572 | 3.4572 |
H2 | 1.0861 | | 1.7427 | 2.1639 | 2.1639 | 3.4594 | 3.4594 | 3.8459 | 4.8179 | 2.5249 | 2.5249 | 2.4641 | 2.4641 | 3.8043 | 3.8043 | 4.2871 | 4.2871 | H3 | 1.0892 | 1.7427 | | 2.1517 | 2.1517 | 2.8126 | 2.8126 | 3.3265 | 4.1274 | 2.4532 | 2.4532 | 3.0459 | 3.0459 | 2.6643 | 2.6643 | 3.8057 | 3.8057 | C4 | 1.5303 | 2.1639 | 2.1517 | | 2.5191 | 2.9023 | 1.5252 | 2.4370 | 3.3111 | 1.0866 | 3.4647 | 1.0867 | 2.8156 | 3.3224 | 2.1510 | 3.8800 | 2.1549 | C5 | 1.5303 | 2.1639 | 2.1517 | 2.5191 | | 1.5252 | 2.9023 | 2.4370 | 3.3111 | 3.4647 | 1.0866 | 2.8156 | 1.0867 | 2.1510 | 3.3224 | 2.1549 | 3.8800 | C6 | 2.5149 | 3.4594 | 2.8126 | 2.9023 | 1.5252 | | 2.4205 | 1.4532 | 2.0255 | 3.8775 | 2.1528 | 3.3238 | 2.1366 | 1.0959 | 2.7133 | 1.0847 | 3.3437 | C7 | 2.5149 | 3.4594 | 2.8126 | 1.5252 | 2.9023 | 2.4205 | | 1.4532 | 2.0255 | 2.1528 | 3.8775 | 2.1366 | 3.3238 | 2.7133 | 1.0959 | 3.3437 | 1.0847 | N8 | 2.8764 | 3.8459 | 3.3265 | 2.4370 | 2.4370 | 1.4532 | 1.4532 | | 0.9975 | 3.3770 | 3.3770 | 2.7051 | 2.7051 | 2.1229 | 2.1229 | 2.0713 | 2.0713 | H9 | 3.8146 | 4.8179 | 4.1274 | 3.3111 | 3.3111 | 2.0255 | 2.0255 | 0.9975 | | 4.1451 | 4.1451 | 3.5892 | 3.5892 | 2.4334 | 2.4334 | 2.3252 | 2.3252 | H10 | 2.1653 | 2.5249 | 2.4532 | 1.0866 | 3.4647 | 3.8775 | 2.1528 | 3.3770 | 4.1451 | | 4.2967 | 1.7498 | 3.8169 | 4.1142 | 2.4475 | 4.9008 | 2.5022 | H11 | 2.1653 | 2.5249 | 2.4532 | 3.4647 | 1.0866 | 2.1528 | 3.8775 | 3.3770 | 4.1451 | 4.2967 | | 3.8169 | 1.7498 | 2.4475 | 4.1142 | 2.5022 | 4.9008 | H12 | 2.1549 | 2.4641 | 3.0459 | 1.0867 | 2.8156 | 3.3238 | 2.1366 | 2.7051 | 3.5892 | 1.7498 | 3.8169 | | 2.6782 | 4.0055 | 3.0376 | 4.1232 | 2.4356 | H13 | 2.1549 | 2.4641 | 3.0459 | 2.8156 | 1.0867 | 2.1366 | 3.3238 | 2.7051 | 3.5892 | 3.8169 | 1.7498 | 2.6782 | | 3.0376 | 4.0055 | 2.4356 | 4.1232 | H14 | 2.8067 | 3.8043 | 2.6643 | 3.3224 | 2.1510 | 1.0959 | 2.7133 | 2.1229 | 2.4334 | 4.1142 | 2.4475 | 4.0055 | 3.0376 | | 2.5453 | 1.7537 | 3.6871 | H15 | 2.8067 | 3.8043 | 2.6643 | 2.1510 | 3.3224 | 2.7133 | 1.0959 | 2.1229 | 2.4334 | 2.4475 | 4.1142 | 3.0376 | 4.0055 | 2.5453 | | 3.6871 | 1.7537 | H16 | 3.4572 | 4.2871 | 3.8057 | 3.8800 | 2.1549 | 1.0847 | 3.3437 | 2.0713 | 2.3252 | 4.9008 | 2.5022 | 4.1232 | 2.4356 | 1.7537 | 3.6871 | | 4.1328 | H17 | 3.4572 | 4.2871 | 3.8057 | 2.1549 | 3.8800 | 3.3437 | 1.0847 | 2.0713 | 2.3252 | 2.5022 | 4.9008 | 2.4356 | 4.1232 | 3.6871 | 1.7537 | 4.1328 | |
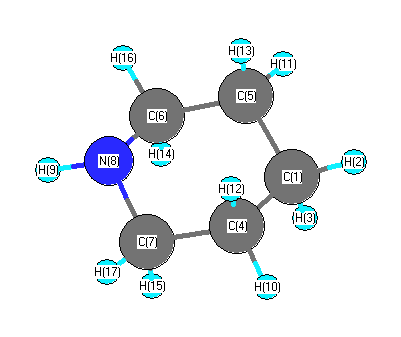 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.787 |
|
C1 |
C4 |
H10 |
110.525 |
| C1 |
C4 |
H12 |
109.692 |
|
C1 |
C5 |
C6 |
110.787 |
| C1 |
C5 |
H11 |
110.525 |
|
C1 |
C5 |
H13 |
109.692 |
| H2 |
C1 |
H3 |
106.473 |
|
H2 |
C1 |
C4 |
110.444 |
| H2 |
C1 |
C5 |
110.444 |
|
H3 |
C1 |
C4 |
109.294 |
| H3 |
C1 |
C5 |
109.294 |
|
C4 |
C1 |
C5 |
110.785 |
| C4 |
C7 |
N8 |
109.798 |
|
C4 |
C7 |
H15 |
109.205 |
| C4 |
C7 |
H17 |
110.175 |
|
C5 |
C6 |
N8 |
109.798 |
| C5 |
C6 |
H14 |
109.205 |
|
C5 |
C6 |
H16 |
110.175 |
| C6 |
C5 |
H11 |
109.890 |
|
C6 |
C5 |
H13 |
108.615 |
| C6 |
N8 |
C7 |
112.784 |
|
C6 |
N8 |
H9 |
110.091 |
| C7 |
C4 |
H10 |
109.890 |
|
C7 |
C4 |
H12 |
108.615 |
| C7 |
N8 |
H9 |
110.091 |
|
N8 |
C6 |
H14 |
112.013 |
| N8 |
C6 |
H16 |
108.538 |
|
N8 |
C7 |
H15 |
112.013 |
| N8 |
C7 |
H17 |
108.538 |
|
H10 |
C4 |
H12 |
107.245 |
| H11 |
C5 |
H13 |
107.245 |
|
H14 |
C6 |
H16 |
107.070 |
| H15 |
C7 |
H17 |
107.070 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.430 |
|
|
|
| 2 |
H |
0.211 |
|
|
|
| 3 |
H |
0.197 |
|
|
|
| 4 |
C |
-0.392 |
|
|
|
| 5 |
C |
-0.392 |
|
|
|
| 6 |
C |
-0.236 |
|
|
|
| 7 |
C |
-0.236 |
|
|
|
| 8 |
N |
-0.631 |
|
|
|
| 9 |
H |
0.320 |
|
|
|
| 10 |
H |
0.204 |
|
|
|
| 11 |
H |
0.204 |
|
|
|
| 12 |
H |
0.210 |
|
|
|
| 13 |
H |
0.210 |
|
|
|
| 14 |
H |
0.175 |
|
|
|
| 15 |
H |
0.175 |
|
|
|
| 16 |
H |
0.206 |
|
|
|
| 17 |
H |
0.206 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.827 |
-0.502 |
0.000 |
0.968 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.731 |
-0.302 |
0.000 |
| y |
-0.302 |
-36.958 |
0.000 |
| z |
0.000 |
0.000 |
-38.429 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.037 |
-0.302 |
0.000 |
| y |
-0.302 |
3.121 |
0.000 |
| z |
0.000 |
0.000 |
0.916 |
|
| Polar |
|---|
| 3z2-r2 | 1.831 |
|---|
| x2-y2 | -4.772 |
|---|
| xy | -0.302 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.856 |
0.239 |
0.000 |
| y |
0.239 |
8.709 |
0.000 |
| z |
0.000 |
0.000 |
9.108 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
Jump to
S1C1
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -250.233570 |
| Energy at 298.15K | -250.248268 |
| Nuclear repulsion energy | 259.680794 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3755 |
3396 |
1.28 |
|
|
|
| 2 |
A' |
3227 |
2919 |
101.55 |
|
|
|
| 3 |
A' |
3209 |
2902 |
126.68 |
|
|
|
| 4 |
A' |
3205 |
2898 |
79.09 |
|
|
|
| 5 |
A' |
3177 |
2873 |
54.91 |
|
|
|
| 6 |
A' |
3159 |
2857 |
35.28 |
|
|
|
| 7 |
A' |
3155 |
2853 |
29.56 |
|
|
|
| 8 |
A' |
1643 |
1486 |
1.80 |
|
|
|
| 9 |
A' |
1628 |
1472 |
6.07 |
|
|
|
| 10 |
A' |
1621 |
1466 |
4.56 |
|
|
|
| 11 |
A' |
1535 |
1388 |
9.58 |
|
|
|
| 12 |
A' |
1515 |
1371 |
0.66 |
|
|
|
| 13 |
A' |
1461 |
1322 |
0.23 |
|
|
|
| 14 |
A' |
1395 |
1261 |
10.75 |
|
|
|
| 15 |
A' |
1301 |
1176 |
3.36 |
|
|
|
| 16 |
A' |
1132 |
1023 |
3.01 |
|
|
|
| 17 |
A' |
1091 |
987 |
12.93 |
|
|
|
| 18 |
A' |
982 |
888 |
19.70 |
|
|
|
| 19 |
A' |
937 |
847 |
36.07 |
|
|
|
| 20 |
A' |
869 |
786 |
10.32 |
|
|
|
| 21 |
A' |
844 |
763 |
96.01 |
|
|
|
| 22 |
A' |
598 |
541 |
1.92 |
|
|
|
| 23 |
A' |
468 |
423 |
1.16 |
|
|
|
| 24 |
A' |
413 |
374 |
6.39 |
|
|
|
| 25 |
A' |
256 |
231 |
4.74 |
|
|
|
| 26 |
A" |
3224 |
2915 |
75.61 |
|
|
|
| 27 |
A" |
3199 |
2893 |
124.47 |
|
|
|
| 28 |
A" |
3171 |
2868 |
6.16 |
|
|
|
| 29 |
A" |
3151 |
2850 |
25.86 |
|
|
|
| 30 |
A" |
1649 |
1492 |
14.54 |
|
|
|
| 31 |
A" |
1624 |
1469 |
2.36 |
|
|
|
| 32 |
A" |
1613 |
1458 |
3.18 |
|
|
|
| 33 |
A" |
1515 |
1370 |
0.53 |
|
|
|
| 34 |
A" |
1505 |
1361 |
3.00 |
|
|
|
| 35 |
A" |
1474 |
1333 |
1.19 |
|
|
|
| 36 |
A" |
1410 |
1276 |
0.59 |
|
|
|
| 37 |
A" |
1323 |
1196 |
13.40 |
|
|
|
| 38 |
A" |
1233 |
1115 |
26.94 |
|
|
|
| 39 |
A" |
1214 |
1098 |
0.46 |
|
|
|
| 40 |
A" |
1132 |
1024 |
2.57 |
|
|
|
| 41 |
A" |
1031 |
932 |
4.17 |
|
|
|
| 42 |
A" |
933 |
843 |
0.06 |
|
|
|
| 43 |
A" |
876 |
792 |
0.32 |
|
|
|
| 44 |
A" |
486 |
440 |
0.69 |
|
|
|
| 45 |
A" |
252 |
228 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37293.0 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 33727.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.638 |
1.324 |
0.000 |
| H2 |
0.565 |
2.407 |
0.000 |
| H3 |
1.701 |
1.086 |
0.000 |
| C4 |
-0.009 |
0.741 |
1.260 |
| C5 |
-0.009 |
0.741 |
-1.260 |
| C6 |
-0.009 |
-0.789 |
-1.211 |
| C7 |
-0.009 |
-0.789 |
1.211 |
| N8 |
-0.601 |
-1.337 |
0.000 |
| H9 |
-1.585 |
-1.164 |
0.000 |
| H10 |
0.509 |
1.087 |
2.151 |
| H11 |
0.509 |
1.087 |
-2.151 |
| H12 |
-1.035 |
1.097 |
1.339 |
| H13 |
-1.035 |
1.097 |
-1.339 |
| H14 |
1.015 |
-1.152 |
-1.272 |
| H15 |
1.015 |
-1.152 |
1.272 |
| H16 |
-0.533 |
-1.201 |
-2.066 |
| H17 |
-0.533 |
-1.201 |
2.066 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0860 | 1.0891 | 1.5311 | 1.5311 | 2.5198 | 2.5198 | 2.9354 | 3.3359 | 2.1674 | 2.1674 | 2.1548 | 2.1548 | 2.8092 | 2.8092 | 3.4663 | 3.4663 |
H2 | 1.0860 | | 1.7421 | 2.1662 | 2.1662 | 3.4661 | 3.4661 | 3.9222 | 4.1684 | 2.5242 | 2.5242 | 2.4638 | 2.4638 | 3.8068 | 3.8068 | 4.3007 | 4.3007 | H3 | 1.0891 | 1.7421 | | 2.1513 | 2.1513 | 2.8117 | 2.8117 | 3.3423 | 3.9821 | 2.4585 | 2.4585 | 3.0461 | 3.0461 | 2.6646 | 2.6646 | 3.8068 | 3.8068 | C4 | 1.5311 | 2.1662 | 2.1513 | | 2.5195 | 2.9063 | 1.5313 | 2.5018 | 2.7750 | 1.0871 | 3.4668 | 1.0895 | 2.8165 | 3.3233 | 2.1525 | 3.8872 | 2.1675 | C5 | 1.5311 | 2.1662 | 2.1513 | 2.5195 | | 1.5313 | 2.9063 | 2.5018 | 2.7750 | 3.4668 | 1.0871 | 2.8165 | 1.0895 | 2.1525 | 3.3233 | 2.1675 | 3.8872 | C6 | 2.5198 | 3.4661 | 2.8117 | 2.9063 | 1.5313 | | 2.4218 | 1.4551 | 2.0226 | 3.8844 | 2.1615 | 3.3337 | 2.1516 | 1.0876 | 2.7101 | 1.0847 | 3.3444 | C7 | 2.5198 | 3.4661 | 2.8117 | 1.5313 | 2.9063 | 2.4218 | | 1.4551 | 2.0226 | 2.1615 | 3.8844 | 2.1516 | 3.3337 | 2.7101 | 1.0876 | 3.3444 | 1.0847 | N8 | 2.9354 | 3.9222 | 3.3423 | 2.5018 | 2.5018 | 1.4551 | 1.4551 | | 0.9994 | 3.4258 | 3.4258 | 2.8123 | 2.8123 | 2.0646 | 2.0646 | 2.0719 | 2.0719 | H9 | 3.3359 | 4.1684 | 3.9821 | 2.7750 | 2.7750 | 2.0226 | 2.0226 | 0.9994 | | 3.7521 | 3.7521 | 2.6844 | 2.6844 | 2.8943 | 2.8943 | 2.3188 | 2.3188 | H10 | 2.1674 | 2.5242 | 2.4585 | 1.0871 | 3.4668 | 3.8844 | 2.1615 | 3.4258 | 3.7521 | | 4.3012 | 1.7454 | 3.8160 | 4.1214 | 2.4580 | 4.9097 | 2.5159 | H11 | 2.1674 | 2.5242 | 2.4585 | 3.4668 | 1.0871 | 2.1615 | 3.8844 | 3.4258 | 3.7521 | 4.3012 | | 3.8160 | 1.7454 | 2.4580 | 4.1214 | 2.5159 | 4.9097 | H12 | 2.1548 | 2.4638 | 3.0461 | 1.0895 | 2.8165 | 3.3337 | 2.1516 | 2.8123 | 2.6844 | 1.7454 | 3.8160 | | 2.6773 | 4.0100 | 3.0444 | 4.1385 | 2.4624 | H13 | 2.1548 | 2.4638 | 3.0461 | 2.8165 | 1.0895 | 2.1516 | 3.3337 | 2.8123 | 2.6844 | 3.8160 | 1.7454 | 2.6773 | | 3.0444 | 4.0100 | 2.4624 | 4.1385 | H14 | 2.8092 | 3.8068 | 2.6646 | 3.3233 | 2.1525 | 1.0876 | 2.7101 | 2.0646 | 2.8943 | 4.1214 | 2.4580 | 4.0100 | 3.0444 | | 2.5444 | 1.7405 | 3.6803 | H15 | 2.8092 | 3.8068 | 2.6646 | 2.1525 | 3.3233 | 2.7101 | 1.0876 | 2.0646 | 2.8943 | 2.4580 | 4.1214 | 3.0444 | 4.0100 | 2.5444 | | 3.6803 | 1.7405 | H16 | 3.4663 | 4.3007 | 3.8068 | 3.8872 | 2.1675 | 1.0847 | 3.3444 | 2.0719 | 2.3188 | 4.9097 | 2.5159 | 4.1385 | 2.4624 | 1.7405 | 3.6803 | | 4.1326 | H17 | 3.4663 | 4.3007 | 3.8068 | 2.1675 | 3.8872 | 3.3444 | 1.0847 | 2.0719 | 2.3188 | 2.5159 | 4.9097 | 2.4624 | 4.1385 | 3.6803 | 1.7405 | 4.1326 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.729 |
|
C1 |
C4 |
H10 |
110.603 |
| C1 |
C4 |
H12 |
109.465 |
|
C1 |
C5 |
C6 |
110.729 |
| C1 |
C5 |
H11 |
110.603 |
|
C1 |
C5 |
H13 |
109.465 |
| H2 |
C1 |
H3 |
106.437 |
|
H2 |
C1 |
C4 |
110.567 |
| H2 |
C1 |
C5 |
110.567 |
|
H3 |
C1 |
C4 |
109.218 |
| H3 |
C1 |
C5 |
109.218 |
|
C4 |
C1 |
C5 |
110.723 |
| C4 |
C7 |
N8 |
113.773 |
|
C4 |
C7 |
H15 |
109.384 |
| C4 |
C7 |
H17 |
110.741 |
|
C5 |
C6 |
N8 |
113.773 |
| C5 |
C6 |
H14 |
109.384 |
|
C5 |
C6 |
H16 |
110.741 |
| C6 |
C5 |
H11 |
110.124 |
|
C6 |
C5 |
H13 |
109.204 |
| C6 |
N8 |
C7 |
112.644 |
|
C6 |
N8 |
H9 |
109.582 |
| C7 |
C4 |
H10 |
110.124 |
|
C7 |
C4 |
H12 |
109.204 |
| C7 |
N8 |
H9 |
109.582 |
|
N8 |
C6 |
H14 |
107.698 |
| N8 |
C6 |
H16 |
108.451 |
|
N8 |
C7 |
H15 |
107.698 |
| N8 |
C7 |
H17 |
108.451 |
|
H10 |
C4 |
H12 |
106.618 |
| H11 |
C5 |
H13 |
106.618 |
|
H14 |
C6 |
H16 |
106.497 |
| H15 |
C7 |
H17 |
106.497 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.425 |
|
|
|
| 2 |
H |
0.211 |
|
|
|
| 3 |
H |
0.200 |
|
|
|
| 4 |
C |
-0.405 |
|
|
|
| 5 |
C |
-0.405 |
|
|
|
| 6 |
C |
-0.263 |
|
|
|
| 7 |
C |
-0.263 |
|
|
|
| 8 |
N |
-0.593 |
|
|
|
| 9 |
H |
0.304 |
|
|
|
| 10 |
H |
0.208 |
|
|
|
| 11 |
H |
0.208 |
|
|
|
| 12 |
H |
0.191 |
|
|
|
| 13 |
H |
0.191 |
|
|
|
| 14 |
H |
0.207 |
|
|
|
| 15 |
H |
0.207 |
|
|
|
| 16 |
H |
0.213 |
|
|
|
| 17 |
H |
0.213 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.280 |
1.219 |
0.000 |
1.250 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.288 |
-0.316 |
0.000 |
| y |
-0.316 |
-44.399 |
0.000 |
| z |
0.000 |
0.000 |
-38.378 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.100 |
-0.316 |
0.000 |
| y |
-0.316 |
-6.566 |
0.000 |
| z |
0.000 |
0.000 |
2.466 |
|
| Polar |
|---|
| 3z2-r2 | 4.932 |
|---|
| x2-y2 | 7.111 |
|---|
| xy | -0.316 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.099 |
0.200 |
0.000 |
| y |
0.200 |
8.305 |
0.000 |
| z |
0.000 |
0.000 |
9.113 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
