Jump to
S1C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -251.209144 |
| Energy at 298.15K | -251.223620 |
| Nuclear repulsion energy | 259.802477 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3562 |
3373 |
0.09 |
|
|
|
| 2 |
A' |
3142 |
2975 |
66.53 |
|
|
|
| 3 |
A' |
3122 |
2957 |
8.29 |
|
|
|
| 4 |
A' |
3121 |
2956 |
47.70 |
|
|
|
| 5 |
A' |
3080 |
2917 |
15.25 |
|
|
|
| 6 |
A' |
3064 |
2902 |
29.15 |
|
|
|
| 7 |
A' |
2969 |
2812 |
139.00 |
|
|
|
| 8 |
A' |
1550 |
1468 |
3.81 |
|
|
|
| 9 |
A' |
1531 |
1450 |
8.59 |
|
|
|
| 10 |
A' |
1523 |
1442 |
10.32 |
|
|
|
| 11 |
A' |
1453 |
1376 |
1.15 |
|
|
|
| 12 |
A' |
1404 |
1330 |
1.00 |
|
|
|
| 13 |
A' |
1341 |
1270 |
0.62 |
|
|
|
| 14 |
A' |
1316 |
1246 |
2.49 |
|
|
|
| 15 |
A' |
1192 |
1129 |
6.08 |
|
|
|
| 16 |
A' |
1089 |
1031 |
6.77 |
|
|
|
| 17 |
A' |
1075 |
1018 |
10.87 |
|
|
|
| 18 |
A' |
934 |
885 |
1.42 |
|
|
|
| 19 |
A' |
893 |
846 |
21.25 |
|
|
|
| 20 |
A' |
851 |
806 |
1.47 |
|
|
|
| 21 |
A' |
805 |
762 |
79.02 |
|
|
|
| 22 |
A' |
557 |
528 |
20.18 |
|
|
|
| 23 |
A' |
432 |
409 |
8.10 |
|
|
|
| 24 |
A' |
418 |
396 |
1.25 |
|
|
|
| 25 |
A' |
258 |
244 |
1.95 |
|
|
|
| 26 |
A" |
3137 |
2971 |
10.41 |
|
|
|
| 27 |
A" |
3121 |
2956 |
67.39 |
|
|
|
| 28 |
A" |
3080 |
2917 |
33.48 |
|
|
|
| 29 |
A" |
2967 |
2810 |
27.83 |
|
|
|
| 30 |
A" |
1537 |
1455 |
0.01 |
|
|
|
| 31 |
A" |
1514 |
1434 |
0.10 |
|
|
|
| 32 |
A" |
1508 |
1428 |
8.99 |
|
|
|
| 33 |
A" |
1408 |
1334 |
0.01 |
|
|
|
| 34 |
A" |
1385 |
1312 |
16.84 |
|
|
|
| 35 |
A" |
1371 |
1299 |
12.79 |
|
|
|
| 36 |
A" |
1321 |
1251 |
1.96 |
|
|
|
| 37 |
A" |
1208 |
1144 |
9.14 |
|
|
|
| 38 |
A" |
1193 |
1130 |
4.96 |
|
|
|
| 39 |
A" |
1176 |
1114 |
6.25 |
|
|
|
| 40 |
A" |
1093 |
1035 |
3.24 |
|
|
|
| 41 |
A" |
997 |
944 |
0.94 |
|
|
|
| 42 |
A" |
910 |
862 |
0.62 |
|
|
|
| 43 |
A" |
832 |
788 |
0.20 |
|
|
|
| 44 |
A" |
448 |
424 |
0.92 |
|
|
|
| 45 |
A" |
259 |
245 |
0.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35572.8 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 33691.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.642 |
1.321 |
0.000 |
| H2 |
0.584 |
2.415 |
0.000 |
| H3 |
1.709 |
1.062 |
0.000 |
| C4 |
-0.012 |
0.746 |
1.256 |
| C5 |
-0.012 |
0.746 |
-1.256 |
| C6 |
-0.012 |
-0.777 |
-1.203 |
| C7 |
-0.012 |
-0.777 |
1.203 |
| N8 |
-0.716 |
-1.217 |
0.000 |
| H9 |
-0.782 |
-2.229 |
0.000 |
| H10 |
0.509 |
1.090 |
2.156 |
| H11 |
0.509 |
1.090 |
-2.156 |
| H12 |
-1.050 |
1.089 |
1.324 |
| H13 |
-1.050 |
1.089 |
-1.324 |
| H14 |
1.034 |
-1.137 |
-1.232 |
| H15 |
1.034 |
-1.137 |
1.232 |
| H16 |
-0.526 |
-1.187 |
-2.077 |
| H17 |
-0.526 |
-1.187 |
2.077 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0952 | 1.0979 | 1.5283 | 1.5283 | 2.5049 | 2.5049 | 2.8781 | 3.8250 | 2.1728 | 2.1728 | 2.1610 | 2.1610 | 2.7775 | 2.7775 | 3.4600 | 3.4600 |
H2 | 1.0952 | | 1.7596 | 2.1716 | 2.1716 | 3.4620 | 3.4620 | 3.8568 | 4.8403 | 2.5320 | 2.5320 | 2.4859 | 2.4859 | 3.7865 | 3.7865 | 4.3036 | 4.3036 | H3 | 1.0979 | 1.7596 | | 2.1539 | 2.1539 | 2.7910 | 2.7910 | 3.3276 | 4.1276 | 2.4678 | 2.4678 | 3.0605 | 3.0605 | 2.6098 | 2.6098 | 3.7910 | 3.7910 | C4 | 1.5283 | 2.1716 | 2.1539 | | 2.5118 | 2.8920 | 1.5236 | 2.4342 | 3.3198 | 1.0956 | 3.4689 | 1.0954 | 2.8017 | 3.2910 | 2.1545 | 3.8873 | 2.1627 | C5 | 1.5283 | 2.1716 | 2.1539 | 2.5118 | | 1.5236 | 2.8920 | 2.4342 | 3.3198 | 3.4689 | 1.0956 | 2.8017 | 1.0954 | 2.1545 | 3.2910 | 2.1627 | 3.8873 | C6 | 2.5049 | 3.4620 | 2.7910 | 2.8920 | 1.5236 | | 2.4056 | 1.4614 | 2.0369 | 3.8778 | 2.1595 | 3.3075 | 2.1381 | 1.1067 | 2.6743 | 1.0944 | 3.3452 | C7 | 2.5049 | 3.4620 | 2.7910 | 1.5236 | 2.8920 | 2.4056 | | 1.4614 | 2.0369 | 2.1595 | 3.8778 | 2.1381 | 3.3075 | 2.6743 | 1.1067 | 3.3452 | 1.0944 | N8 | 2.8781 | 3.8568 | 3.3276 | 2.4342 | 2.4342 | 1.4614 | 1.4614 | | 1.0144 | 3.3866 | 3.3866 | 2.6792 | 2.6792 | 2.1413 | 2.1413 | 2.0858 | 2.0858 | H9 | 3.8250 | 4.8403 | 4.1276 | 3.3198 | 3.3198 | 2.0369 | 2.0369 | 1.0144 | | 4.1629 | 4.1629 | 3.5819 | 3.5819 | 2.4509 | 2.4509 | 2.3375 | 2.3375 | H10 | 2.1728 | 2.5320 | 2.4678 | 1.0956 | 3.4689 | 3.8778 | 2.1595 | 3.3866 | 4.1629 | | 4.3126 | 1.7680 | 3.8134 | 4.0883 | 2.4674 | 4.9171 | 2.5027 | H11 | 2.1728 | 2.5320 | 2.4678 | 3.4689 | 1.0956 | 2.1595 | 3.8778 | 3.3866 | 4.1629 | 4.3126 | | 3.8134 | 1.7680 | 2.4674 | 4.0883 | 2.5027 | 4.9171 | H12 | 2.1610 | 2.4859 | 3.0605 | 1.0954 | 2.8017 | 3.3075 | 2.1381 | 2.6792 | 3.5819 | 1.7680 | 3.8134 | | 2.6472 | 3.9786 | 3.0507 | 4.1253 | 2.4540 | H13 | 2.1610 | 2.4859 | 3.0605 | 2.8017 | 1.0954 | 2.1381 | 3.3075 | 2.6792 | 3.5819 | 3.8134 | 1.7680 | 2.6472 | | 3.0507 | 3.9786 | 2.4540 | 4.1253 | H14 | 2.7775 | 3.7865 | 2.6098 | 3.2910 | 2.1545 | 1.1067 | 2.6743 | 2.1413 | 2.4509 | 4.0883 | 2.4674 | 3.9786 | 3.0507 | | 2.4640 | 1.7752 | 3.6588 | H15 | 2.7775 | 3.7865 | 2.6098 | 2.1545 | 3.2910 | 2.6743 | 1.1067 | 2.1413 | 2.4509 | 2.4674 | 4.0883 | 3.0507 | 3.9786 | 2.4640 | | 3.6588 | 1.7752 | H16 | 3.4600 | 4.3036 | 3.7910 | 3.8873 | 2.1627 | 1.0944 | 3.3452 | 2.0858 | 2.3375 | 4.9171 | 2.5027 | 4.1253 | 2.4540 | 1.7752 | 3.6588 | | 4.1539 | H17 | 3.4600 | 4.3036 | 3.7910 | 2.1627 | 3.8873 | 3.3452 | 1.0944 | 2.0858 | 2.3375 | 2.5027 | 4.9171 | 2.4540 | 4.1253 | 3.6588 | 1.7752 | 4.1539 | |
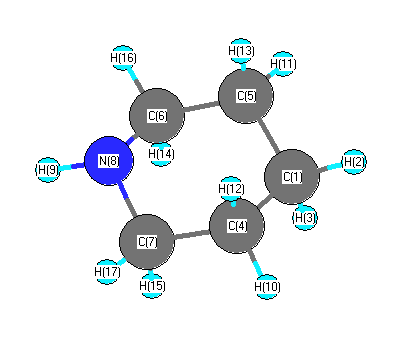 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.321 |
|
C1 |
C4 |
H10 |
110.719 |
| C1 |
C4 |
H12 |
109.806 |
|
C1 |
C5 |
C6 |
110.321 |
| C1 |
C5 |
H11 |
110.719 |
|
C1 |
C5 |
H13 |
109.806 |
| H2 |
C1 |
H3 |
106.706 |
|
H2 |
C1 |
C4 |
110.654 |
| H2 |
C1 |
C5 |
110.654 |
|
H3 |
C1 |
C4 |
109.100 |
| H3 |
C1 |
C5 |
109.100 |
|
C4 |
C1 |
C5 |
110.527 |
| C4 |
C7 |
N8 |
109.250 |
|
C4 |
C7 |
H15 |
108.955 |
| C4 |
C7 |
H17 |
110.321 |
|
C5 |
C6 |
N8 |
109.250 |
| C5 |
C6 |
H14 |
108.955 |
|
C5 |
C6 |
H16 |
110.321 |
| C6 |
C5 |
H11 |
109.992 |
|
C6 |
C5 |
H13 |
108.338 |
| C6 |
N8 |
C7 |
110.781 |
|
C6 |
N8 |
H9 |
109.390 |
| C7 |
C4 |
H10 |
109.992 |
|
C7 |
C4 |
H12 |
108.338 |
| C7 |
N8 |
H9 |
109.390 |
|
N8 |
C6 |
H14 |
112.248 |
| N8 |
C6 |
H16 |
108.540 |
|
N8 |
C7 |
H15 |
112.248 |
| N8 |
C7 |
H17 |
108.540 |
|
H10 |
C4 |
H12 |
107.593 |
| H11 |
C5 |
H13 |
107.593 |
|
H14 |
C6 |
H16 |
107.512 |
| H15 |
C7 |
H17 |
107.512 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -251.207818 |
| Energy at 298.15K | -251.222236 |
| Nuclear repulsion energy | 259.220442 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3537 |
3350 |
0.84 |
|
|
|
| 2 |
A' |
3134 |
2968 |
80.51 |
|
|
|
| 3 |
A' |
3125 |
2960 |
32.08 |
|
|
|
| 4 |
A' |
3115 |
2950 |
29.12 |
|
|
|
| 5 |
A' |
3070 |
2907 |
40.41 |
|
|
|
| 6 |
A' |
3064 |
2902 |
23.11 |
|
|
|
| 7 |
A' |
3060 |
2898 |
28.28 |
|
|
|
| 8 |
A' |
1543 |
1461 |
4.47 |
|
|
|
| 9 |
A' |
1527 |
1446 |
16.82 |
|
|
|
| 10 |
A' |
1523 |
1443 |
2.60 |
|
|
|
| 11 |
A' |
1418 |
1343 |
2.03 |
|
|
|
| 12 |
A' |
1406 |
1332 |
2.16 |
|
|
|
| 13 |
A' |
1368 |
1296 |
0.03 |
|
|
|
| 14 |
A' |
1311 |
1241 |
9.83 |
|
|
|
| 15 |
A' |
1218 |
1153 |
4.29 |
|
|
|
| 16 |
A' |
1074 |
1017 |
0.97 |
|
|
|
| 17 |
A' |
1038 |
983 |
9.65 |
|
|
|
| 18 |
A' |
948 |
898 |
13.29 |
|
|
|
| 19 |
A' |
883 |
836 |
51.29 |
|
|
|
| 20 |
A' |
840 |
796 |
0.16 |
|
|
|
| 21 |
A' |
802 |
760 |
94.93 |
|
|
|
| 22 |
A' |
562 |
532 |
2.23 |
|
|
|
| 23 |
A' |
439 |
416 |
1.01 |
|
|
|
| 24 |
A' |
404 |
383 |
6.30 |
|
|
|
| 25 |
A' |
248 |
235 |
5.03 |
|
|
|
| 26 |
A" |
3132 |
2966 |
19.56 |
|
|
|
| 27 |
A" |
3116 |
2951 |
61.00 |
|
|
|
| 28 |
A" |
3067 |
2905 |
20.23 |
|
|
|
| 29 |
A" |
3059 |
2897 |
37.14 |
|
|
|
| 30 |
A" |
1528 |
1447 |
4.97 |
|
|
|
| 31 |
A" |
1520 |
1440 |
7.40 |
|
|
|
| 32 |
A" |
1505 |
1426 |
7.79 |
|
|
|
| 33 |
A" |
1406 |
1331 |
0.00 |
|
|
|
| 34 |
A" |
1394 |
1320 |
1.15 |
|
|
|
| 35 |
A" |
1370 |
1297 |
1.99 |
|
|
|
| 36 |
A" |
1326 |
1256 |
0.34 |
|
|
|
| 37 |
A" |
1243 |
1177 |
13.63 |
|
|
|
| 38 |
A" |
1167 |
1106 |
15.66 |
|
|
|
| 39 |
A" |
1143 |
1083 |
0.80 |
|
|
|
| 40 |
A" |
1092 |
1034 |
4.36 |
|
|
|
| 41 |
A" |
965 |
914 |
5.40 |
|
|
|
| 42 |
A" |
901 |
853 |
0.36 |
|
|
|
| 43 |
A" |
819 |
775 |
0.02 |
|
|
|
| 44 |
A" |
450 |
427 |
0.92 |
|
|
|
| 45 |
A" |
245 |
232 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35551.5 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 33670.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.648 |
1.317 |
0.000 |
| H2 |
0.593 |
2.411 |
0.000 |
| H3 |
1.715 |
1.056 |
0.000 |
| C4 |
-0.008 |
0.744 |
1.256 |
| C5 |
-0.008 |
0.744 |
-1.256 |
| C6 |
-0.008 |
-0.786 |
-1.206 |
| C7 |
-0.008 |
-0.786 |
1.206 |
| N8 |
-0.618 |
-1.346 |
0.000 |
| H9 |
-1.608 |
-1.117 |
0.000 |
| H10 |
0.507 |
1.093 |
2.158 |
| H11 |
0.507 |
1.093 |
-2.158 |
| H12 |
-1.045 |
1.100 |
1.322 |
| H13 |
-1.045 |
1.100 |
-1.322 |
| H14 |
1.027 |
-1.150 |
-1.245 |
| H15 |
1.027 |
-1.150 |
1.245 |
| H16 |
-0.523 |
-1.205 |
-2.077 |
| H17 |
-0.523 |
-1.205 |
2.077 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0951 | 1.0979 | 1.5287 | 1.5287 | 2.5121 | 2.5121 | 2.9486 | 3.3195 | 2.1746 | 2.1746 | 2.1591 | 2.1591 | 2.7891 | 2.7891 | 3.4708 | 3.4708 |
H2 | 1.0951 | | 1.7593 | 2.1723 | 2.1723 | 3.4697 | 3.4697 | 3.9469 | 4.1586 | 2.5303 | 2.5303 | 2.4798 | 2.4798 | 3.7969 | 3.7969 | 4.3167 | 4.3167 | H3 | 1.0979 | 1.7593 | | 2.1545 | 2.1545 | 2.7955 | 2.7955 | 3.3476 | 3.9703 | 2.4737 | 2.4737 | 3.0600 | 3.0600 | 2.6243 | 2.6243 | 3.7987 | 3.7987 | C4 | 1.5287 | 2.1723 | 2.1545 | | 2.5115 | 2.8987 | 1.5305 | 2.5126 | 2.7568 | 1.0965 | 3.4705 | 1.0981 | 2.8014 | 3.3030 | 2.1575 | 3.8947 | 2.1762 | C5 | 1.5287 | 2.1723 | 2.1545 | 2.5115 | | 1.5305 | 2.8987 | 2.5126 | 2.7568 | 3.4705 | 1.0965 | 2.8014 | 1.0981 | 2.1575 | 3.3030 | 2.1762 | 3.8947 | C6 | 2.5121 | 3.4697 | 2.7955 | 2.8987 | 1.5305 | | 2.4130 | 1.4631 | 2.0313 | 3.8884 | 2.1688 | 3.3205 | 2.1552 | 1.0974 | 2.6859 | 1.0947 | 3.3499 | C7 | 2.5121 | 3.4697 | 2.7955 | 1.5305 | 2.8987 | 2.4130 | | 1.4631 | 2.0313 | 2.1688 | 3.8884 | 2.1552 | 3.3205 | 2.6859 | 1.0974 | 3.3499 | 1.0947 | N8 | 2.9486 | 3.9469 | 3.3476 | 2.5126 | 2.5126 | 1.4631 | 1.4631 | | 1.0163 | 3.4456 | 3.4456 | 2.8125 | 2.8125 | 2.0723 | 2.0723 | 2.0840 | 2.0840 | H9 | 3.3195 | 4.1586 | 3.9703 | 2.7568 | 2.7568 | 2.0313 | 2.0313 | 1.0163 | | 3.7442 | 3.7442 | 2.6422 | 2.6422 | 2.9147 | 2.9147 | 2.3452 | 2.3452 | H10 | 2.1746 | 2.5303 | 2.4737 | 1.0965 | 3.4705 | 3.8884 | 2.1688 | 3.4456 | 3.7442 | | 4.3169 | 1.7624 | 3.8108 | 4.1093 | 2.4769 | 4.9276 | 2.5196 | H11 | 2.1746 | 2.5303 | 2.4737 | 3.4705 | 1.0965 | 2.1688 | 3.8884 | 3.4456 | 3.7442 | 4.3169 | | 3.8108 | 1.7624 | 2.4769 | 4.1093 | 2.5196 | 4.9276 | H12 | 2.1591 | 2.4798 | 3.0600 | 1.0981 | 2.8014 | 3.3205 | 2.1552 | 2.8125 | 2.6422 | 1.7624 | 3.8108 | | 2.6445 | 3.9928 | 3.0588 | 4.1399 | 2.4806 | H13 | 2.1591 | 2.4798 | 3.0600 | 2.8014 | 1.0981 | 2.1552 | 3.3205 | 2.8125 | 2.6422 | 3.8108 | 1.7624 | 2.6445 | | 3.0588 | 3.9928 | 2.4806 | 4.1399 | H14 | 2.7891 | 3.7969 | 2.6243 | 3.3030 | 2.1575 | 1.0974 | 2.6859 | 2.0723 | 2.9147 | 4.1093 | 2.4769 | 3.9928 | 3.0588 | | 2.4905 | 1.7596 | 3.6663 | H15 | 2.7891 | 3.7969 | 2.6243 | 2.1575 | 3.3030 | 2.6859 | 1.0974 | 2.0723 | 2.9147 | 2.4769 | 4.1093 | 3.0588 | 3.9928 | 2.4905 | | 3.6663 | 1.7596 | H16 | 3.4708 | 4.3167 | 3.7987 | 3.8947 | 2.1762 | 1.0947 | 3.3499 | 2.0840 | 2.3452 | 4.9276 | 2.5196 | 4.1399 | 2.4806 | 1.7596 | 3.6663 | | 4.1541 | H17 | 3.4708 | 4.3167 | 3.7987 | 2.1762 | 3.8947 | 3.3499 | 1.0947 | 2.0840 | 2.3452 | 2.5196 | 4.9276 | 2.4806 | 4.1399 | 3.6663 | 1.7596 | 4.1541 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.403 |
|
C1 |
C4 |
H10 |
110.789 |
| C1 |
C4 |
H12 |
109.469 |
|
C1 |
C5 |
C6 |
110.403 |
| C1 |
C5 |
H11 |
110.789 |
|
C1 |
C5 |
H13 |
109.469 |
| H2 |
C1 |
H3 |
106.681 |
|
H2 |
C1 |
C4 |
110.679 |
| H2 |
C1 |
C5 |
110.679 |
|
H3 |
C1 |
C4 |
109.121 |
| H3 |
C1 |
C5 |
109.121 |
|
C4 |
C1 |
C5 |
110.462 |
| C4 |
C7 |
N8 |
114.122 |
|
C4 |
C7 |
H15 |
109.265 |
| C4 |
C7 |
H17 |
110.895 |
|
C5 |
C6 |
N8 |
114.122 |
| C5 |
C6 |
H14 |
109.265 |
|
C5 |
C6 |
H16 |
110.895 |
| C6 |
C5 |
H11 |
110.201 |
|
C6 |
C5 |
H13 |
109.042 |
| C6 |
N8 |
C7 |
111.098 |
|
C6 |
N8 |
H9 |
108.685 |
| C7 |
C4 |
H10 |
110.201 |
|
C7 |
C4 |
H12 |
109.042 |
| C7 |
N8 |
H9 |
108.685 |
|
N8 |
C6 |
H14 |
107.196 |
| N8 |
C6 |
H16 |
108.268 |
|
N8 |
C7 |
H15 |
107.196 |
| N8 |
C7 |
H17 |
108.268 |
|
H10 |
C4 |
H12 |
106.854 |
| H11 |
C5 |
H13 |
106.854 |
|
H14 |
C6 |
H16 |
106.779 |
| H15 |
C7 |
H17 |
106.779 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
