return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for SiH2(CH3)2 (dimethylsilane)

using model chemistry: CISD/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at CISD/6-311G*
 hartrees
Energy at 0K-369.720553
Energy at 298.15K-369.728555
Nuclear repulsion energy113.963747
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CISD/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3181 2944 17.28      
2 A1 3102 2870 5.64      
3 A1 2257 2088 127.91      
4 A1 1533 1419 5.35      
5 A1 1383 1280 17.57      
6 A1 1004 929 132.08      
7 A1 908 840 32.04      
8 A1 680 629 4.90      
9 A1 214 198 1.43      
10 A2 3177 2939 0.00      
11 A2 1525 1411 0.00      
12 A2 931 862 0.00      
13 A2 613 567 0.00      
14 A2 150 139 0.00      
15 B1 3178 2941 29.87      
16 B1 2248 2081 179.20      
17 B1 1538 1423 14.13      
18 B1 934 865 89.52      
19 B1 482 446 16.40      
20 B1 167 155 0.00      
21 B2 3181 2943 10.18      
22 B2 3100 2869 9.90      
23 B2 1527 1413 3.16      
24 B2 1376 1274 43.16      
25 B2 972 899 271.46      
26 B2 752 696 13.49      
27 B2 677 627 16.52      

Unscaled Zero Point Vibrational Energy (zpe) 20394.2 cm-1
Scaled (by 0.9253) Zero Point Vibrational Energy (zpe) 18870.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CISD/6-311G*
ABC
0.55704 0.19627 0.16192

See section I.F.4 to change rotational constant units
Geometric Data calculated at CISD/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Si1 0.000 0.000 0.551
H2 -1.199 0.000 1.428
H3 1.199 0.000 1.428
C4 0.000 1.549 -0.512
C5 0.000 -1.549 -0.512
H6 0.000 2.453 0.099
H7 0.000 -2.453 0.099
H8 0.880 1.588 -1.156
H9 -0.880 1.588 -1.156
H10 -0.880 -1.588 -1.156
H11 0.880 -1.588 -1.156

Atom - Atom Distances (Å)
  Si1 H2 H3 C4 C5 H6 H7 H8 H9 H10 H11
Si11.48571.48571.87821.87822.49402.49402.49162.49162.49162.4916
H21.48572.39842.75662.75663.03643.03643.67693.04923.04923.6769
H31.48572.39842.75662.75663.03643.03643.04923.67693.67693.0492
C41.87822.75662.75663.09771.09084.04791.09131.09133.32123.3212
C51.87822.75662.75663.09774.04791.09083.32123.32121.09131.0913
H62.49403.03643.03641.09084.04794.90551.75961.75964.32174.3217
H72.49403.03643.03644.04791.09084.90554.32174.32171.75961.7596
H82.49163.67693.04921.09133.32121.75964.32171.76043.63153.1763
H92.49163.04923.67691.09133.32121.75964.32171.76043.17633.6315
H102.49163.04923.67693.32121.09134.32171.75963.63153.17631.7604
H112.49163.67693.04923.32121.09134.32171.75963.17633.63151.7604

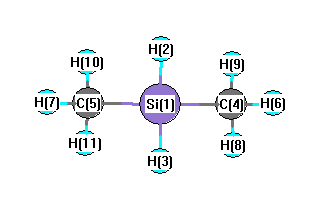
picture of dimethylsilane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Si1 C4 H6 111.512 Si1 C4 H8 111.310
Si1 C4 H9 111.310 Si1 C5 H7 111.512
Si1 C5 H10 111.310 Si1 C5 H11 111.310
H2 Si1 H3 107.637 H2 Si1 C4 109.507
H2 Si1 C5 109.507 H3 Si1 C4 109.507
H3 Si1 C5 109.507 C4 Si1 C5 111.107
H6 C4 H8 107.487 H6 C4 H9 107.487
H7 C5 H10 107.487 H7 C5 H11 107.487
H8 C4 H9 107.526 H10 C5 H11 107.526
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability