return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6OS (Vinyl sulfoxide)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-628.367425
Energy at 298.15K-628.373686
Nuclear repulsion energy291.010427
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3294 3120 0.50      
2 A' 3220 3050 5.32      
3 A' 3184 3016 2.44      
4 A' 1658 1570 3.39      
5 A' 1424 1349 17.18      
6 A' 1291 1223 0.62      
7 A' 1114 1055 98.14      
8 A' 1022 968 8.33      
9 A' 997 944 46.08      
10 A' 922 873 28.15      
11 A' 706 668 75.62      
12 A' 651 617 7.53      
13 A' 518 490 0.73      
14 A' 307 290 2.63      
15 A' 228 216 2.58      
16 A' 100 95 0.70      
17 A" 3294 3120 0.50      
18 A" 3216 3046 2.99      
19 A" 3184 3015 2.73      
20 A" 1650 1563 4.38      
21 A" 1420 1344 4.20      
22 A" 1274 1207 5.82      
23 A" 1008 954 0.01      
24 A" 985 933 55.24      
25 A" 915 866 15.66      
26 A" 659 624 15.29      
27 A" 558 528 7.89      
28 A" 486 461 9.00      
29 A" 271 257 8.09      
30 A" 159 151 0.94      

Unscaled Zero Point Vibrational Energy (zpe) 19857.2 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 18806.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.15720 0.07777 0.06932

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.591 -0.566 0.000
O2 1.346 0.731 0.000
C3 -0.631 -0.467 1.311
C4 -0.631 -0.467 -1.311
C5 -0.631 0.605 2.105
C6 -0.631 0.605 -2.105
H7 -1.282 -1.330 1.422
H8 -1.282 -1.330 -1.422
H9 -1.329 0.697 2.929
H10 -1.329 0.697 -2.929
H11 0.077 1.407 1.921
H12 0.077 1.407 -1.921

Atom - Atom Distances (Å)
  S1 O2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
S11.50091.79491.79492.70072.70072.47252.47253.72263.72262.80142.8014
O21.50092.65792.65792.89072.89073.63033.63033.96703.96702.40002.4000
C31.79492.65792.62221.33373.57991.08712.93892.11144.45192.09413.8024
C41.79492.65792.62223.57991.33372.93891.08714.45192.11143.80242.0941
C52.70072.89071.33373.57994.20912.15324.07471.08385.08251.08554.1650
C62.70072.89073.57991.33374.20914.07472.15325.08251.08384.16501.0855
H72.47253.63031.08712.93892.15324.07472.84322.52654.79993.09694.5291
H82.47253.63032.93891.08714.07472.15322.84324.79992.52654.52913.0969
H93.72263.96702.11144.45191.08385.08252.52654.79995.85811.86995.0991
H103.72263.96704.45192.11145.08251.08384.79992.52655.85815.09911.8699
H112.80142.40002.09413.80241.08554.16503.09694.52911.86995.09913.8415
H122.80142.40003.80242.09414.16501.08554.52913.09695.09911.86993.8415

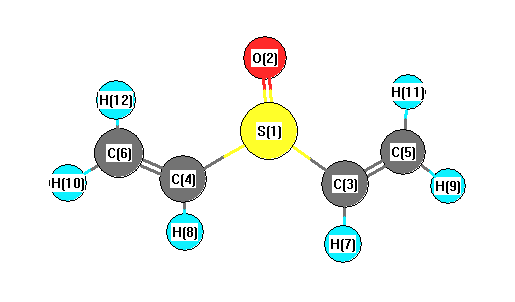
picture of Vinyl sulfoxide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C3 C5 118.618 S1 C3 H7 115.982
S1 C4 C6 118.618 S1 C4 H8 115.982
O2 S1 C3 107.162 O2 S1 C4 107.162
C3 S1 C4 93.847 C3 C5 H9 121.364
C3 C5 H11 119.555 C4 C6 H10 121.364
C4 C6 H12 119.555 C5 C3 H7 125.308
C6 C4 H8 125.308 H9 C5 H11 119.080
H10 C6 H12 119.080
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability