return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (Cyclobutane, methylene-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-194.744926
Energy at 298.15K-194.753445
HF Energy-193.975002
Nuclear repulsion energy167.905236
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3182 3014 40.16      
2 A' 3176 3008 6.67      
3 A' 3153 2986 11.39      
4 A' 3111 2947 39.96      
5 A' 3089 2926 10.35      
6 A' 1754 1661 14.40      
7 A' 1541 1459 0.22      
8 A' 1515 1435 4.53      
9 A' 1455 1378 1.46      
10 A' 1272 1205 1.72      
11 A' 1239 1174 0.07      
12 A' 1099 1041 0.96      
13 A' 998 946 0.67      
14 A' 915 866 0.19      
15 A' 850 805 49.63      
16 A' 764 724 2.54      
17 A' 670 635 1.79      
18 A' 379 359 6.27      
19 A' 125 119 0.37      
20 A" 3272 3099 10.20      
21 A" 3157 2990 18.22      
22 A" 3087 2923 39.84      
23 A" 1500 1420 6.00      
24 A" 1295 1226 0.04      
25 A" 1265 1198 0.34      
26 A" 1215 1151 1.26      
27 A" 1197 1134 2.68      
28 A" 1059 1003 0.24      
29 A" 949 898 1.81      
30 A" 926 877 0.34      
31 A" 827 783 0.20      
32 A" 661 626 0.12      
33 A" 351 332 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 25523.2 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 24173.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.34294 0.15500 0.11767

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.245 -0.412 1.086
C2 0.245 -0.412 -1.086
C3 0.000 0.621 0.000
C4 -0.013 -1.494 0.000
C5 -0.477 1.866 0.000
H6 -0.402 -0.392 1.966
H7 -0.402 -0.392 -1.966
H8 1.291 -0.416 1.410
H9 1.291 -0.416 -1.410
H10 -0.676 2.393 0.928
H11 -0.676 2.393 -0.928
H12 0.633 -2.375 0.000
H13 -1.058 -1.810 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C12.17151.51861.55392.62531.09283.11971.09542.70662.95723.57422.27582.1979
C22.17151.51861.55392.62533.11971.09282.70661.09543.57422.95722.27582.1979
C31.51861.51862.11451.33372.24782.24782.17492.17492.11192.11193.06142.6514
C41.55391.55392.11453.39192.28702.28702.20262.20264.05104.05101.09201.0921
C52.62532.62531.33373.39192.99512.99513.21283.21281.08561.08564.38363.7223
H61.09283.11972.24782.28702.99513.93191.78253.77742.98524.02592.97772.5112
H73.11971.09282.24782.28702.99513.93193.77741.78254.02592.98522.97772.5112
H81.09542.70662.17492.20263.21281.78253.77742.82093.46314.15062.50203.0745
H92.70661.09542.17492.20263.21283.77741.78252.82094.15063.46312.50203.0745
H102.95723.57422.11194.05101.08562.98524.02593.46314.15061.85585.03064.3219
H113.57422.95722.11194.05101.08564.02592.98524.15063.46311.85585.03064.3219
H122.27582.27583.06141.09204.38362.97772.97772.50202.50205.03065.03061.7823
H132.19792.19792.65141.09213.72232.51122.51123.07453.07454.32194.32191.7823

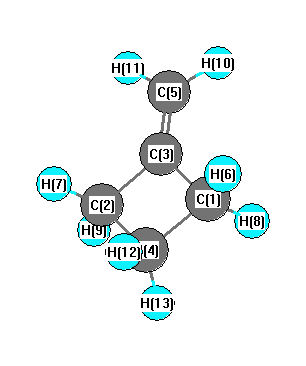
picture of Cyclobutane, methylene- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C2 91.283 C1 C3 C5 133.866
C1 C4 C2 88.651 C1 C4 H12 117.602
C1 C4 H13 111.131 C2 C3 C5 133.866
C2 C4 H12 117.602 C2 C4 H13 111.131
C3 C1 C4 86.968 C3 C1 H6 117.887
C3 C1 H8 111.590 C3 C2 C4 86.968
C3 C2 H7 117.887 C3 C2 H9 111.590
C3 C5 H10 121.267 C3 C5 H11 121.267
C4 C1 H6 118.520 C4 C1 H8 111.308
C4 C2 H7 118.520 C4 C2 H9 111.308
H6 C1 H8 109.093 H7 C2 H9 109.093
H10 C5 H11 117.466 H12 C4 H13 109.375
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability