return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6S (Thiophene, 2,3-dihydro-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-553.368636
Energy at 298.15K-553.375620
HF Energy-552.500951
Nuclear repulsion energy217.590039
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3257 3085 8.10      
2 A 3231 3060 0.56      
3 A 3184 3015 9.75      
4 A 3139 2973 19.89      
5 A 3102 2938 20.58      
6 A 3055 2894 33.06      
7 A 1621 1535 8.98      
8 A 1521 1441 2.03      
9 A 1504 1424 4.34      
10 A 1364 1292 2.01      
11 A 1360 1288 5.16      
12 A 1309 1240 20.21      
13 A 1227 1162 2.60      
14 A 1180 1117 0.26      
15 A 1132 1072 1.85      
16 A 1046 991 6.90      
17 A 1031 976 3.73      
18 A 950 900 2.99      
19 A 900 852 0.36      
20 A 884 838 2.54      
21 A 815 772 24.79      
22 A 711 674 3.42      
23 A 686 650 23.32      
24 A 650 616 45.07      
25 A 506 479 1.20      
26 A 412 390 2.60      
27 A 206 195 1.18      

Unscaled Zero Point Vibrational Energy (zpe) 19991.3 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 18933.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.23097 0.15689 0.09939

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.063 -1.240 0.249
H2 -0.085 -1.416 1.328
H3 0.146 -2.175 -0.271
C4 -1.366 -0.580 -0.207
H5 -2.225 -1.010 0.317
H6 -1.517 -0.736 -1.284
C7 0.105 1.272 0.091
H8 0.473 2.286 0.198
C9 -1.183 0.891 0.067
H10 -2.011 1.588 0.144
S11 1.267 -0.037 -0.102

Atom - Atom Distances (Å)
  C1 H2 H3 C4 H5 H6 C7 H8 C9 H10 S11
C11.09311.09071.52942.17472.17192.52183.56622.41433.43551.8271
H21.09311.78502.16612.40093.05492.96463.91042.84953.75992.4030
H31.09071.78502.19812.70572.42073.46604.49743.35914.35742.4200
C41.52942.16612.19811.09351.09912.38363.42921.50812.28962.6900
H52.17472.40092.70571.09351.77153.26864.26072.18252.61293.6486
H62.17193.05492.42071.09911.77152.92423.90972.14122.77233.1035
C72.52182.96463.46602.38363.26862.92421.08411.34302.14021.7609
H83.56623.91044.49743.42924.26073.90971.08412.16872.58072.4731
C92.41432.84953.35911.50812.18252.14121.34302.16871.08502.6253
H103.43553.75994.35742.28962.61292.77232.14022.58071.08503.6671
S111.82712.40302.42002.69003.64863.10351.76092.47312.62533.6671

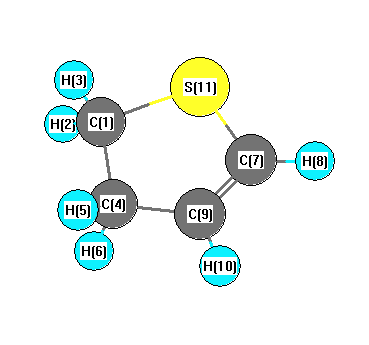
picture of Thiophene, 2,3-dihydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H5 110.918 C1 C4 H6 110.363
C1 C4 C9 105.273 C1 S11 C7 89.296
H2 C1 H3 109.648 H2 C1 C4 110.264
H2 C1 S11 108.108 H3 C1 C4 112.986
H3 C1 S11 109.475 C4 C1 S11 106.198
C4 C9 C7 113.315 C4 C9 H10 123.172
H5 C4 H6 107.789 H5 C4 C9 113.077
H6 C4 C9 109.422 C7 C9 H10 123.282
H8 C7 C9 126.316 H8 C7 S11 118.817
C9 C7 S11 114.857
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability