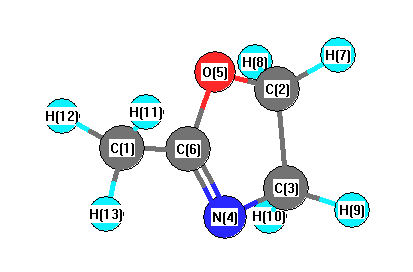
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3313 |
2996 |
14.78 |
|
|
|
| 2 |
A |
3238 |
2928 |
71.50 |
|
|
|
| 3 |
A |
3210 |
2903 |
64.38 |
|
|
|
| 4 |
A |
3205 |
2898 |
19.32 |
|
|
|
| 5 |
A |
1930 |
1745 |
231.71 |
|
|
|
| 6 |
A |
1677 |
1517 |
1.92 |
|
|
|
| 7 |
A |
1650 |
1492 |
0.22 |
|
|
|
| 8 |
A |
1611 |
1457 |
8.46 |
|
|
|
| 9 |
A |
1563 |
1414 |
53.59 |
|
|
|
| 10 |
A |
1522 |
1376 |
6.39 |
|
|
|
| 11 |
A |
1481 |
1339 |
24.64 |
|
|
|
| 12 |
A |
1371 |
1240 |
139.44 |
|
|
|
| 13 |
A |
1197 |
1083 |
8.05 |
|
|
|
| 14 |
A |
1143 |
1033 |
111.31 |
|
|
|
| 15 |
A |
1083 |
979 |
0.54 |
|
|
|
| 16 |
A |
1035 |
936 |
38.37 |
|
|
|
| 17 |
A |
994 |
899 |
17.21 |
|
|
|
| 18 |
A |
830 |
751 |
7.15 |
|
|
|
| 19 |
A |
703 |
636 |
3.32 |
|
|
|
| 20 |
A |
372 |
337 |
6.38 |
|
|
|
| 21 |
A |
3282 |
2969 |
75.93 |
|
|
|
| 22 |
A |
3259 |
2948 |
21.17 |
|
|
|
| 23 |
A |
3235 |
2926 |
18.85 |
|
|
|
| 24 |
A |
1609 |
1455 |
8.25 |
|
|
|
| 25 |
A |
1369 |
1238 |
0.00 |
|
|
|
| 26 |
A |
1340 |
1212 |
4.38 |
|
|
|
| 27 |
A |
1234 |
1116 |
0.14 |
|
|
|
| 28 |
A |
1169 |
1057 |
2.73 |
|
|
|
| 29 |
A |
902 |
816 |
1.71 |
|
|
|
| 30 |
A |
662 |
599 |
9.47 |
|
|
|
| 31 |
A |
259 |
234 |
11.43 |
|
|
|
| 32 |
A |
157 |
142 |
1.02 |
|
|
|
| 33 |
A |
77 |
69 |
0.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25839.7 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 23369.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.660 |
-0.563 |
|
-0.724 |
| 2 |
C |
-0.195 |
0.029 |
|
0.083 |
| 3 |
C |
-0.337 |
0.457 |
|
0.273 |
| 4 |
N |
-0.431 |
-0.799 |
|
-0.765 |
| 5 |
O |
-0.438 |
-0.469 |
|
-0.483 |
| 6 |
C |
0.479 |
0.889 |
|
0.934 |
| 7 |
H |
0.214 |
0.050 |
|
0.047 |
| 8 |
H |
0.214 |
0.050 |
|
0.047 |
| 9 |
H |
0.218 |
-0.046 |
|
0.007 |
| 10 |
H |
0.218 |
-0.046 |
|
0.007 |
| 11 |
H |
0.235 |
0.148 |
|
0.190 |
| 12 |
H |
0.235 |
0.148 |
|
0.190 |
| 13 |
H |
0.250 |
0.153 |
|
0.193 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.012 |
-1.009 |
0.000 |
1.429 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.995 |
-1.077 |
0.000 |
1.466 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.830 |
1.914 |
0.000 |
| y |
1.914 |
-29.748 |
0.000 |
| z |
0.000 |
0.000 |
-35.279 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.316 |
1.914 |
0.000 |
| y |
1.914 |
8.806 |
0.000 |
| z |
0.000 |
0.000 |
0.510 |
|
| Polar |
|---|
| 3z2-r2 | 1.019 |
|---|
| x2-y2 | -12.082 |
|---|
| xy | 1.914 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.171 |
0.736 |
0.000 |
| y |
0.736 |
8.323 |
0.000 |
| z |
0.000 |
0.000 |
5.547 |
<r2> (average value of r
2) Å
2
| <r2> |
146.511 |
| (<r2>)1/2 |
12.104 |