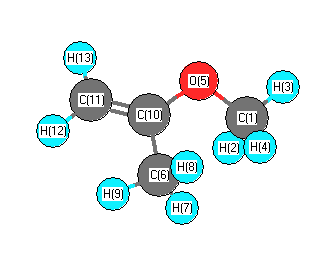
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3261 |
3140 |
10.10 |
|
|
|
| 2 |
A |
3168 |
3050 |
1.54 |
|
|
|
| 3 |
A |
3145 |
3028 |
8.58 |
|
|
|
| 4 |
A |
3143 |
3026 |
37.16 |
|
|
|
| 5 |
A |
3090 |
2975 |
21.98 |
|
|
|
| 6 |
A |
3076 |
2962 |
51.82 |
|
|
|
| 7 |
A |
3037 |
2923 |
26.76 |
|
|
|
| 8 |
A |
3012 |
2900 |
58.32 |
|
|
|
| 9 |
A |
1729 |
1664 |
120.92 |
|
|
|
| 10 |
A |
1523 |
1466 |
10.79 |
|
|
|
| 11 |
A |
1515 |
1458 |
10.40 |
|
|
|
| 12 |
A |
1502 |
1446 |
12.10 |
|
|
|
| 13 |
A |
1491 |
1435 |
6.53 |
|
|
|
| 14 |
A |
1490 |
1435 |
10.49 |
|
|
|
| 15 |
A |
1435 |
1381 |
12.01 |
|
|
|
| 16 |
A |
1410 |
1358 |
29.68 |
|
|
|
| 17 |
A |
1315 |
1266 |
181.65 |
|
|
|
| 18 |
A |
1215 |
1170 |
0.72 |
|
|
|
| 19 |
A |
1179 |
1135 |
4.63 |
|
|
|
| 20 |
A |
1141 |
1098 |
107.18 |
|
|
|
| 21 |
A |
1070 |
1030 |
2.03 |
|
|
|
| 22 |
A |
1022 |
984 |
4.01 |
|
|
|
| 23 |
A |
938 |
903 |
6.24 |
|
|
|
| 24 |
A |
842 |
811 |
67.58 |
|
|
|
| 25 |
A |
818 |
787 |
9.08 |
|
|
|
| 26 |
A |
742 |
715 |
4.32 |
|
|
|
| 27 |
A |
539 |
519 |
7.77 |
|
|
|
| 28 |
A |
481 |
463 |
3.11 |
|
|
|
| 29 |
A |
424 |
408 |
0.97 |
|
|
|
| 30 |
A |
308 |
297 |
1.24 |
|
|
|
| 31 |
A |
247 |
238 |
0.80 |
|
|
|
| 32 |
A |
198 |
191 |
1.71 |
|
|
|
| 33 |
A |
88 |
85 |
7.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24796.2 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 23871.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.481 |
|
|
|
| 2 |
H |
0.213 |
|
|
|
| 3 |
H |
0.233 |
|
|
|
| 4 |
H |
0.216 |
|
|
|
| 5 |
O |
-0.326 |
|
|
|
| 6 |
C |
-0.748 |
|
|
|
| 7 |
H |
0.238 |
|
|
|
| 8 |
H |
0.244 |
|
|
|
| 9 |
H |
0.241 |
|
|
|
| 10 |
C |
0.273 |
|
|
|
| 11 |
C |
-0.536 |
|
|
|
| 12 |
H |
0.211 |
|
|
|
| 13 |
H |
0.222 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.922 |
1.595 |
0.538 |
1.919 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.103 |
-0.176 |
0.366 |
| y |
-0.176 |
-31.501 |
-0.507 |
| z |
0.366 |
-0.507 |
-33.433 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.364 |
-0.176 |
0.366 |
| y |
-0.176 |
-0.733 |
-0.507 |
| z |
0.366 |
-0.507 |
-3.631 |
|
| Polar |
|---|
| 3z2-r2 | -7.262 |
|---|
| x2-y2 | 3.398 |
|---|
| xy | -0.176 |
|---|
| xz | 0.366 |
|---|
| yz | -0.507 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
128.355 |
| (<r2>)1/2 |
11.329 |