return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O (Furan, 2,3-dihydro-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-230.658374
Energy at 298.15K-230.665707
HF Energy-229.834360
Nuclear repulsion energy174.712085
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3292 3117 6.76      
2 A 3265 3092 2.48      
3 A 3189 3020 28.44      
4 A 3147 2981 19.99      
5 A 3086 2923 49.34      
6 A 3074 2911 34.72      
7 A 1666 1578 44.99      
8 A 1551 1469 1.51      
9 A 1530 1449 1.35      
10 A 1429 1353 1.93      
11 A 1356 1284 2.93      
12 A 1322 1252 7.84      
13 A 1267 1200 14.09      
14 A 1239 1173 21.65      
15 A 1180 1117 63.81      
16 A 1114 1055 21.63      
17 A 1091 1033 19.69      
18 A 1034 979 8.29      
19 A 972 920 14.00      
20 A 965 914 32.78      
21 A 898 851 5.95      
22 A 872 826 7.13      
23 A 843 799 7.23      
24 A 717 679 44.74      
25 A 661 626 1.06      
26 A 460 436 25.50      
27 A 190 180 1.85      

Unscaled Zero Point Vibrational Energy (zpe) 20704.0 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 19608.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.27306 0.26011 0.14231

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 0.771 2.184 0.146
C2 0.418 1.164 0.082
H3 2.271 -0.021 0.037
C4 1.192 0.072 0.013
O5 0.522 -1.109 -0.125
H6 -1.359 0.940 -1.133
H7 -1.720 1.174 0.584
C8 -1.014 0.721 -0.116
H9 -1.475 -1.419 -0.506
H10 -1.062 -1.020 1.187
C11 -0.864 -0.785 0.135

Atom - Atom Distances (Å)
  H1 C2 H3 C4 O5 H6 H7 C8 H9 H10 C11
H11.08112.66922.15783.31402.77852.72342.32194.29553.83543.3895
C21.08112.19961.34022.28532.16492.19651.51173.25642.86052.3338
H32.66922.19961.08302.06603.93294.20183.37074.03533.66473.2283
C42.15781.34021.08301.36512.92863.16552.30293.09972.76602.2310
O53.31402.28532.06601.36512.95893.27792.38982.05722.05901.4475
H62.77852.16493.93292.92862.95891.77061.09682.44323.05152.1974
H72.72342.19654.20183.16553.27791.77061.09252.82332.36842.1844
C82.32191.51173.37072.30292.38981.09681.09252.22402.17511.5344
H94.29553.25644.03533.09972.05722.44322.82332.22401.78771.0894
H103.83542.86053.66472.76602.05903.05152.36842.17511.78771.0956
C113.38952.33383.22832.23101.44752.19742.18441.53441.08941.0956

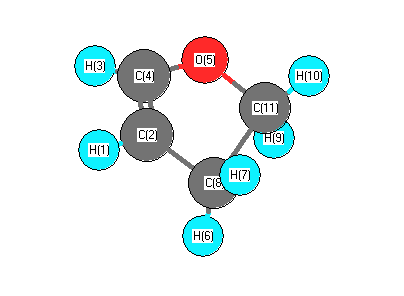
picture of Furan, 2,3-dihydro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 125.709 H1 C2 C8 126.354
C2 C4 H3 130.085 C2 C4 O5 115.287
C2 C8 H6 111.186 C2 C8 H7 114.047
C2 C8 C11 100.016 H3 C4 O5 114.621
C4 C2 C8 107.549 C4 O5 C11 104.934
O5 C11 C8 106.496 O5 C11 H9 107.532
O5 C11 H10 107.312 H6 C8 H7 107.952
H6 C8 C11 112.191 H7 C8 C11 111.406
C8 C11 H9 114.854 C8 C11 H10 110.481
H9 C11 H10 109.808
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability